


Product Monograph - Comirnaty Omicron Xbb.1.5
RECENT MAJOR LABEL CHANGES
TABLE OF CONTENTS
Sections or subsections that are not applicable at the time of authorization are not listed.
RECENT MAJOR LABEL CHANGES .......................................................................................... 2
TABLE OF CONTENTS ............................................................................................................ 2
PART I: HEALTH PROFESSIONAL INFORMATION .................................................................... 6
1 INDICATIONS ........................................................................................................... 6
1.1 Pediatrics ..................................................................................................................... 6
1.2 Geriatrics ..................................................................................................................... 6
2 CONTRAINDICATIONS .............................................................................................. 6
3 SERIOUS WARNINGS AND PRECAUTIONS ................................................................. 6
4 DOSAGE AND ADMINISTRATION.............................................................................. 6
4.1 Dosing Considerations ................................................................................................ 6
4.2 Recommended Dose and Dosage Adjustment ........................................................... 8
4.2.1 Vaccination Schedule for Individuals 12 Years of Age and Older .......................... 8
4.2.2 Vaccination Schedule for Individuals Aged 5 Years to <12 Years .......................... 8
4.2.3 Vaccination Schedule for Individuals Aged 6 Months to <5 Years ......................... 8
4.3 Reconstitution ............................................................................................................. 9
4.3.1 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Gray Cap and Gray Label Border) .................................................................... 9
4.3.2 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border) .......................................................................... 13
Imported and distributed by: Pfizer Canada ULC 17,300 Trans-Canada Highway Kirkland, Quebec, Canada H9J 2M5 |
4.3.3 For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border)................................................................... 18
4.3.4 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) ....................................................................... 21
4.4 Administration .......................................................................................................... 26
4.4.1 For 12 Years of Age and Older .............................................................................. 26
4.4.2 For Age 5 Years to <12 Years ................................................................................ 27
4.4.3 For Age 6 Months to <5 Years .............................................................................. 28
5 OVERDOSAGE ........................................................................................................ 29
6 DOSAGE FORMS, STRENGTHS, COMPOSITION AND PACKAGING ............................ 29
7 WARNINGS AND PRECAUTIONS ............................................................................. 32
7.1 Special Populations ................................................................................................... 33
7.1.1 Pregnant Women ................................................................................................. 33
7.1.2 Breast-feeding ...................................................................................................... 34
7.1.3 Pediatrics .............................................................................................................. 34
7.1.4 Geriatrics .............................................................................................................. 34
8 ADVERSE REACTIONS ............................................................................................ 34
8.1 Adverse Reaction Overview ...................................................................................... 34
8.1.1 COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg) ............................ 34
8.1.2 COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg) ................................ 35
8.1.3 COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) .......................... 35
8.1.4 COMIRNATY (30 mcg) ........................................................................................... 36
8.1.5 COMIRNATY (10 mcg) ........................................................................................... 37
8.1.6 COMIRNATY (3 mcg) ............................................................................................. 38
8.2 Clinical Trial Adverse Reactions ................................................................................ 38
8.2.1 COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg) ............................ 39
8.2.2 COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg) ................................ 41
8.2.3 COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) .......................... 43
8.2.4 COMIRNATY (30 mcg) ........................................................................................... 47
8.2.5 COMIRNATY (10 mcg) ........................................................................................... 56
8.2.6 COMIRNATY (3 mcg) ............................................................................................. 61
8.5 Post-Market Adverse Reactions ................................................................................ 65
9 DRUG INTERACTIONS ............................................................................................ 65
10 CLINICAL PHARMACOLOGY .................................................................................... 65
10.1 Mechanism of Action ................................................................................................ 65
11 STORAGE, STABILITY AND DISPOSAL ...................................................................... 65
12 SPECIAL HANDLING INSTRUCTIONS........................................................................ 68
PART II: SCIENTIFIC INFORMATION ..................................................................................... 69
13 PHARMACEUTICAL INFORMATION ........................................................................ 69
14 CLINICAL TRIALS .................................................................................................... 70
14.1 Trial Design and Study Demographics ...................................................................... 70
14.1.1 COMIRNATY Original & Omicron BA.4/BA.5 (15/15mcg) .................................... 70
14.1.2 COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) ................................. 70
14.1.3 COMIRNATY (30 mcg) ........................................................................................... 70
14.1.4 COMIRNATY (10 mcg) ........................................................................................... 72
14.2 Study Results ............................................................................................................. 74
14.2.1 COMIRNATY Original & Omicron BA.4/BA.5 (15/15mcg) .................................... 74
14.2.1.1 Immunogenicity in Participants 12 Years of Age and Older – After Second Booster Dose (Fourth Dose) ................................................................................... 74
14.2.2 COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) ................................. 78
14.2.2.1 Immunogenicity in Participants 6 Months Through <5 Years of Age – After Booster (Fourth Dose) ............................................................................................ 78
14.2.3 COMIRNATY (30 mcg) ........................................................................................... 79
14.2.3.1 Efficacy and Immunogenicity in Participants 16 Years of Age and Older ....... 79
14.2.3.1.1 Efficacy in Participants 16 Years of Age and Older – After Two Doses .... 79
14.2.3.1.2 Efficacy and Immunogenicity in Participants 16 Years of Age and Older – After Booster Dose ............................................................................................... 81
14.2.3.2 Efficacy and Immunogenicity in Adolescents 12 to 15 Years of Age .............. 84
14.2.3.2.1 Efficacy and Immunogenicity in Adolescents 12 to 15 Years of Age – After Two Doses ................................................................................................... 84
14.2.4 COMIRNATY (10 mcg) ........................................................................................... 86
14.2.4.1 Efficacy and Immunogenicity in Children 5 Years Through <12 Years of Age . 86
14.2.4.1.1 Efficacy and Immunogenicity in Children 5 Years Through <12 Years of Age – After Two Doses ......................................................................................... 86
14.2.4.2 Immunogenicity in Children 5 Years Through <12 Years of Age – After Booster Dose ........................................................................................................................ 88
14.2.5 COMIRNATY (3 mcg) ............................................................................................. 89
14.2.5.1 Immunogenicity in Children 6 Months Through <5 Years of Age ................... 89
14.2.5.1.1 Immunogenicity in Children 6 Months Through <5 Years of Age – After a 3-Dose Primary Series .......................................................................................... 89
15 MICROBIOLOGY .................................................................................................... 93
16 NON-CLINICAL TOXICOLOGY .................................................................................. 93
PATIENT MEDICATION INFORMATION ................................................................................ 94
PART I: HEALTH PROFESSIONAL INFORMATION
1 INDICATIONS
COMIRNATY Omicron XBB.1.5 (COVID-19 mRNA Vaccine, Monovalent (Omicron XBB.1.5)) is indicated for active immunization against coronavirus disease 2019 (COVID-19) caused by severe acute respiratory syndrome coronavirus 2 (SARS- CoV-2) in individuals 6 months of age and older (see 4.2 Recommended Dose and Dosage Adjustment).
The safety and effectiveness of COMIRNATY Omicron XBB.1.5 for individuals 6 months of age and older are inferred from studies which evaluated the primary series and booster vaccination with COMIRNATY and supported by studies which evaluated a booster dose of COMIRNATY Original & Omicron BA.4/BA.5 in individuals 6 months of age and older.
1.1 Pediatrics
The safety and efficacy of COMIRNATY Omicron XBB.1.5 in children under 6 months of age have not yet been established.
1.2 Geriatrics
Clinical studies of COMIRNATY and COMIRNATY Original & Omicron BA.4/BA.5 included participants 65 years of age and older and their data contributes to the overall assessment of the safety and efficacy of COMIRNATY Omicron XBB.1.5 (see 8 ADVERSE REACTIONS and 14 CLINICAL TRIALS). The safety and efficacy of COMIRNATY Omicron XBB.1.5 in geriatric individuals have not yet been established in clinical trials.
2 CONTRAINDICATIONS
COMIRNATY Omicron XBB.1.5 is contraindicated in individuals who are hypersensitive to the active substance or to any ingredient in the formulation. For a complete listing see 6 DOSAGE FORMS, STRENGTHS, COMPOSITION AND PACKAGING.
3 SERIOUS WARNINGS AND PRECAUTIONS
At the time of authorization, there are no known serious warnings or precautions associated with this product.
4 DOSAGE AND ADMINISTRATION
4.1 Dosing Considerations
The storage, preparation and administration information differ depending on which presentation of the vaccine is considered. Careful attention should be paid to the vial cap and label border colour and information on the label, and the appropriate corresponding instructions must be followed under the subsections below.
For 12 Years of Age and Older
COMIRNATY Omicron XBB.1.5 is a suspension for intramuscular injection. DO NOT DILUTE (Single Dose or Multiple dose Vials with Gray Cap and Gray Label Border).
A single dose is 0.3 mL.
For Age 5 Years to <12 Years
There are two formulations of COMIRNATY Omicron XBB.1.5 for use in individuals 5 to <12 years of age, with different dosing.
DILUTE PRIOR TO USE: A suspension for intramuscular injection which must be diluted prior to administration (Multiple Dose Vials with Orange Cap and Orange Label Border).
- After preparation, a single dose is 0.2 mL.
DO NOT DILUTE: A ready to use suspension for intramuscular injection (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border).
- A single dose is 0.3 mL.
For Age 6 Months to <5 Years
COMIRNATY Omicron XBB.1.5 is a suspension for intramuscular injection which must be diluted prior to administration. DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border).
After preparation, a single dose is 0.2 mL.
| Dosage Form | Vial Cap and Label Border Colour | Age Range | Dilution Information | Dose(s) Per Vial | Dose Volume |
|---|---|---|---|---|---|
COMIRNATY Omicron XBB.1.5 Single Dose Vial (for 12 years of age and older: DO NOT DILUTE) | Gray | 12 years and older | DO NOT DILUTE prior to use | 1 | 0.3 mL |
COMIRNATY Omicron XBB.1.5 Multiple Dose Vial (for 12 years of age and older: DO NOT DILUTE) | Gray | 12 years and older | DO NOT DILUTE prior to use | 6 | 0.3 mL |
| COMIRNATY Omicron XBB.1.5 Multiple Dose Vial (for age 5 years to <12 years: DILUTE PRIOR TO USE) | Orange | 5 to <12 years | Dilute with 1.3 mL sterile 0.9% Sodium Chloride Injection, USP prior to use | 10 | 0.2 mL |
| COMIRNATY Omicron XBB.1.5 Single Dose Vial (for age 5 years to <12 years: DO NOT DILUTE) | Blue | 5 to <12 years | DO NOT DILUTE prior to use | 1 | 0.3 mL |
| COMIRNATY Omicron XBB.1.5 Multiple Dose Vial (for age 5 years to <12 years: DO NOT DILUTE) | Blue | 5 to <12 years | DO NOT DILUTE prior to use | 6 | 0.3 mL |
| COMIRNATY Omicron XBB.1.5 Multiple Dose Vial (for age 6 | Maroon | 6 months to <5 years | Dilute with 2.2 mL sterile 0.9% Sodium Chloride | 10 | 0.2 mL |
| Dosage Form | Vial Cap and Label Border Colour | Age Range | Dilution Information | Dose(s) Per Vial | Dose Volume |
|---|---|---|---|---|---|
| months to <5 years: DILUTE PRIOR TO USE) | Injection, USP prior to use | ||||
4.2 Recommended Dose and Dosage Adjustment
4.2.1 Vaccination Schedule for Individuals 12 Years of Age and Older
COMIRNATY Omicron XBB.1.5 is administered intramuscularly as a single dose of 0.3 mL, regardless of prior COVID-19 vaccination status.
For individuals who have previously been vaccinated with a COVID-19 vaccine, COMIRNATY Omicron XBB.1.5 should be administered at least 3 to 6 months after the most recent dose of a COVID-19 vaccine.
Vials of COMIRNATY Omicron XBB.1.5 intended for individuals 12 years of age and older (gray cap/gray label border) cannot be used to prepare doses for individuals aged 6 months to <12 years of age.
4.2.2 Vaccination Schedule for Individuals Aged 5 Years to <12 Years
COMIRNATY Omicron XBB.1.5 is administered intramuscularly as a single dose, regardless of prior COVID-19 vaccination status.
- DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border): After dilution, a single dose is 0.2 mL.
- DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border): A single dose is 0.3 mL.
For individuals who have previously been vaccinated with a COVID-19 vaccine, COMIRNATY Omicron XBB.1.5 should be administered at least 6 months after the most recent dose of a COVID-19 vaccine.
Vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 5 years to <12 years (orange cap/orange label border or blue cap/blue label border) cannot be used to prepare doses for individuals 6 months to < 5 years or 12 years of age and older.
4.2.3 Vaccination Schedule for Individuals Aged 6 Months to <5 Years
Infants and children 6 months to <5 years of age without history of completion of a COVID-19 primary course
COMIRNATY Omicron XBB.1.5 is administered intramuscularly as a three-dose course (0.2 mL each). It is recommended to administer the second dose 3 weeks after the first dose, followed by a third dose administered at least 8 weeks after the second dose.
If an infant or child starts a primary vaccination course with COMIRNATY or COMIRNATY Original & Omicron BA.4/BA.5, they may complete the three-dose course with COMIRNATY Omicron XBB.1.5.
The interchangeability of COMIRNATY Omicron XBB.1.5 with COVID-19 vaccines from other manufacturers to complete the three-dose course has not been established. Individuals who have received a dose of COMIRNATY Omicron XBB.1.5 should receive COMIRNATY Omicron XBB.1.5 to complete the three-dose course.
Infants and children 6 months to <5 years of age with history of completion of a COVID-19 primary course
COMIRNATY Omicron XBB.1.5 is administered intramuscularly as a single dose of 0.2 mL for infants and children 6 months to <5 years of age.
For individuals who have previously been vaccinated with a COVID-19 vaccine, COMIRNATY Omicron XBB.1.5 should be administered at least 6 months after the most recent dose of a COVID-19 vaccine.
Vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 6 months to <5 years (maroon cap/maroon label border) cannot be used to prepare doses for individuals 5 years of age and older.
4.3 Reconstitution
4.3.1 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Gray Cap and Gray Label Border)
The COMIRNATY Omicron XBB.1.5 single dose or multiple dose vial with a gray cap and gray label border MUST NOT BE DILUTED prior to administration. Instructions on the handling and dose preparation of the vaccine prior to administration are provided below.
Preparation for Administration
Single Dose Vials
DO NOT DILUTE
- The COMIRNATY Omicron XBB.1.5 single dose vial with a gray cap and a gray label border is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed prior to administration. DO NOT DILUTE prior to use.
- Single dose vials of COMIRNATY Omicron XBB.1.5 intended for 12 years of age or older with a gray cap/gray label border cannot be used to prepare doses for individuals aged 6 months to <12 years.
- Single dose vials may be thawed in the refrigerator (2°C to 8°C [35°F to 46°F]) or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- The thawed suspension may contain white to off-white opaque amorphous particles.
- One single dose vial contains 1 dose of 0.3 mL.
- Refer to thawing and dose preparation instructions in the panels below.
Multiple Dose Vials
DO NOT DILUTE
- The COMIRNATY Omicron XBB.1.5 multiple dose vial with a gray cap and a gray label border contains a volume of 2.25 mL, and is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed prior to administration. DO NOT DILUTE prior to use.
- Multiple dose vials of COMIRNATY Omicron XBB.1.5 intended for 12 years of age or older with a gray cap/gray label border cannot be used to prepare doses for individuals aged 6 months to <12 years.
- Multiple dose vials may be thawed in the refrigerator (2°C to 8°C [35°F to 46°F]) or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- The thawed suspension may contain white to off-white opaque amorphous particles.
- One multiple dose vial contains 6† doses of 0.3 mL.
- Refer to thawing and dose preparation instructions in the panels below.
COMIRNATY Omicron XBB.1.5 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Gray Cap and Gray Label Border) | ||
|---|---|---|
| VIAL AND DOSE VERIFICATION | ||
| Gray plastic cap and label with gray border |
| |
† Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
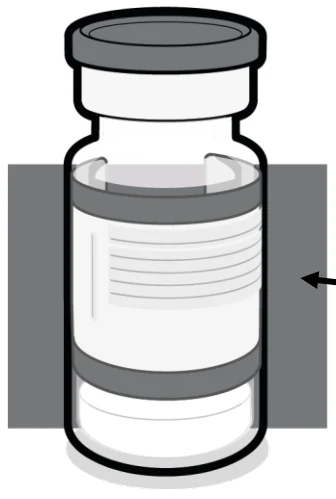 COMIRNATY Omicron XBB.1.5 Do not dilute |
|---|
COMIRNATY Omicron XBB.1.5 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Gray Cap and Gray Label Border) | |
|---|---|
| THAWING PRIOR TO USE | |
| |
| |
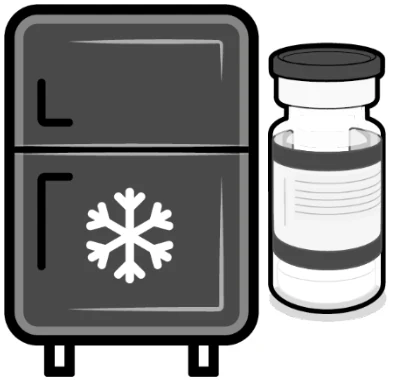 Can be stored in the refrigerator (2°C to 8°C) for up to 10 weeks prior to use within the expiry date 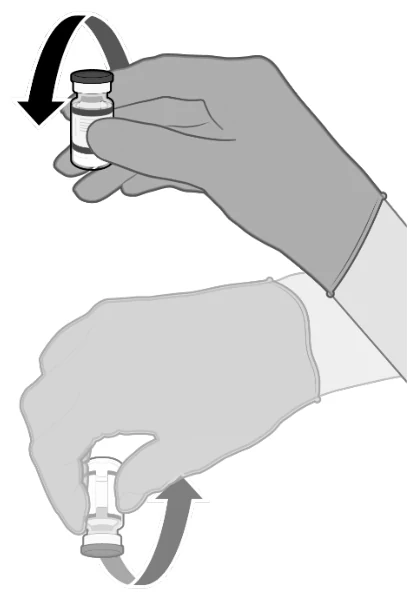 Gently × 10 |
|---|
COMIRNATY Omicron XBB.1.5 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Gray Cap and Gray Label Border) | |
|---|---|
| PREPARATION OF INDIVIDUAL 0.3 mL DOSES | |
Single dose vials
Multiple dose vials
| |
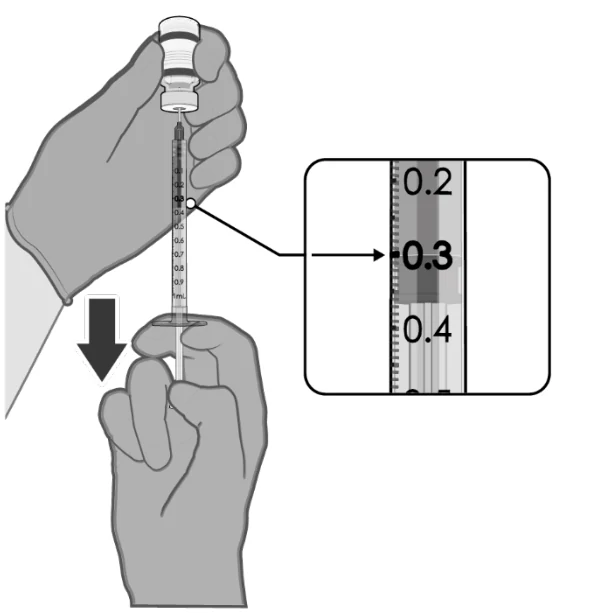 Withdraw 0.3 mL dose of vaccine COMIRNATY Omicron XBB.1.5 For 12 Years of Age and Older: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Gray Cap and Gray Label Border) | |
|---|---|
Multiple Dose Vial Only:
| |
4.3.2 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border)
Preparation for Administration
Prior to Dilution:
- The COMIRNATY Omicron XBB.1.5 multiple dose vial (for ages 5 years to <12 years) has an orange cap and an orange label border and contains a volume of 1.3 mL. It is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed and diluted prior to administration.
- Multiple dose vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 5 years to <12 years cannot be used to prepare doses for individuals aged 6 months to <5 years or 12 years and older.
- Multiple dose vials may be thawed in the refrigerator at 2°C to 8°C [35°F to 46°F] or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- Prior to dilution, the thawed suspension may contain opaque amorphous particles.
- Refer to thawing instructions in the panels below.
Dilution:
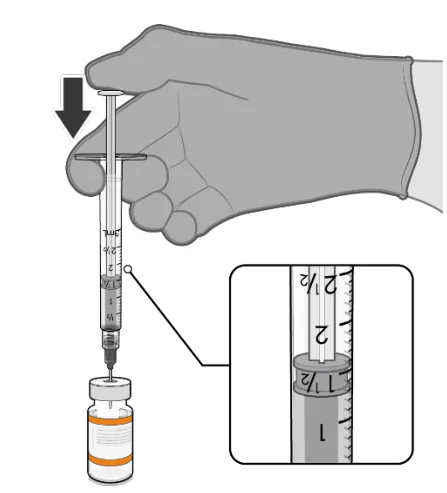
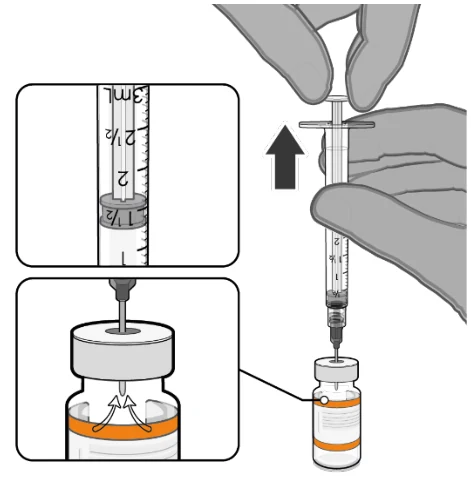
- Dilute the multiple dose vial contents using 1.3 mL of sterile 0.9% Sodium Chloride Injection, USP to form COMIRNATY Omicron XBB.1.5. Do not add more than 1.3 mL of diluent.
- ONLY use 0.9% Sodium Chloride Injection, USP as the diluent. This diluent is not packaged with the vaccine and must be sourced separately. Do not use bacteriostatic 0.9% Sodium Chloride Injection or any other diluent.
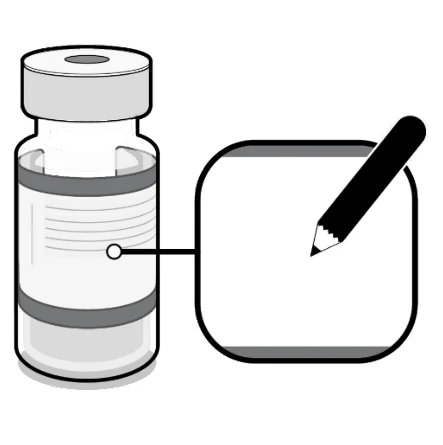 Record the discard time on the vial label. Use within 12 hours after first puncture. |
|---|
- After dilution, one multiple dose vial contains 10* doses of 0.2 mL.
- After dilution, the vaccine will be a white to off-white suspension. Inspect vials to confirm there are no particulates and no discolouration is observed.
- Strict adherence to aseptic techniques must be followed.
- Refer to dilution and dose preparation instructions in the panels below.
| COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border) | ||
|---|---|---|
| VIAL AND DOSE VERIFICATION | ||
| Orange plastic cap and label with orange border |
| |
| THAWING PRIOR TO DILUTION | ||
| ||
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
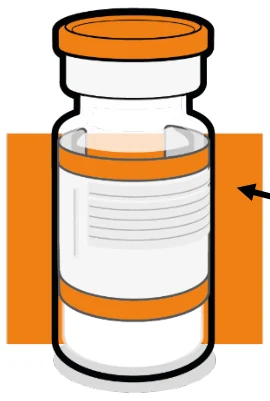 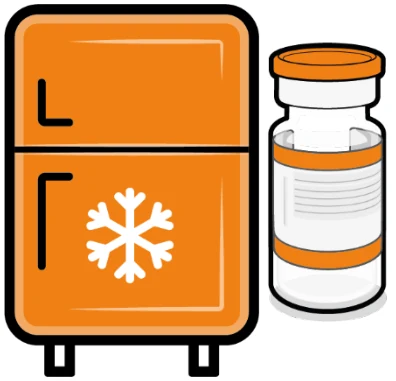 Can be stored in the refrigerator (2°C to 8°C) for up to 10 weeks prior to use within the expiry date. COMIRNATY Omicron XBB.1.5 10 doses after dilution |
|---|
| COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border) | |||
|---|---|---|---|
| Gently x 10 |
| ||
| DILUTION | |||
| Add 1.3 mL of 0.9% Sodium Chloride Injection, USP. |
| ||
| COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border) | ||
|---|---|---|
| Pull back plunger to 1.3 mL to remove air from the vial. |
| |
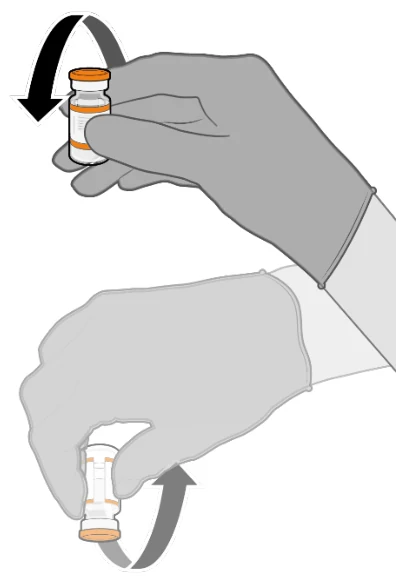
| Gently x 10 |
| |
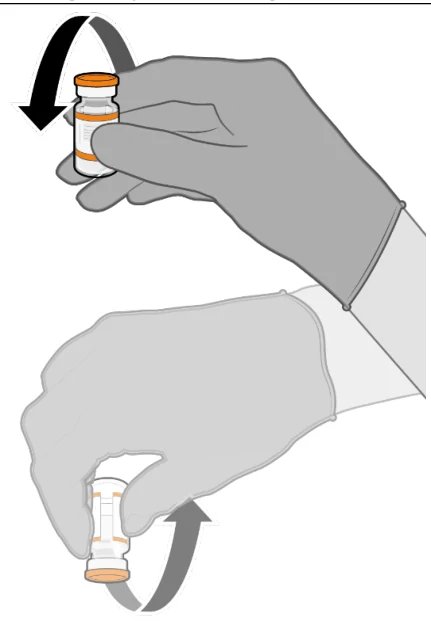    COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border) | ||
|---|---|---|
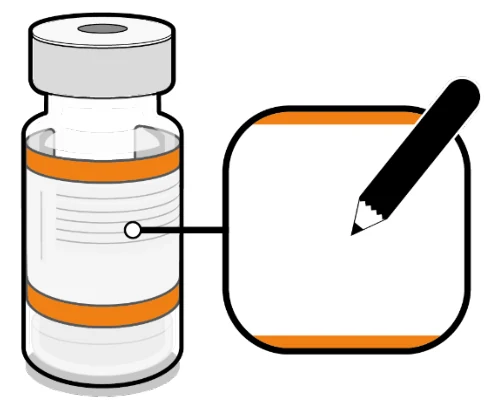 Record the discard time on the vial label. Use within 12 hours after dilution. |
| |
| PREPARATION OF INDIVIDUAL 0.2 mL DOSES | ||
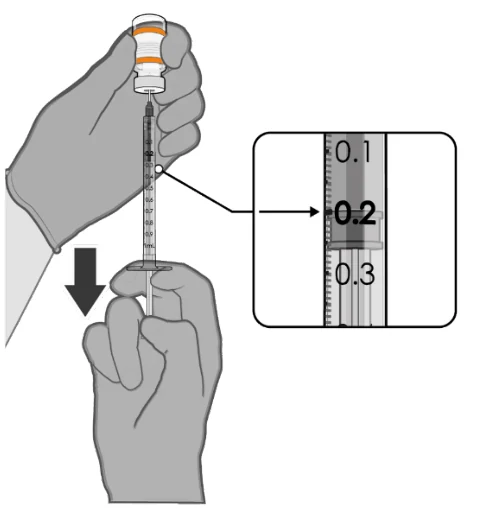 Withdraw 0.2 mL dose of vaccine |
| |
4.3.3 For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border)
The COMIRNATY Omicron XBB.1.5 single dose or multiple dose vial with a blue cap and blue label border MUST NOT BE DILUTED prior to administration. Instructions on the handling and dose preparation of the vaccine prior to administration are provided below.
Preparation for Administration
Single Dose Vials
DO NOT DILUTE
- The COMIRNATY Omicron XBB.1.5 single dose vial with a blue cap and a blue label border is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed prior to administration. DO NOT DILUTE prior to use.
- Single dose vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 5 years to <12 years of age cannot be used to prepare doses for individuals aged 6 months to <5 years or 12 years and older.
- Single dose vials may be thawed in the refrigerator (2°C to 8°C [35°F to 46°F]) or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- The thawed suspension may contain white to off-white opaque amorphous particles.
- One single dose vial contains 1 dose of 0.3 mL.
- Refer to thawing and dose preparation instructions in the panels below.
Multiple Dose Vials
DO NOT DILUTE
- The COMIRNATY Omicron XBB.1.5 multiple dose vial with a blue cap and a blue label border contains a volume of 2.25 mL, and is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed prior to administration. DO NOT DILUTE prior to use.
- Multiple dose vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 5 years to <12 years of age cannot be used to prepare doses for individuals aged 6 months to <5 years or 12 years and older.
- Multiple dose vials may be thawed in the refrigerator (2°C to 8°C [35°F to 46°F]) or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- The thawed suspension may contain white to off-white opaque amorphous particles.
- One multiple dose vial contains 6† doses of 0.3 mL.
- Refer to thawing and dose preparation instructions in the panels below.
† Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Blue Cap and Blue Label Border) | ||
|---|---|---|
| VIAL AND DOSE VERIFICATION | ||
| Blue plastic cap and label with blue border |
| |
| THAWING PRIOR TO USE | ||
| ||
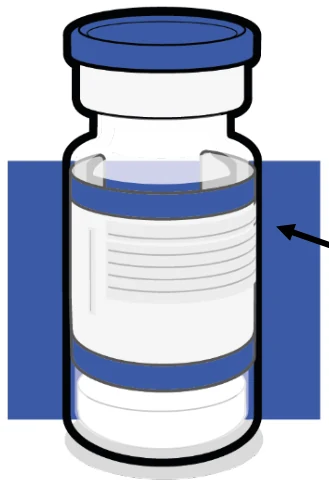 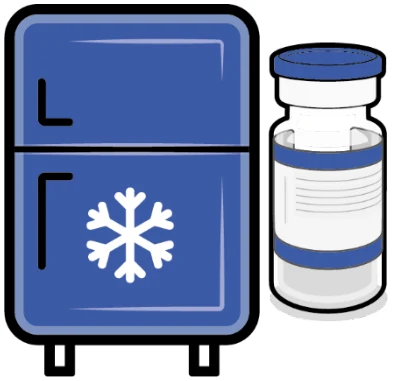 Can be stored in the refrigerator (2°C to 8°C) for up to 10 weeks prior to use within the expiry date. COMIRNATY Omicron XBB.1.5 Do not dilute |
|---|
COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Blue Cap and Blue Label Border) | |
|---|---|
| |
| PREPARATION OF INDIVIDUAL 0.3 mL DOSES | |
Single dose vials
Multiple dose vials
| |
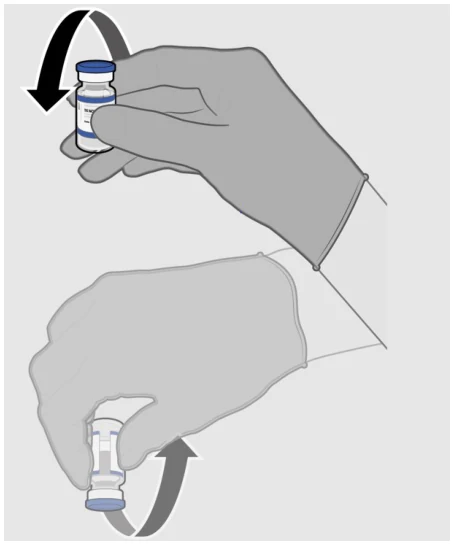 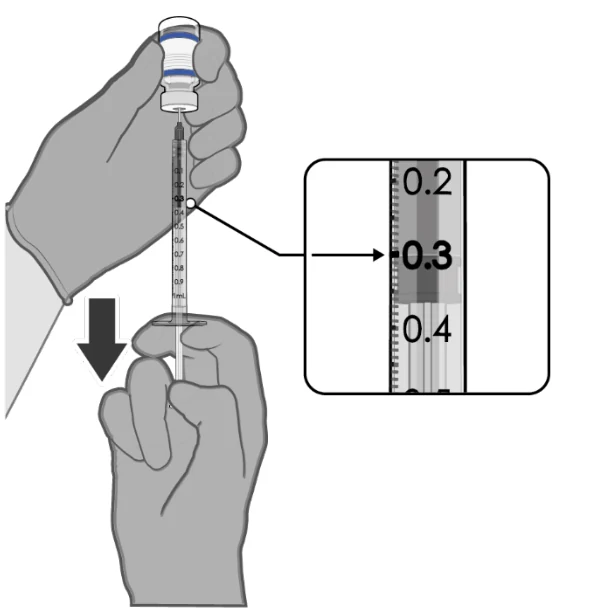 Withdraw 0.3 mL dose of vaccine Gently × 10 COMIRNATY Omicron XBB.1.5 For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose & Multiple Dose Vials with Blue Cap and Blue Label Border) | ||
|---|---|---|
| ||
Multiple Dose Vial Only:
| ||
4.3.4 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border)
Preparation for Administration
Prior to Dilution:
- The COMIRNATY Omicron XBB.1.5 multiple dose vial (for age 6 months to <5 years) has a maroon cap and a maroon label border and contains a volume of 0.4 mL. It is supplied as a frozen suspension that does not contain preservative. Each vial must be thawed and diluted prior to administration.
- Multiple dose vials of COMIRNATY Omicron XBB.1.5 intended for individuals aged 6 months to <5 years cannot be used to prepare doses for individuals aged 5 years and older.
- Multiple dose vials may be thawed in the refrigerator at 2°C to 8°C [35°F to 46°F] or at room temperature (up to 25°C [77°F]) (see 11 STORAGE, STABILITY AND DISPOSAL).
- Prior to dilution, the thawed suspension may contain opaque amorphous particles.
- Refer to thawing instructions in the panels below.
Dilution:
- Dilute the multiple dose vial contents using 2.2 mL of sterile 0.9% Sodium Chloride Injection, USP to form COMIRNATY Omicron XBB.1.5. Do not add more than 2.2 mL of diluent.
- ONLY use 0.9% Sodium Chloride Injection, USP as the diluent. This diluent is not packaged with the vaccine and must be sourced separately. Do not use bacteriostatic 0.9% Sodium Chloride Injection or any other diluent.
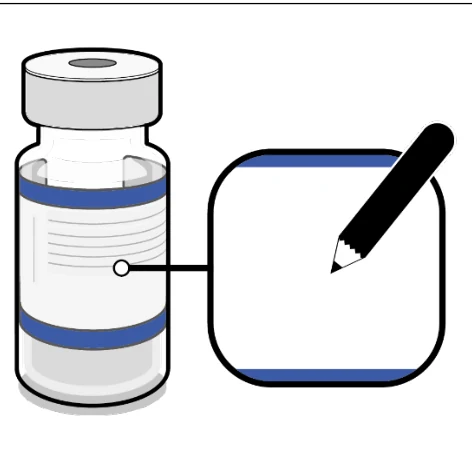 Record the discard time on the vial label. Use within 12 hours after first puncture. |
|---|
- After dilution, one multiple dose vial contains 10* doses of 0.2 mL.
- After dilution, the vaccine will be a white to off-white suspension. Inspect vials to confirm there are no particulates and no discolouration is observed.
- Strict adherence to aseptic techniques must be followed.
- Refer to dilution and dose preparation instructions in the panels below.
| COMIRNATY Omicron XBB.1.5 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) | ||
|---|---|---|
| VIAL AND DOSE VERIFICATION | ||
| Maroon plastic cap and maroon label border |
| |
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
 COMIRNATY Omicron XBB.1.5 10 doses after dilution |
|---|
| COMIRNATY Omicron XBB.1.5 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) | ||
|---|---|---|
| THAWING PRIOR TO DILUTION | ||
| ||
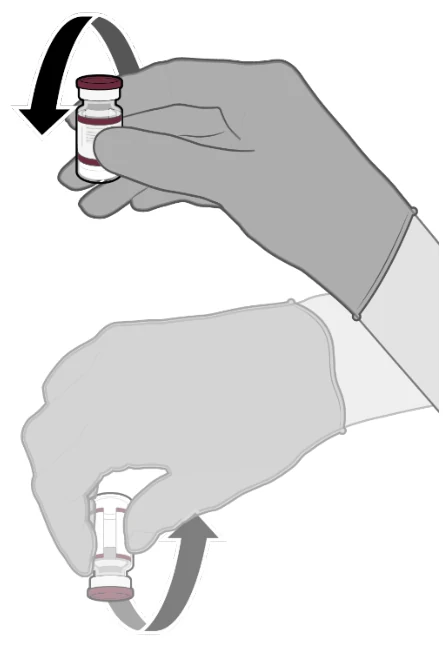
| Gently × 10 |
| |
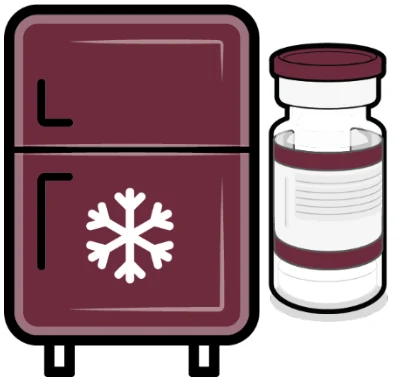 Can be stored in the refrigerator (2°C to 8°C) for up to 10 weeks prior to use |
|---|
| COMIRNATY Omicron XBB.1.5 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) | ||
|---|---|---|
| DILUTION | ||
| ||
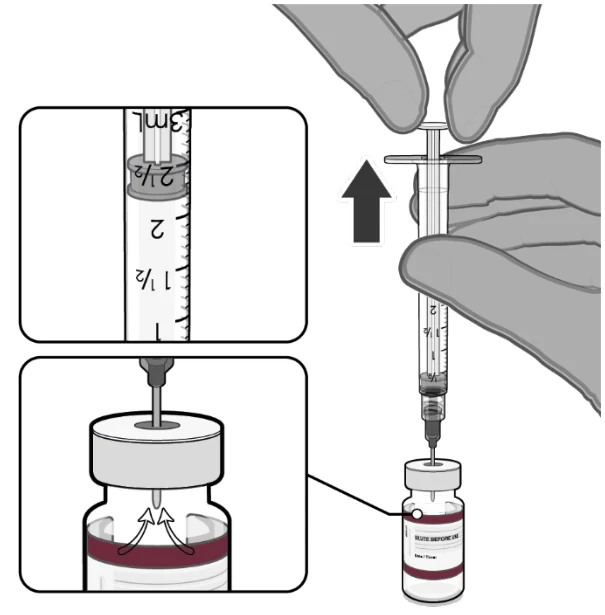
| Pull back plunger to 2.2 mL to remove air from vial |
| |
 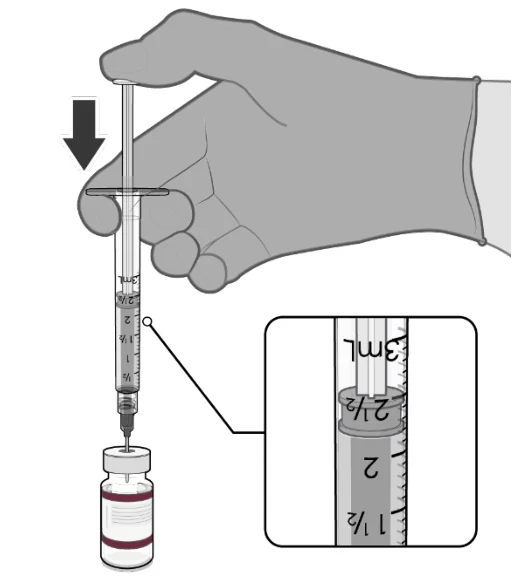 Add 2.2 mL of sterile 0.9% Sodium Chloride Injection, USP |
|---|
| COMIRNATY Omicron XBB.1.5 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) | ||
|---|---|---|
| ||
Record the discard time on the vial label. Use within 12 hours after dilution. |
| |
 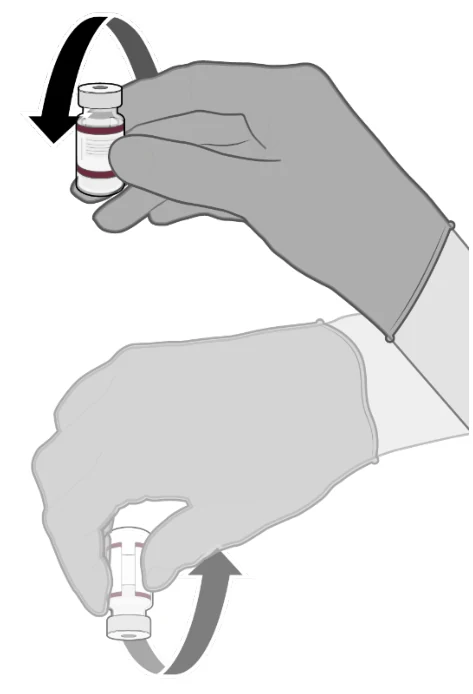 Gently × 10 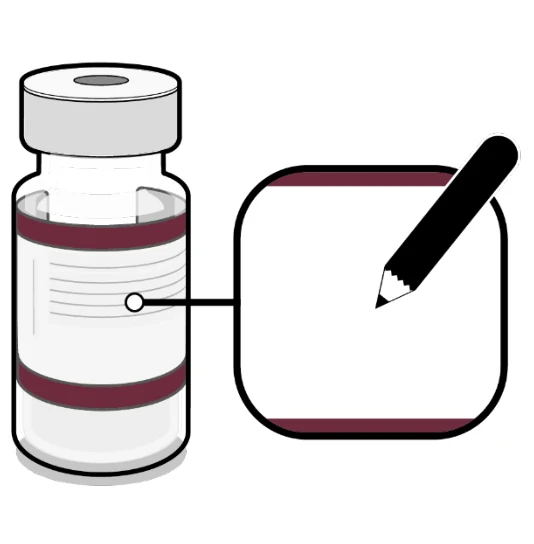 COMIRNATY Omicron XBB.1.5 For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border) | ||
|---|---|---|
| PREPARATION OF INDIVIDUAL 0.2 mL DOSES | ||
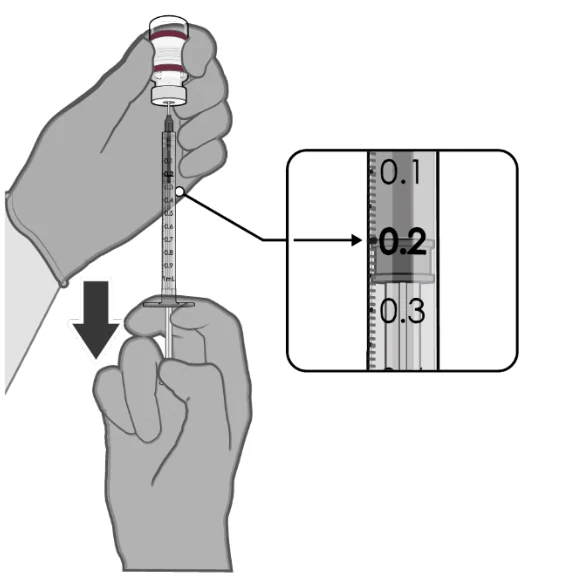 Withdraw 0.2 mL dose of vaccine |
| |
4.4 Administration
4.4.1 For 12 Years of Age and Older
DO NOT DILUTE (Single Dose or Multiple Dose Vials with Gray Cap and Gray Label Border)
Administer a single 0.3 mL dose of COMIRNATY Omicron XBB.1.5 intramuscularly, preferably in the deltoid muscle.
Do not inject the vaccine intravascularly, subcutaneously or intradermally.
Visually inspect each dose in the dosing syringe prior to administration. The vaccine will be an off-white suspension. During the visual inspection:
- Verify the final dosing volume of 0.3 mL.
- Confirm there are no particulates and that no discolouration is observed.
- Do not administer if vaccine is discoloured or contains particulate matter.
Single Dose Vials
Single dose vials of COMIRNATY Omicron XBB.1.5 with gray caps and gray label borders contain 1 dose of 0.3 mL of vaccine.
- Withdraw a single 0.3 mL dose of vaccine.
- Discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
Multiple Dose Vials
Multiple dose vials of COMIRNATY Omicron XBB.1.5 with gray caps and gray label borders contain 6 doses of 0.3 mL of vaccine.
Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. In order to ensure consistent withdrawal of 6 doses of 0.3 mL, it is important to adhere to minimizing volume loss during dose extraction. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial. Irrespective of the type of syringe and needle:
- Each dose must contain 0.3 mL of vaccine.
- If the amount of vaccine remaining in the vial cannot provide a full dose of 0.3 mL, discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
4.4.2 For Age 5 Years to <12 Years
DILUTE PRIOR TO USE (Multiple Dose Vial with Orange Cap and Orange Label Border)
Administer a single 0.2 mL dose of COMIRNATY Omicron XBB.1.5 intramuscularly, preferably in the deltoid muscle.
Do not inject the vaccine intravascularly, subcutaneously or intradermally.
DO NOT administer COMIRNATY Omicron XBB.1.5 with gray cap and gray label border or maroon cap and maroon label border to children 5 years to <12 years.
Visually inspect each dose in the dosing syringe prior to administration. The diluted vaccine will be a white to off-white suspension. During the visual inspection:
- Verify the final dosing volume of 0.2 mL.
- Confirm there are no particulates and that no discolouration is observed.
- Do not administer if vaccine is discoloured or contains particulate matter.
After dilution, multiple dose vials of COMIRNATY Omicron XBB.1.5 (for age 5 years to <12 years) contain 10 doses of 0.2 mL of vaccine. Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. In order to ensure consistent withdrawal of 10 doses of 0.2 mL, it is important to adhere to minimizing volume loss during dose extraction. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial. Irrespective of the type of syringe and needle:
- Each dose must contain 0.2 mL of vaccine.
- If the amount of vaccine remaining in the vial cannot provide a full dose of 0.2 mL, discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border)
Administer a single 0.3 mL dose of COMIRNATY Omicron XBB.1.5 intramuscularly, preferably in the deltoid muscle.
Do not inject the vaccine intravascularly, subcutaneously or intradermally.
Visually inspect each dose in the dosing syringe prior to administration. The vaccine will be an off-white suspension. During the visual inspection:
- Verify the final dosing volume of 0.3 mL.
- Confirm there are no particulates and that no discolouration is observed.
- Do not administer if vaccine is discoloured or contains particulate matter.
Single Dose Vials
Single dose vials of COMIRNATY Omicron XBB.1.5 with blue caps and blue label borders contain 1 dose of 0.3 mL of vaccine.
- Withdraw a single 0.3 mL dose of vaccine.
- Discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
Multiple Dose Vials
Multiple dose vials of COMIRNATY Omicron XBB.1.5 with blue caps and blue label borders contain 6 doses of 0.3 mL of vaccine.
Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. In order to ensure consistent withdrawal of 6 doses of 0.3 mL, it is important to adhere to minimizing volume loss during dose extraction. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial. Irrespective of the type of syringe and needle:
- Each dose must contain 0.3 mL of vaccine.
- If the amount of vaccine remaining in the vial cannot provide a full dose of 0.3 mL, discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
4.4.3 For Age 6 Months to <5 Years
Administer a single 0.2 mL dose of COMIRNATY Omicron XBB.1.5 intramuscularly. In individuals from 6 to less than 12 months of age, the recommended injection site is the anterolateral aspect of the thigh. In individuals 1 year of age and older, the recommended injection site is the anterolateral aspect of the thigh or the deltoid muscle.
Do not inject the vaccine intravascularly, subcutaneously or intradermally.
DO NOT administer COMIRNATY Omicron XBB.1.5 with gray cap and gray label border, blue cap and blue label or orange cap and orange label border to infants and children 6 months to <5 years.
DILUTE PRIOR TO USE (Multiple Dose Vial with Maroon Cap and Maroon Label Border).
Visually inspect each dose in the dosing syringe prior to administration. The diluted vaccine will be a white to off-white suspension. During the visual inspection:
- Verify the final dosing volume of 0.2 mL.
- Confirm there are no particulates and that no discolouration is observed.
- Do not administer if vaccine is discoloured or contains particulate matter.
After dilution, multiple dose vials of COMIRNATY Omicron XBB.1.5 (for age 6 months to <5 years) contain 10 doses of 0.2 mL of vaccine. Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. In order to ensure consistent withdrawal of 10 doses of 0.2 mL, it is important to adhere to minimizing volume loss during dose extraction. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial. Irrespective of the type of syringe and needle:
- Each dose must contain 0.2 mL of vaccine.
- If the amount of vaccine remaining in the vial cannot provide a full dose of 0.2 mL, discard the vial and any excess volume.
- Do not pool excess vaccine from multiple vials.
5 OVERDOSAGE
In the event of suspected overdose, monitoring of vital functions and symptomatic treatment is recommended. Contact your regional poison control centre.
6 DOSAGE FORMS, STRENGTHS, COMPOSITION AND PACKAGING
To help ensure the traceability of vaccines for patient immunization record-keeping as well as safety monitoring, health professionals should record the time and date of administration, quantity of administered dose (if applicable), anatomical site and route of administration, brand name and generic name of the vaccine, the product lot number and expiry date.
For 12 Years of Age and Older: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Gray Cap and Gray Label Border)
COMIRNATY Omicron XBB.1.5 is supplied as a frozen suspension in single dose or multiple dose vials with gray caps and labels with gray borders. Do not dilute. Each single dose vial contains 1 dose of 0.3 mL and each multiple dose vial contains 6† doses of 0.3 mL. Each 0.3 mL dose of COMIRNATY Omicron XBB.1.5 contains 30 mcg of a nucleoside modified messenger RNA (modRNA) encoding the viral spike (S) protein of SARS-CoV-2 Omicron variant lineage XBB.1.5 (Omicron XBB.1.5). Each dose also includes the non-medicinal ingredients listed in Table 1.
COMIRNATY Omicron XBB.1.5 does not contain preservative. The vial stoppers are not made with natural rubber latex.
COMIRNATY Omicron XBB.1.5 single dose or multiple dose vials (with gray cap and gray label border) are supplied in a carton containing 10 single dose or multiple dose vials.
† Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
| Table 1: Dosage Forms, Strengths, Composition and Packaging (DO NOT DILUTE: For Age 5 Years to <12 Years and For 12 Years of Age and Older) | |||
| Route of Administration | Dosage Form / Strength/Composition | Non-medicinal Ingredients | |
| Intramuscular injection | DO NOT DILUTE (Vials with Gray Cap and Gray Label Border) Suspension (do not dilute) Raxtozinameran (mRNA) encodes for the viral spike (S) protein of SARS-CoV-2 Omicron XBB.1.5 strain Single dose vial: (each vial contains 1 dose of 0.3 mL) Multiple dose vial: (each vial contains 6† doses of 0.3 mL) |
| |
| Intramuscular injection | DO NOT DILUTE (Vials with Blue Cap and Blue Label Border) Suspension (do not dilute) Raxtozinameran (mRNA) encodes for the viral spike (S) protein of SARS-CoV-2 Omicron XBB.1.5 strain Single dose vial: (each vial contains 1 dose of 0.3 mL) Multiple dose vial: (each vial 6† doses of 0.3 mL) | ||
For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Vials with Orange Cap and Orange Label Border)
COMIRNATY Omicron XBB.1.5 is supplied as a frozen suspension in multiple dose vials with an orange cap and an orange label border. Each multiple dose vial must be diluted with 1.3 mL of sterile 0.9% Sodium Chloride Injection, USP prior to use to form the vaccine, and contains 10* doses of 0.2 mL after dilution. Each 0.2 mL dose of COMIRNATY Omicron XBB.1.5 contains 10 mcg of a nucleoside modified messenger RNA (modRNA) encoding the viral spike (S) protein of SARS-CoV-2 Omicron variant lineage XBB.1.5 (Omicron XBB.1.5) and the non-medicinal ingredients listed in Table 2. COMIRNATY Omicron XBB.1.5 does not contain preservative. The vial stoppers are not made with natural rubber latex.
COMIRNATY Omicron XBB.1.5 multiple dose vials (with orange cap and orange label border) are
† Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
supplied in a carton containing 10 multiple dose vials.
For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border)
COMIRNATY Omicron XBB.1.5 is supplied as a frozen suspension in single dose or multiple dose vials with blue caps and labels with blue borders. Do not dilute. Each single dose vial contains 1 dose of 0.3 mL and each multiple dose vial contains 6† doses of 0.3 mL. Each 0.3 mL dose of COMIRNATY Omicron XBB.1.5 contains 10 mcg of a nucleoside modified messenger RNA (modRNA) encoding the viral spike (S) protein of SARS-CoV-2 Omicron variant lineage XBB.1.5 (Omicron XBB.1.5). Each dose also includes the non-medicinal ingredients listed in Table 1.
COMIRNATY Omicron XBB.1.5 does not contain preservative. The vial stoppers are not made with natural rubber latex.
COMIRNATY Omicron XBB.1.5 single dose or multiple dose vials (with blue cap and blue label border) are supplied in a carton containing 10 single dose or multiple dose vials.
Table 2: Dosage Forms, Strengths, Composition and Packaging (DILUTE PRIOR TO USE: For Age 5 Years to <12 Years and For Age 6 Months to <5 Years)
† Low dead-volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
| Route of Administration | Dosage Form / Strength/Composition | Non-medicinal Ingredients |
| Intramuscular injection | DILUTE PRIOR TO USE (Vials with Orange Cap and Orange Label Border) Suspension (to be diluted before use) Raxtozinameran (mRNA) encodes for the viral spike (S) protein of SARS-CoV-2 Omicron XBB.1.5 strain Multiple dose vial (after dilution, each vial contains10* doses of 0.2 mL) |
|
| Intramuscular injection | DILUTE PRIOR TO USE (Vials with Maroon Cap and Maroon Label Border) Suspension (to be diluted before use) Raxtozinameran (mRNA) encodes for the viral spike (S) protein of SARS-CoV-2 | |
For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Vials with Maroon Cap and Maroon Label Border)
COMIRNATY Omicron XBB.1.5 is supplied as a frozen suspension in multiple dose vials with a maroon cap and a maroon label border. Each multiple dose vial must be diluted with 2.2 mL of sterile 0.9% Sodium Chloride Injection, USP prior to use to form the vaccine, and contains 10* doses of 0.2 mL after dilution. Each 0.2 mL dose of COMIRNATY Omicron XBB.1.5 contains 3 mcg of a nucleoside modified messenger RNA (modRNA) encoding the viral spike (S) protein of SARS-CoV-2 Omicron variant lineage XBB.1.5 (Omicron XBB.1.5) and the non-medicinal ingredients listed in Table 2. COMIRNATY Omicron XBB.1.5 does not contain preservative. The vial stoppers are not made with natural rubber latex.
COMIRNATY Omicron XBB.1.5 multiple dose vials (with maroon cap and maroon label border) are supplied in a carton containing 10 multiple dose vials.
7 WARNINGS AND PRECAUTIONS
General
The administration of COMIRNATY Omicron XBB.1.5 should be postponed in individuals suffering from acute severe febrile illness.
Fainting may occur in association with administration of injectable vaccines. Individuals should be advised to bring symptoms (e.g., dizziness, increases in heart rate, feeling short of breath, tingling sensations or sweating) to the attention of the vaccination provider for evaluation. Procedures should be in place to avoid injury from fainting.
As with any vaccine, vaccination with COMIRNATY Omicron XXB.1.5 may not protect all recipients.
Acute Allergic Reactions
Anaphylaxis has been reported. As with all vaccines, training for immunizers, appropriate medical treatment and supervision after immunization should always be readily available in case of a rare anaphylactic event following the administration of this vaccine.
Vaccine recipients should be kept under observation for at least 15 minutes after immunization; 30 minutes is a preferred interval when there is a specific concern about a possible vaccine reaction.
COMIRNATY Omicron XBB.1.5 should not be given to those who have experienced anaphylaxis after a prior dose of COMIRNATY, COMIRNATY Original/Omicron BA.1, or COMIRNATY Original & Omicron BA.4/BA.5.
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
| Route of Administration | Dosage Form / Strength/Composition | Non-medicinal Ingredients | |
Omicron XBB.1.5 strain Multiple dose vial (after dilution, each vial contains 10* doses of 0.2 mL) | |||
Cardiovascular
Myocarditis and Pericarditis
Very rare cases of myocarditis and/or pericarditis following vaccination with COMIRNATY have been reported during post-authorization use. These cases occurred more commonly after the second dose and in adolescents and young adults. Typically, the onset of symptoms has been within a few days following receipt of COMIRNATY. Based on accumulating data, the reporting rates of myocarditis and pericarditis after COMIRNATY primary series in children ages 5 through <12 years are lower than in ages 12 through 17 years. Available short-term follow-up data suggest that the symptoms resolve in most individuals, but information on long-term sequelae is lacking. The decision to administer COMIRNATY Omicron XBB.1.5 to an individual with a history of myocarditis or pericarditis should take into account the individual’s clinical circumstances.
Healthcare professionals are advised to consider the possibility of myocarditis and/or pericarditis in their differential diagnosis if individuals present with chest pain, shortness of breath, palpitations or other signs and symptoms of myocarditis and/or pericarditis following immunization with a COVID-19 vaccine. This could allow for early diagnosis and treatment. Cardiology consultation for management and follow up should be considered.
Driving and Operating Machinery
COMIRNATY Omicron XBB.1.5 has no or negligible influence on the ability to drive and use machines. However, some of the effects mentioned under 8 ADVERSE REACTIONS may temporarily affect the ability to drive or use machines.
Fertility
It is unknown whether this vaccine has an impact on fertility. Animal studies do not indicate direct or indirect harmful effects with respect to female fertility or reproductive toxicity (see 16 NON-CLINICAL TOXICOLOGY).
Hematologic
Individuals receiving anticoagulant therapy or those with a bleeding disorder that would contraindicate intramuscular injection should not be given the vaccine unless the potential benefit clearly outweighs the risk of administration.
Immune
The efficacy and safety of the vaccine has not been assessed in immunocompromised individuals, including those receiving immunosuppressant therapy. Immunocompromised persons, including individuals receiving immunosuppressant therapy, may have a diminished immune response to the vaccine.
7.1 Special Populations
7.1.1 Pregnant Women
No data are available yet regarding the use of COMIRNATY Omicron XBB.1.5 during pregnancy.
Animal studies do not indicate direct or indirect harmful effects with respect to pregnancy, embryo/ fetal development, parturition, or post-natal development (see 16 NON-CLINICAL TOXICOLOGY).
7.1.2 Breast-feeding
No data are available yet regarding the use of COMIRNATY Omicron XBB.1.5 during breast- feeding.
It is unknown whether COMIRNATY Omicron XBB.1.5 is excreted in human milk. A risk to the newborns/infants cannot be excluded.
The developmental and health benefits of breastfeeding should be considered along with the mother’s clinical need for immunization against COVID-19.
7.1.3 Pediatrics
The safety and efficacy of COMIRNATY Omicron XBB.1.5 in children under 6 months of age have not yet been established.
7.1.4 Geriatrics
Clinical studies of COMIRNATY and COMIRNATY Original & Omicron BA.4/BA.5 included participants 65 years of age and older and their data contributes to the overall assessment of the safety and efficacy of COMIRNATY Omicron XBB.1.5 (See 8 ADVERSE REACTIONS and 14 CLINICAL TRIALS). The safety and efficacy of COMIRNATY Omicron XBB.1.5 in geriatric individuals have not yet been established in clinical trials.
8 ADVERSE REACTIONS
8.1 Adverse Reaction Overview
The safety of COMIRNATY Omicron XBB.1.5 is inferred from safety data of the prior COMIRNATY and COMIRNATY Original & Omicron BA.4/BA.5 vaccines.
Safety data accrued with the COMIRNATY and COMIRNATY Original & Omicron BA.4/BA.5 formulations are relevant to the COMIRNATY Omicron XBB.1.5 vaccine because these vaccines are manufactured using the same process.
8.1.1 COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg)
Participants ≥12 Years of Age – After a Dose of COMIRNATY Original & Omicron BA.4/BA.5 as a Second Booster (4th Dose)
Study C4591044 (Study 5) is an ongoing Phase 2/3 study to evaluate the safety, tolerability, and immunogenicity of new bivalent vaccines including COMIRNATY Original & Omicron BA.4/BA.5. In Cohorts 2 and 3 of the study 317 participants 12 years and older and 410 participants 18 years and older, respectively, received COMIRNATY Original & Omicron BA.4/BA.5 30 mcg (15/15 mcg) as a second booster dose following a previous primary series and one booster dose of COMIRNATY. The safety evaluation of participants in the study is ongoing. All participants were monitored for solicited local and systemic reactions and use of antipyretic medication after vaccination with an electronic diary during the 7 days following the dose of vaccination. Participants continue to be monitored for unsolicited adverse events (AEs), including serious adverse events (SAEs), throughout the study [from Dose 1 to 1 month after the last dose (all AEs) and 6 months (SAEs) after the last vaccination].
In a substudy from Study 5, 108 participants 12 to 17 years of age, 313 participants 18 to 55 years of age and 306 participants ≥56 years of age who had completed 3 doses of COMIRNATY, received a booster (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 (15/15 mcg) 5.4 to 16.9 months after receiving Dose 3. Participants who received a booster (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 had a median follow-up time of at least 1.5 months.
The overall safety profile for the COMIRNATY Original & Omicron BA.4/BA.5 booster (fourth dose) was similar to that seen after 3 doses of COMIRNATY. The most frequent adverse reactions in participants 12 years of age and older were injection site pain (>60%), fatigue (>50%), headache (>40%), muscle pain (>20%), chills (>10%), and joint pain (>10%).
8.1.2 COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg)
Participants 5 to <12 Years of Age – After a Dose of COMIRNATY Original & Omicron BA.4/BA.5 as a Second Booster (4th Dose)
Study C4591048 (Study 6) is an ongoing study to evaluate the safety, tolerability, and immunogenicity of new bivalent vaccines including COMIRNATY Original & Omicron BA.4/BA.5.
In a subset from Study 6 (Phase 3), 113 participants 5 to <12 years of age who had completed 3 doses of COMIRNATY, received a booster (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 (5/5 mcg) 2.6 to 8.5 months after receiving Dose 3. Participants who received a booster (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 had a median follow-up time of at least 1.6 months.
The overall safety profile for the COMIRNATY Original & Omicron BA.4/BA.5 booster (fourth dose) was similar to that seen after 3 doses of COMIRNATY. The most frequent adverse reactions in participants 5 to <12 years were injection site pain (>60%), fatigue (>40%), headache (>20%), and muscle pain (>10%).
8.1.3 COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg)
Study C4591048 (Study 6) is an ongoing study evaluating the safety, tolerability and immunogenicity of COMIRNATY Original & Omicron BA.4/BA.5. The safety of a 3-dose primary series of COMIRNATY Original & Omicron BA.4/BA.5 at 1.5 mcg/1.5 mcg in children 6 months to < 5 years of age is inferred primarily from the safety profile of COMIRNATY at 3 mcg administered as a 3-dose primary series in this age bracket. Safety data from study 6 in children 6 months to <5 years of age using the COMIRNATY Original & Omicron BA.4/BA.5 formulation at 1.5 mcg/1.5 mcg administered as a booster (fourth) dose are considered supportive.
Participants 2 Years Through <5 Years of Age – After a Dose of COMIRNATY Original & Omicron BA.4/BA.5 as a Booster (4th Dose)
In a subset from Study 6 (Phase 3), 124 participants 2 through <5 years of age who had completed 3 doses of COMIRNATY, received a booster of COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) 2.2 to 8.6 months after receiving Dose 3. Participants who received a booster of COMIRNATY Original & Omicron BA.4/BA.5 had a median follow-up time of at least 1.8 months up to a data cut-off date of 30 Nov 2022.
The overall safety profile for the COMIRNATY Original & Omicron BA.4/BA.5 booster was similar to that seen after 3 doses of COMIRNATY. The most frequent adverse reactions in participants 2 through <5 years of age were injection site pain (>30%) and fatigue (>20%).
Participants 6 months Through <2 Years of Age – After a Dose of COMIRNATY Original & Omicron BA.4/BA.5 as a Booster (4th Dose)
In a subset from Study 6 (Phase 3), 39 participants 6 months to <2 years of age who had completed 3 doses of COMIRNATY, received a booster of COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg)
2.1 to 8.6 months after receiving Dose 3. Participants who received a booster of COMIRNATY Original & Omicron BA.4/BA.5 had a median follow-up time of at least 1.7 months up to data cut-off of 30 Nov 2022.
The overall safety profile for the COMIRNATY Original & Omicron BA.4/BA.5 booster was similar to that seen after 3 doses of COMIRNATY. The most frequent adverse reactions in participants 6 months to <2 years of age were irritability (>20%) and decreased appetite (>10%).
8.1.4 COMIRNATY (30 mcg)
Study BNT162-01 (Study 1) was a Phase 1/2, two-part dose-escalation trial that enrolled 60 participants 18 through 55 years of age and 36 participants 56 through 85 years of age.
Study C4591001 (Study 2) is a Phase 1/2/3, multicenter, multinational, randomized, saline placebo-controlled, observer-blind, dose-finding, vaccine candidate-selection (Phase 1) and efficacy (Phase 2/3) study that has enrolled approximately 46,000 participants, 12 years of age or older. Of these, approximately 44,047 participants (22,026 COMIRNATY; 22,021 placebo) in Phase 2/3 are 16 years of age or older (including 378 and 376 adolescents 16 and 17 years of age in the vaccine and placebo groups, respectively) and 2,260 adolescents are 12 to 15 years of age (1,131 and 1,129 in the vaccine and placebo groups, respectively). Of the total number of COMIRNATY recipients in the study, 20.7% were 65 years of age and older. Study 2 also included 200 participants with confirmed stable human immunodeficiency virus (HIV) infection. HIV-positive participants are included in safety population disposition but are summarized separately in safety analyses.
Additionally, 306 existing Phase 3 participants 18 through 55 years of age received a booster dose of COMIRNATY approximately 6 months (range of 4.8 to 8.0 months) after completing the second dose in the non-placebo-controlled booster dose portion of Study 2. The overall safety profile for the booster dose was similar to that seen after 2 doses.
In Study C4591031 (Study 4), a placebo-controlled booster study, 5,081 participants 16 years of age and older were recruited from Study 2 to receive a booster dose of COMIRNATY at least 6 months after the second dose. The overall safety profile for the booster dose was similar to that seen after 2 doses.
The safety evaluation of participants in Study 2 and Study 4 is ongoing. In Study 2, all participants 12 to 15 years of age and 16 years of age and older in the reactogenicity subset, and a subset of 306 participants 18 through 55 years of age who received a booster dose in Study 2, were monitored for solicited local and systemic reactions and use of antipyretic medication after each vaccination with an electronic diary during the 7 days following any dose of vaccination. Participants, including those who received a booster in Study 4, continue to be monitored for unsolicited adverse events (AEs), including serious adverse events (SAEs), throughout the study [from Dose 1 to 1 month after the last dose (all AEs) and 6 months (SAEs) after the last vaccination].
Participants 12 Years of Age and Older
At the time of the analysis of Study 2 (data accrued through March 13, 2021), a total of 25,651 (58.2%) participants (13,031 in vaccine group and 12,620 in placebo group) 16 years of age and older had been followed up for at least 4 months, with 3,082 (7.0%) participants (1,778 in vaccine group and 1,304 in placebo group) followed up for at least 6 months after the second dose during the blinded placebo-controlled follow-up period. A total of 12,006 (54.5%) participants originally randomized to the vaccine group in Study 2 had been followed up for at least 6 months after the second dose including the blinded and open-label periods.
In an analysis of Study 2, based on data up to the cut-off date of March 13, 2021, a total of 2,260 adolescents (1,131 COMIRNATY; 1,129 placebo) were 12 to 15 years of age. Of these,1,559 (786 COMIRNATY and 773 placebo) adolescents have been followed for ≥4 months after the second dose of COMIRNATY.
In clinical studies with a data cut-off of March 13, 2021, and where 2 doses were administered 3 weeks apart, the most common adverse reactions in the reactogenicity subset (n=4,924) of participants 16 years of age and older after any dose included injection site pain (84.3%), fatigue (64.7%), headache (57.1%), muscle pain (40.2%), chills (34.7%), joint pain (25.0%), fever (15.2%), injection site swelling (11.1%), and injection site redness (9.9%). Additional AEs reported in the safety population (n=21,926) of participants 16 years of age and older from dose 1 to 1 month after dose 2 included nausea (1.2%), malaise (0.6%), lymphadenopathy (0.4%), asthenia (0.3%), decreased appetite (0.2%), hyperhidrosis (0.1%), lethargy (0.1%), and night sweats (0.1%). Adverse reactions were usually mild or moderate in intensity and resolved within a few days after vaccination.
The safety profile in 545 participants receiving COMIRNATY that were seropositive for SARS-CoV-2 at baseline was similar to that seen in the general population.
In a clinical study with a data cut-off date of 02 September 2021, the most commonly reported (≥8%) adverse reactions in adolescents 12 through 15 years of age following any dose were pain at the injection site (90.5%), fatigue (77.5%), headache (75.5%), chills (49.2%), muscle pain (42.2%), fever (24.3%), joint pain (20.2%), injection site swelling (9.2%), and injection site redness (8.6%).
In a clinical study of participants 18 through 55 years of age (N=306), 289 participants (94%) completed the e-diary recording adverse reactions. The most commonly reported adverse reactions (≥10%) following administration of a booster dose were pain at the injection site (83.0%), fatigue (63.7%), headache (48.4%), muscle pain (39.1%), chills (29.1%), and joint pain (25.3%).
In a clinical study of approximately 10,000 participants 16 years of age and older, unsolicited adverse reactions following administration of a booster dose included headache (5%), fever (4.8%), lymphadenopathy (2.8%), pain in extremity (1.1%), nausea (0.9%), malaise (0.7%), and decreased appetite (0.2%).
8.1.5 COMIRNATY (10 mcg)
Study C4591007 (Study 3) is a Phase 1/2/3 study comprised of an open-label vaccine dose-finding portion (Phase 1) and a multicenter, multinational, randomized, saline placebo-controlled, observer-blind immunogenicity and efficacy portion (Phase 2/3) that has enrolled approximately 4,600 participants 5 years through <12 years of age. Of these, approximately 3,100 participants received COMIRNATY 10 mcg and approximately 1,500 participants received placebo in the Phase 2/3 part of the study. Study 3 also enrolled 1,776 participants 6 months through <2 years of age (1,178 COMIRNATY 3 mcg; 598 placebo), and 2,750 participants 2 through <5 years of age (1,835 COMIRNATY 3 mcg; 915 placebo) in Phase 2/3.
In a subset of Study 3 Phase 2/3 participants, 401 participants 5 years through <12 years of age received a booster dose of COMIRNATY at least 5 months (range 5 to 9 months) after completing the primary series. The overall safety profile for the booster dose was similar to that seen after the primary series. The analysis of the Study 3 Phase 2/3 subset is based on data up to the cut-off date of March 22, 2022 (median follow-up time of 1.3 months).
Participants 5 Years Through <12 Years of Age
In an analysis of Study 3 Phase 2/3, based on data up to the cut-off date of October 8, 2021, 2,268 participants (initial enrolment group: 1,518 COMIRNATY 10 mcg and 750 placebo) were 5 years through <12 years of age. Of these, 2,171 (95.7%) (1,456 COMIRNATY 10 mcg and 715 placebo) participants have been followed for at least 3 months after Dose 2. An analysis of Study 3 Phase 2/3 adverse event data also included another 2,379 participants (safety expansion group: 1,591 COMIRNATY 10 mcg and 788 placebo), of whom 71.2% had a follow-up period for at least 2 weeks after Dose 2. The safety evaluation in Study 3 is ongoing.
Adverse reactions following administration of any dose in the initial enrolment safety population (n = 1,518) of children 5 years through <12 years of age included pain at the injection site (84.3%), fatigue (51.7%), headache (38.2%), injection site redness (26.4%), injection site swelling (20.4%), muscle pain (17.5%), chills (12.4%), fever (8.3%), joint pain (7.6%), lymphadenopathy (0.9%), rash (0.5%), nausea (0.4%), malaise (0.1%), and decreased appetite (0.1%).
The most frequent adverse reactions in participants 5 years through <12 years of age following the booster dose (data cut-off date of March 22, 2022; median follow-up time of 1.3 months) were injection site pain (73.9%), fatigue (45.6%), headache (34.0%), myalgia (18.3%), chills (10.5%), injection site redness (15.6%), and swelling (16.4%). The most frequently reported unsolicited adverse event was lymphadenopathy (2.5%).
8.1.6 COMIRNATY (3 mcg)
Participants 2 Through <5 Years of Age
Study 3 (Phase 2/3) enrolled 2,750 participants 2 through <5 years of age (1,835 COMIRNATY 3 mcg; 915 placebo). Of these, 2,726 participants (1,819 COMIRNATY 3 mcg; 907 placebo) received 2 doses and 1,369 (50.2%; 910 COMIRNATY 3 mcg and 459 placebo) participants have been followed for at least 4 months after the second dose; 886 participants received a 3-dose primary series (606 COMIRNATY 3 mcg; 280 placebo) and have been followed for a median of 1.4 months after the third dose, based on data in the blinded, placebo-controlled follow-up period up to the cut-off date of April 29, 2022. Adverse reactions following administration of any dose included pain at the injection site (47.0%), fatigue (44.8%), injection site redness (18.9%), fever (10.5%), headache (8.7%), injection site swelling (8.4%), chills (5.7%), muscle pain (5.0%), joint pain (2.4%), and lymphadenopathy (0.1%).
Participants 6 Months Through <2 Years of Age
Study 3 (Phase 2/3) also enrolled 1,776 participants 6 months through <2 years of age (1,178 COMIRNATY 3 mcg; 598 placebo). Of these, 1,762 participants (1,166 COMIRNATY 3 mcg; 596 placebo) received 2 doses and 1,207 (68.5%; 801 COMIRNATY 3 mcg and 406 placebo) participants have been followed for at least 4 months after the second dose; 570 participants received a 3-dose primary series (386 COMIRNATY 3 mcg; 184 placebo) and have been followed for a median of 1.3 months after the third dose, based on data in the blinded, placebo-controlled follow-up period up to the cut-off date of April 29, 2022. Adverse reactions following administration of any dose included irritability (68.4%), decreased appetite (38.6%), tenderness at the injection site (26.4%), injection site redness (17.8%), fever (14.4%), injection site swelling (7.3%), and lymphadenopathy (0.2%).
8.2 Clinical Trial Adverse Reactions
Clinical trials are conducted under very specific conditions. The adverse reaction rates observed in the clinical trials, therefore, may not reflect the rates observed in practice and should not be compared to the rates in the clinical trials of another drug. Adverse reaction information from clinical trials may be useful in identifying and approximating rates of adverse drug reactions in real-world use.
8.2.1 COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg)
Participants 12 Years of Age and Older
Solicited Local Adverse Reactions
Table 3 presents the frequency of reported solicited local reactions within 7 days of a second booster dose with COMIRNATY Original & Omicron BA.4/BA.5.
Most local reactions were mild or moderate in severity and no Grade 4 local reactions were reported in any group. The median onset for all local reactions was 1 to 3 days, and all events resolved within a median duration of 1 to 3 days after onset.
| Table 3: Study 5 - Solicited Local Adverse Reactions Reported for Vaccine Groups Within 7 Days After Study Vaccination | ||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg) | |||||||||||||
12– 17 years Na=107 nb | 18 – 55 years (Na=310) nb | >55 years (Na=300) nb | ||||||||||||
| Rednessc | ||||||||||||||
| Any | 6 (5.6) | 21 (6.8) | 11 (3.7) | |||||||||||
| Severe | 0 | 0 | 0 | |||||||||||
| Swellingc | ||||||||||||||
| Any | 8 (7.5) | 23 (7.4)) | 8 (2.7) | |||||||||||
| Severe | 0 | 0 | 0 | |||||||||||
| Pain at the injection sited | ||||||||||||||
| Any | 75 (70.1) | 236 (76.1) | 172 (57.1)e | |||||||||||
| Severe | 1 (0.9) | 0 | 1 (0.3)e | |||||||||||
| Any local reactionf | 75 (70.1) | 240 (77.4) | 174 (57.8)e | |||||||||||
a. N = number of participants reporting at least 1 yes or no response for the specified reaction after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. Mild: >2.0 to 5.0 cm; moderate: >5.0 to 10.0 cm; severe: >10.0 cm; Grade 4: necrosis (redness and swelling categories) or exfoliative dermatitis (redness category only).
d. Mild: does not interfere with activity; moderate: interferes with activity; severe: prevents daily activity; Grade 4: emergency room visit or hospitalization for severe pain at the injection site.
e. N=301
f. Any local reaction: any redness >2.0 cm, any swelling >2.0 cm, or any pain at the injection site.
Solicited Systemic Adverse Reactions
Table 4 presents the frequency of reported systemic reactions within 7 days of a second booster dose of COMIRNATY Original & Omicron BA.4/BA.5. Most systemic reactions were mild or moderate in severity and no Grade 4 systemic reactions were reported in any group. The median onset for all systemic reactions was 2 to 4 days, and all events resolved within a median duration of 1 to 2 days after onset.
| Table 4: Study 5 - Solicited Systemic Adverse Reactions Reported for Vaccine Groups Within 7 Days After Study Vaccination | |||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (15 mcg/15 mcg) | ||||||||||||||
12– 17 years Na=107 nb | 18 – 55 years (Na=309) nb | >55 years (Na=300) nb | |||||||||||||
| Fever | |||||||||||||||
| ≥38.0°C | 10 (9.3) | 15 (4.9) | 13 (4.3) | ||||||||||||
| ≥38.9°C to 40.0°C | 1 (0.9) | 0 | 0 | ||||||||||||
| Fatiguec | |||||||||||||||
| Any | 72 (67.3) | 189 (61.2) | 116 (38.5)d | ||||||||||||
| Severe | 0 | 6 (1.9) | 4 (1.3)d | ||||||||||||
| Headachec | |||||||||||||||
| Any | 54 (50.5) | 144 (46.6) | 92 (30.7) | ||||||||||||
| Severe | 0 | 2 (0.6) | 0 | ||||||||||||
| Chillsc | |||||||||||||||
| Any | 25 (23.4) | 68 (22.0) | 36 (12.0) | ||||||||||||
| Severe | 0 | 2 (0.6) | 1 (0.3) | ||||||||||||
| Vomitinge | |||||||||||||||
| Any | 3 (2.8) | 6 (1.9) | 29 (2.7) | ||||||||||||
| Severe | 0 | 0 | 0 | ||||||||||||
| Diarrheaf | |||||||||||||||
| Any | 7 (6.5) | 33 (10.7) | 29 (9.6)d | ||||||||||||
| Severe | 0 | 1 (0.3) | 0d | ||||||||||||
| New or worsened muscle painc | |||||||||||||||
| Any | 28 (26.2) | 94 (30.4) | 54 (18.0) | ||||||||||||
| Severe | 0 | 0 | 0 | ||||||||||||
| New or worsened joint painc | |||||||||||||||
| Any | 13 (12.1)) | 46 (14.9) | 36 (12.0) | ||||||||||||
| Severe | 1 (0.9) | 0 | 0 | ||||||||||||
| Any systemic eventg | 86 (80.4) | 229 (74.1) | 12 (53.8)d | ||||||||||||
| Use of antipyretic or pain medicationh | 36 (33.6) | 105 (34.0) | 74 (24.7) | ||||||||||||
a. N = number of participants reporting at least 1 yes or no response for the specified event after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. Mild: does not interfere with activity; moderate: some interference with activity; severe: prevents daily activity; Grade 4: emergency room visit or hospitalization for severe fatigue, severe headache, severe chills, severe muscle pain, or severe joint pain.
d. N=301
e. Mild: 1 to 2 times in 24 hours; moderate: >2 times in 24 hours; severe: requires intravenous hydration; Grade 4: emergency room visit or hospitalization for severe vomiting.
f. Mild: 2 to 3 loose stools in 24 hours; moderate: 4 to 5 loose stools in 24 hours; severe: 6 or more loose stools in 24 hours; Grade 4: emergency room visit or hospitalization for severe diarrhea.
g. Any systemic event: any fever, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new or worsened joint pain.
h. Severity was not collected for use of antipyretic or pain medication.
Unsolicited Adverse Events
Among participants 12 years of age and older, unsolicited adverse events were reported by 48 (6.6%) participants who received a second booster dose through 1 month after the booster dose. Lymphadenopathy occurred in 7 (1.0%) participants.
8.2.2 COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg)
Participants 5 to <12 Years of Age
Solicited Local Adverse Reactions
Table 5 presents the frequency of reported solicited local reactions within 7 days of a second booster dose with COMIRNATY Original & Omicron BA.4/BA.5.
All local reactions were mild or moderate in severity. The median onset for all local reactions was 1 to 2 days, and all events resolved within a median duration of 2 days after onset.
| Table 5: Study 6 - Solicited Local Adverse Reactions Reported Within 7 Days After Study Vaccination | |||||
|---|---|---|---|---|---|
| Local Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg) (N=111)a nb % | ||||
| Rednessc | |||||
| Any | 8 (7.2) | ||||
| Severe | 0 | ||||
| Swellingc | |||||
| Any | 5 (4.5) | ||||
| Severe | 0 | ||||
| Pain at the injection sited | |||||
| Any | 71 (64.0) | ||||
| Severe | 0 | ||||
| Any local reactione | 74 (66.7) | ||||
a. N = number of participants reporting at least 1 yes or no response for the specified reaction after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. Mild: ≥0.5 to 2.0 cm; moderate: >2.0 to 7.0 cm; severe: >7.0 cm; Grade 4: necrosis (redness and swelling categories) or exfoliative dermatitis (redness category only).
d. Mild: does not interfere with activity; moderate: interferes with activity; severe: prevents daily activity; Grade 4: emergency room visit or hospitalization for severe pain at the injection site.
e. Any local reaction: any redness ≥0.5 cm, any swelling ≥0.5 cm, or any pain at the injection site.
Solicited Systemic Adverse Reactions
Table 6 presents the frequency of reported solicited systemic reactions within 7 days of a second booster dose with COMIRNATY Original & Omicron BA.4/BA.5.
Most systemic events were mild or moderate in severity, and no Grade 4 systemic events were reported. The median onset for all systemic events was 2 to 4 days, and all events resolved within a median duration of 1 to 2 days after onset.
| Table 6: Study 6 - Solicited Systemic Adverse Reactions Reported Within 7 Days After Study Vaccination | |||||
|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (5 mcg/5 mcg) (N=111)a nb % | ||||
| Fever | |||||
| ≥38.0°C | 5 (4.5) | ||||
| ≥38.9°C to 40.0°C | 2 (1.8) | ||||
| Fatiguec | |||||
| Any | 45 (40.5) | ||||
| Severe | 1 (0.9) | ||||
| Headachec | |||||
| Any | 28 (25.2) | ||||
| Severe | 1 (0.9) | ||||
| Chillsc | |||||
| Any | 10 (9.0) | ||||
| Severe | 0 | ||||
| Vomitingd | |||||
| Any | 4 (3.6) | ||||
| Severe | 0 | ||||
| Diarrheae | |||||
| Any | 4 (3.6) | ||||
| Severe | 0 | ||||
| New or worsened muscle painc | |||||
| Any | 15 (13.5) | ||||
| Severe | 0 | ||||
| New or worsened joint painc | |||||
| Any | 10 (9.0) | ||||
| Severe | 0 | ||||
| Any systemic eventf | 58 (52.3) | ||||
| Use of antipyretic or pain medicationg | 26 (23.4) | ||||
a. N = number of participants reporting at least 1 yes or no response for the specified event after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. Mild: does not interfere with activity; moderate: some interference with activity; severe: prevents daily activity; Grade 4: emergency room visit or
hospitalization for severe fatigue, severe headache, severe chills, severe muscle pain, or severe joint pain.
d. Mild: 1 to 2 times in 24 hours; moderate: >2 times in 24 hours; severe: requires intravenous hydration; Grade 4: emergency room visit or hospitalization
for severe vomiting.
e. Mild: 2 to 3 loose stools in 24 hours; moderate: 4 to 5 loose stools in 24 hours; severe: 6 or more loose stools in 24 hours; Grade 4: emergency room visit
or hospitalization for severe diarrhea.
f. Any systemic event: any fever ≥38.0℃, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new
or worsened joint pain.
g. Severity was not collected for use of antipyretic or pain medication.
Unsolicited Adverse Events
Among participants 5 to <12 years of age, unsolicited adverse events were reported by 4 (3.5%) participants who received a second booster dose through 1 month after the booster dose. Lymphadenopathy occurred in 1 (0.9%) participant.
8.2.3 COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg)
Participants 2 Through <5 Years of Age – After a Dose of COMIRNATY Original & Omicron BA.4/BA.5 as a Booster (4th Dose)
Solicited Local Adverse Reactions
Table 7 presents the frequency of solicited local reactions within 7 days of a booster (fourth) dose with COMIRNATY Original & Omicron BA.4/BA.5. Most local reactions were mild or moderate in severity. No severe or Grade 4 local reactions were reported. The onset for all local reactions was 1 to 2 days, and all events resolved within 1 to 3 days after onset.
| Table 7: Study 6 - Solicited Local Adverse Reactions Reported Within 7 Days After a Booster (Fourth Dose) – Participants 2 Through <5 Years of Age – Safety Population | |||||
|---|---|---|---|---|---|
| Local Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) (N=124)a nb % | ||||
| Rednessc | |||||
| Any | 10 (8.1)) | ||||
| Severe | 0 | ||||
| Swellingc | |||||
| Any | 7 (5.6) | ||||
| Severe | 0 | ||||
| Pain at the injection sited | |||||
| Any | 39 (31.5) | ||||
| Severe | 0 | ||||
| Any local reactione | 48 (38.7) | ||||
a. N = number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose.
b. n = Number of participants with the specified characteristic.
c. Mild: ≥0.5 to 2.0 cm; moderate: >2.0 to 7.0 cm; severe: >7.0 cm; Grade 4: necrosis (redness and swelling categories) or exfoliative dermatitis (redness category only).
d. Mild: does not interfere with activity; moderate: interferes with activity; severe: prevents daily activity; Grade 4: emergency room visit or hospitalization for severe pain at the injection site.
e. Any local reaction: any redness ≥0.5 cm, any swelling ≥0.5 cm, or any pain at the injection site.
Solicited Systemic Adverse Reactions
Table 8 presents the frequency of solicited systemic reactions within 7 days of a booster (fourth) dose with COMIRNATY Original & Omicron BA.4/BA.5. Most systemic reactions were mild or moderate in severity. No severe or Grade 4 systemic reactions were reported. The median onset for most systemic reactions was 1 to 6 days, and most events resolved within a median duration of 1 to 2 days after onset.
| Table 8: Study 6 - Solicited Systemic Reactions Reported Within 7 Days After a Booster (Fourth Dose) – Participants 2 Through <5 Years of Age – Safety Population | |||||
|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) (N=123)a nb % | ||||
| Fever | |||||
| ≥38.0°C | 6 (4.8)c | ||||
| ≥38.9°C to 40.0°C | 0 c | ||||
| Fatigued | |||||
| Any | 36 (29.3) | ||||
| Severe | 0 | ||||
| Headached | |||||
| Any | 5 (4.1) | ||||
| Severe | 2 (1.6) | ||||
| Chillsd | |||||
| Any | 7 (5.7) | ||||
| Severe | 0 | ||||
| Vomitinge | |||||
| Any | 7 (5.7) | ||||
| Severe | 0 | ||||
| Diarrheaf | |||||
| Any | 6 (4.9) | ||||
| Severe | 0 | ||||
| New or worsened muscle paind | |||||
| Any | 4 (3.3) | ||||
| Severe | 0 | ||||
| New or worsened joint paind | |||||
| Any | 2 (1.6) | ||||
| Severe | 0 | ||||
| Any systemic eventg | 45 (36.3)c | ||||
| Use of antipyretic or pain medicationh | 14 (11.3)c | ||||
a. N = number of participants reporting at least 1 yes or no response for the specified event after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. N = 124
d. Mild: does not interfere with activity; moderate: some interference with activity; severe: prevents daily activity; Grade 4: emergency room visit or hospitalization for severe fatigue, severe headache, severe chills, severe new or worsened muscle pain, or severe new or worsened joint pain.
e. Mild: 1 to 2 times in 24 hours; moderate: >2 times in 24 hours; severe: requires intravenous hydration; Grade 4: emergency room visit or hospitalization for hypotensive shock.
f. Mild: 2 to 3 loose stools in 24 hours; moderate: 4 to 5 loose stools in 24 hours; severe: 6 or more loose stools in 24 hours; Grade 4: emergency room visit or hospitalization for severe diarrhea.
g. Any systemic event: any fever ≥38.0℃, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new or worsened joint pain.
h. Severity was not collected for use of antipyretic or pain medication.
Participants 6 Months Through <2 Years of Age
Solicited Local Adverse Reactions
Table 9 presents the frequency of solicited local reactions within 7 days of a booster (fourth) dose with COMIRNATY Original & Omicron BA.4/BA.5. All local reactions were mild in severity. No moderate, severe or Grade 4 local reactions were reported. The onset for all local reactions was 1 day, and all events resolved within 1 day after onset.
| Table 9: Study 6 - Solicited Local Adverse Reactions Reported Within 7 Days After a Booster (Fourth Dose) – Participants 6 Months Through <2 Years of Age – Safety Population | ||||
|---|---|---|---|---|
| Local Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) (N=39)a nb % | |||
| Rednessc | ||||
| Any | 2 (5.1) | |||
| Severe | 0 | |||
| Swellingc | ||||
| Any | 1 (2.6) | |||
| Severe | 0 | |||
| Tenderness at the injection sited | ||||
| Any | 2 (5.3)e | |||
| Severe | 0e | |||
| Any local reactionf | 3 (7.7) | |||
a. N = number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose.
b. n = Number of participants with the specified characteristic.
c. Mild: ≥0.5 to 2.0 cm; moderate: >2.0 to 7.0 cm; severe: >7.0 cm; Grade 4: necrosis (redness and swelling categories) or exfoliative dermatitis (redness category only).
d. Mild: hurts if gently touched; moderate: hurts if gently touched with crying; severe: causes limitation of limb movement; Grade 4: emergency room visit or hospitalization for severe pain (tenderness) at the injection site.
e. N = 38
f. Any local reaction: any redness ≥0.5 cm, any swelling ≥0.5 cm, or any pain at the injection site.
Solicited Systemic Adverse Reactions
Table 10 presents the frequency of solicited systemic reactions within 7 days of a booster (fourth) dose with COMIRNATY Original & Omicron BA.4/BA.5. Most systemic reactions were mild or moderate in severity. No severe or Grade 4 systemic reactions were reported. The median onset for all systemic reactions was 2 to 6 days, and most events resolved within a median duration of 1 to 3 days after onset.
| Table 10: Study 6 - Solicited Systemic Reactions Reported Within 7 Days After a Booster (Fourth Dose) – Participants 6 Months Through <2 Years of Age – Safety Population | |||||
|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) (N=39)a nb % | ||||
| Fever | |||||
| ≥38.0°C | 2 (5.1) | ||||
| ≥38.9°C to 40.0°C | 0 | ||||
| Decreased appetitec | |||||
| Any | 7 (18.9)d | ||||
| Severe | 0d | ||||
| Drowsinesse | |||||
| Any | 4 (10.8)d | ||||
| Severe | 0d | ||||
| Irritabilityf | |||||
| Any | 11 (29.7)d | ||||
| Severe | 0d | ||||
| Any systemic eventg | 13 (33.3) | ||||
| Use of antipyretic or pain medicationh | 3 (7.7) | ||||
a. N = number of participants reporting at least 1 yes or no response for the specified event after the study vaccination.
b. n = Number of participants with the specified characteristic.
c. Mild: decreased interest in eating; Moderate: decreased oral intake; Severe: refusal to feed; Grade 4: emergency room visit or hospitalization for severe decreased appetite (loss of appetite). d. N=37
e. Mild: increased or prolonged sleeping bouts; Moderate: slightly subdued interfering with daily activity; Severe: disabling; not interested in usual daily activity; Grade 4: emergency room visit or hospitalization for severe drowsiness (increased sleep).
f. Mild: easily consolable; moderate: requiring increased attention; severe: inconsolable; crying cannot be comforted; Grade 4: emergency room visit or hospitalization for severe irritability (fussiness).
g. Any systemic event: any fever ≥38.0℃, any decreased appetite, any drowsiness, or any irritability.
h. Severity was not collected for use of antipyretic or pain medication.
Unsolicited Adverse Events
The safety population included 39 participants ≥6 months to <2 years of age and 124 participants ≥2 years to <5 years of age who received a fourth dose of COMIRNATY Original & Omicron BA.4/BA.5 at 3 mcg. Overall, median (min, max) follow-up time after study vaccination was 1.8 (1.3, 2.5) months.
Overall, AEs were reported by 6 (15.4%) and 6 (4.8%) participants in the ≥6 months to <2 years of age group and ≥2 to <5 years of age group, respectively. No severe AEs, life-threatening AEs, SAEs, or AEs leading to withdrawal or death were reported from study vaccination to 1 month after study vaccination.
From study vaccination through 1-month postvaccination, no AEs of lymphadenopathy, rash, anaphylaxis/hypersensitivity, appendicitis, Bell’s palsy, and myo/pericarditis were reported.
8.2.4 COMIRNATY (30 mcg)
Participants 16 Years of Age and Older – Primary Series (Two Doses)
Solicited Adverse Reactions
Tables 11 through 14 present the frequency and severity of solicited local and systemic reactions, respectively, within 7 days following each dose of COMIRNATY and placebo in the subset of participants 16 years of age and older (n=9,839) in the safety population who were monitored for reactogenicity with an electronic diary.
| Table 11: Study 2 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose of COMIRNATY– Participants 16-55 Years of Age (Reactogenicity Subset of the Safety Population*) | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | Dose 1 | Dose 2 | |||||||||||
COMIRNATY Na=2,899 nb (%) | Placebo Na=2,908 nb (%) | COMIRNATY Na=2,682 nb (%) | Placebo Na=2,684 nb (%) | ||||||||||
| Redness | |||||||||||||
| Anyc | 156 (5.4) | 28 (1.0) | 151 (5.6) | 18 (0.7) | |||||||||
| Severed | 7 (0.2) | 3 (0.1) | 11 (0.4) | 0 (0.0) | |||||||||
| Swelling | |||||||||||||
| Anyc | 184 (6.3) | 16 (0.6) | 183 (6.8) | 5 (0.2) | |||||||||
| Severed | 6 (0.2) | 2 (0.1) | 7 (0.3) | 0 (0.0) | |||||||||
| Pain at the injection site | |||||||||||||
| Anyc | 2,426 (83.7) | 414 (14.2) | 2,101 (78.3) | 312 (11.6) | |||||||||
| Severee | 39 (1.3) | 3 (0.1) | 39 (1.5) | 0 (0.0) | |||||||||
| Any local reactionc | 2,444 (84.3) | 432 (14.9) | 2,108 (78.6) | 325 (12.1) | |||||||||
*Randomized participants in the safety analysis population who received at least 1 dose of the study intervention.
a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. No Grade 4 solicited local reactions were reported in participants 16-55 years of age.
b. n = Number of participants with the specified reaction.
c. Any local reaction: any redness >2.0 cm, any swelling >2.0 cm, or any pain at the injection site.
d. Severe: >10.0 cm.
e. Severe: prevents daily activity.
| Table 12: Study 2 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose of COMIRNATY – Participants 16-55 Years of Age (Reactogenicity Subset of the Safety Population*) | |||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | Dose 1 | Dose 2 | |||||||||||||
COMIRNATY Na=2,899 nb (%) | Placebo Na=2,908 nb (%) | COMIRNATY Na=2,682 nb (%) | Placebo Na=2,684 nb (%) | ||||||||||||
| Fever | |||||||||||||||
| ≥38.0°C | 119 (4.1) | 25 (0.9) | 440 (16.4) | 11 (0.4) | |||||||||||
| >38.9°C | 8 (0.3) | 4 (0.1) | 40 (1.5) | 2 (0.1) | |||||||||||
| Fatigue | |||||||||||||||
| Any | 1,431 (49.4) | 960 (33.0) | 1,649 (61.5) | 614 (22.9) | |||||||||||
| Severed | 41 (1.4) | 18 (0.6) | 142 (5.3) | 14 (0.5) | |||||||||||
| Headache | |||||||||||||||
| Any | 1,262 (43.5) | 975 (33.5) | 1,448 (54.0) | 652 (24.3) | |||||||||||
| Severed | 33 (1.1) | 24 (0.8) | 91 (3.4) | 18 (0.7) | |||||||||||
| Chills | |||||||||||||||
| Any | 479 (16.5) | 199 (6.8) | 1,015 (37.8) | 114 (4.2) | |||||||||||
| Severed | 15 (0.5) | 2 (0.1) | 69 (2.6) | 2 (0.1) | |||||||||||
| Vomiting | |||||||||||||||
| Any | 34 (1.2) | 36 (1.2) | 58 (2.2) | 30 (1.1) | |||||||||||
| Severee | 0 (0.0) | 1 (0.0) | 4 (0.1) | 0 (0.0) | |||||||||||
| Diarrhea | |||||||||||||||
| Any | 309 (10.7) | 323 (11.1) | 269 (10.0) | 205 (7.6) | |||||||||||
| Severef | 3 (0.1) | 1 (0.0) | 6 (0.2) | 1 (0.0) | |||||||||||
| New or worsened muscle pain | |||||||||||||||
| Any | 664 (22.9) | 329 (11.3) | 1,055 (39.3) | 237 (8.8) | |||||||||||
| Severed | 15 (0.5) | 2 (0.1) | 62 (2.3) | 3 (0.1) | |||||||||||
| New or worsened joint pain | |||||||||||||||
| Any | 342 (11.8) | 168 (5.8) | 638 (23.8) | 147 (5.5) | |||||||||||
| Severed | 5 (0.2) | 1 (0.0) | 27 (1.0) | 4 (0.1) | |||||||||||
| Any systemic reactionc | 1,979 (68.3) | 1,559 (53.6) | 2,034 (75.8) | 1,026 (38.2) | |||||||||||
| Use of antipyretic or pain medication | 805 (27.8) | 398 (13.7) | 1,213 (45.2) | 320 (11.9) | |||||||||||
*Randomized participants in the safety analysis population who received at least 1 dose of the study intervention.
a. N =Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. No Grade 4 solicited systemic reactions were reported in participants 16-55 years of age.
b. n = Number of participants with the specified reaction.
c. Any systemic reaction: any fever ≥38.0°C, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new or worsened joint pain.
d. Severe: prevents daily activity.
e. Severe: requires intravenous hydration.
f. Severe: 6 or more loose stools in 24 hours.
| Table 13: Study 2 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose of COMIRNATY – Participants 56 Years of Age and Older (Reactogenicity Subset of the Safety Population*) | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | Dose 1 | Dose 2 | |||||||||||
COMIRNATY Na=2,008 nb (%) | Placebo Na=1,989 nb (%) | COMIRNATY Na=1,860 nb (%) | Placebo Na=1,833 nb (%) | ||||||||||
| Redness | |||||||||||||
| Anyc | 106 (5.3) | 20 (1.0) | 133 (7.2) | 14 (0.8) | |||||||||
| Severed | 5 (0.2) | 2 (0.1) | 10 (0.5) | 1 (0.1) | |||||||||
| Swelling | |||||||||||||
| Anyc | 141 (7.0) | 23 (1.2) | 145 (7.8) | 13 (0.7) | |||||||||
| Severed | 2 (0.1) | 0 (0.0) | 4 (0.2) | 1 (0.1) | |||||||||
| Pain at the injection site | |||||||||||||
| Anyc | 1,408 (70.1) | 185 (9.3) | 1,230 (66.1) | 143 (7.8) | |||||||||
| Severee | 4 (0.2) | 0 (0.0) | 10 (0.5) | 0 (0.0) | |||||||||
| Any local reactionc | 1,433 (71.4) | 207 (10.4) | 1,243 (66.8) | 158 (8.6) | |||||||||
*Randomized participants in the safety analysis population who received at least 1 dose of the study intervention.
a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. No Grade 4 solicited local reactions were reported in participants 56 years of age and older.
b. n = Number of participants with the specified reaction.
c. Any local reaction: any redness >2.0 cm, any swelling >2.0 cm, or any pain at the injection site.
d. Severe: >10.0 cm.
e. Severe: prevents daily activity.
| Table 14: Study 2 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose of COMIRNATY – Participants 56 Years of Age and Older (Reactogenicity Subset of the Safety Population*) | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | Dose 1 | Dose 2 | |||||||||||
COMIRNATY Na=2,008 nb (%) | Placebo Na=1,989 nb (%) | COMIRNATY Na=1,860 nb (%) | Placebo Na=1,833 nb (%) | ||||||||||
| Fever | |||||||||||||
| ≥38.0°C | 26 (1.3) | 8 (0.4) | 219 (11.8) | 4 (0.2) | |||||||||
| >38.9°C | 1 (0.0) | 2 (0.1) | 7 (0.4) | 1 (0.1) | |||||||||
| Fatigue | |||||||||||||
| Any | 677 (33.7) | 447 (22.5) | 949 (51.0) | 306 (16.7) | |||||||||
| Severed | 3 (0.1) | 3 (0.2) | 60 (3.2) | 2 (0.1) | |||||||||
| Grade 4g | 0 (0.0) | 0 (0.0) | 1 (0.1) | 0 (0.0) | |||||||||
| Headache | |||||||||||||
| Any | 503 (25.0) | 363 (18.3) | 733 (39.4) | 259 (14.1) | |||||||||
| Severed | 2 (0.1) | 3 (0.2) | 13 (0.7) | 5 (0.3) | |||||||||
| Chills | |||||||||||||
| Any | 130 (6.5) | 69 (3.5) | 435 (23.4) | 57 (3.1) | |||||||||
| Table 14: Study 2 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose of COMIRNATY – Participants 56 Years of Age and Older (Reactogenicity Subset of the Safety Population*) | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | Dose 1 | Dose 2 | |||||||||||
COMIRNATY Na=2,008 nb (%) | Placebo Na=1,989 nb (%) | COMIRNATY Na=1,860 nb (%) | Placebo Na=1,833 nb (%) | ||||||||||
| Severed | 0 (0.0) | 1 (0.1) | 21 (1.1) | 0 (0.0) | |||||||||
| Vomiting | |||||||||||||
| Any | 10 (0.5) | 9 (0.5) | 13 (0.7) | 5 (0.3) | |||||||||
| Severee | 0 (0.0) | 0 (0.0) | 2 (0.1) | 0 (0.0) | |||||||||
| Diarrhea | |||||||||||||
| Any | 168 (8.4) | 130 (6.5) | 152 (8.2) | 102 (5.6) | |||||||||
| Severef | 4 (0.2) | 1 (0.1) | 2 (0.1) | 4 (0.2) | |||||||||
| New or worsened muscle pain | |||||||||||||
| Any | 274 (13.6) | 165 (8.3) | 537 (28.9) | 99 (5.4) | |||||||||
| Severed | 1 (0.0) | 3 (0.2) | 20 (1.1) | 1 (0.1) | |||||||||
| New or worsened joint pain | |||||||||||||
| Any | 175 (8.7) | 124 (6.2) | 353 (19.0) | 72 (3.9) | |||||||||
| Severed | 3 (0.1) | 1 (0.1) | 9 (0.5) | 1 (0.1) | |||||||||
| Any systemic reactionc | 984 (49.0) | 749 (37.7) | 1,203 (64.7) | 516 (28.2) | |||||||||
| Use of antipyretic or pain medication | 382 (19.0) | 224 (11.3) | 688 (37.0) | 170 (9.3) | |||||||||
*Randomized participants in the safety analysis population who received at least 1 dose of the study intervention.
a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. The only Grade 4 solicited systemic reaction reported in participants 56 years of age and older was fatigue.
b. n = Number of participants with the specified reaction.
c. Any systemic reaction: any fever ≥38.0°C, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new or worsened joint pain.
d. Severe: prevents daily activity.
e. Severe: requires intravenous hydration.
f. Severe: 6 or more loose stools in 24 hours.
g. Grade 4: emergency room visit or hospitalization.
Study 2 also included 200 participants with confirmed stable human immunodeficiency virus
(HIV) infection. The safety profile of the participants with stable HIV infection receiving COMIRNATY (n = 100) was similar to that seen in the general population.
Unsolicited Adverse Events
The participants were unblinded to offer placebo participants COMIRNATY when they became locally eligible under regulatory approval in December 2020. A total of 25,651 (58.2%) participants (13,031 in vaccine group and 12,620 in placebo group) 16 years of age and older had been followed up for at least 4 months, with 3,082 (7.0%) participants (1,778 in vaccine group and 1,304 in placebo group) followed up for at least 6 months after the second dose during the blinded placebo-controlled follow-up period in Study 2. Adverse events detailed below for participants 16 years of age and older are for the placebo-controlled blinded follow-up period up to the participants’ unblinding dates.
Among participants 16 through 55 years of age who received at least one dose of study vaccine, 12,995 of whom received COMIRNATY and 13,026 of whom received placebo, unsolicited adverse events were reported by 4,396 (33.8%) participants in the COMIRNATY group and 2,136 (16.4%) participants in the placebo group. In a similar analysis in participants 56 years of age and older that included 8,931 COMIRNATY recipients and 8,895 placebo recipients, unsolicited adverse events were reported by 2,551 (28.6%) participants in the COMIRNATY group and 1,432 (16.1%) participants in the placebo group. Among participants with confirmed stable HIV infection that included 100 COMIRNATY recipients and 100 placebo recipients, unsolicited adverse events were reported by 29 (29%) participants in the COMIRNATY group and 15 (15%) participants in the placebo group.
Lymphadenopathy was reported in 87 (0.4%) participants in the vaccine group compared to 8 (<0.1%) participants in the placebo group. Bell’s palsy (facial paralysis and facial paresis) was reported by four participants in the vaccine group and two in the placebo group. In the four vaccinated participants, events began from 3 to 48 days after their last dose, were mild to moderate in severity, and duration ranged from 3 to 68 days. Currently available information is insufficient to determine a causal relationship with the vaccine. There were no other notable patterns or numerical imbalances between treatment groups for specific categories of non-serious adverse events (including other neurologic or neuro-inflammatory, and thrombotic events) that would suggest a causal relationship to COMIRNATY. Cumulative safety follow-up to at least 6 months after Dose 2 for approximately 12,000 participants who received COMIRNATY showed no other safety signals arising from longer-term follow-up of the study.
Serious Adverse Events
In Study 2, among participants 16 through 55 years of age who had received at least 1 dose of vaccine or placebo (COMIRNATY = 12,995; placebo = 13,026), serious adverse events from Dose 1 up to the participant unblinding date in ongoing follow-up were reported by 103 (0.8%) COMIRNATY recipients and 117 (0.9%) placebo recipients. In a similar analysis, in participants 56 years of age and older (COMIRNATY = 8,931; placebo = 8,895), serious adverse events were reported by 165 (1.8%) COMIRNATY recipients and 151 (1.7%) placebo recipients who received at least 1 dose of COMIRNATY or placebo, respectively. Among participants with confirmed stable HIV infection serious adverse events from Dose 1 up to the participant unblinding date in ongoing follow-up were reported by 2 (2%) COMIRNATY recipients and 2 (2%) placebo recipients.
Pericarditis was reported for one participant in the vaccine group, and no case was reported in the placebo group. Appendicitis was reported as a serious adverse event for 27 participants, 15 vaccine participants and 12 placebo participants. Currently available information is insufficient to determine a causal relationship with the vaccine. There were no other notable patterns or numerical imbalances between treatment groups for specific categories of serious adverse events (including neurologic, neuro-inflammatory, thrombotic events, myocarditis or anaphylactic reaction to the vaccine) reported during the blinded placebo-controlled follow-up period of the study.
Participants 16 Years of Age and Older – After Booster Dose
A subset from Study C4591001 (Study 2) Phase 2/3 participants, of 306 adults 18 through 55 years of age who completed the original COMIRNATY 2-dose series, received a booster dose of COMIRNATY approximately 6 months (range of 4.8 to 8.0 months) after receiving Dose 2.
In Study C4591031 (Study 4), a placebo-controlled booster study, participants 16 years of age and older recruited from Study C4591001 (Study 2) received a booster dose of COMIRNATY (5,081 participants), or placebo (5,044 participants) at least 6 months after the second dose of COMIRNATY. Overall, participants who received a booster dose, had a median follow-up time of 2.5 months after the booster dose to the cut-off date (5 October 2021). Among the participants, the median age was 53.0 years
(range 16 through 87 years of age), including 1,175 booster dose recipients (23.1%) who were ≥65 years of age, 49.1% were male and 50.9% were female, 79.0% were White, 14.9% were Hispanic/Latino, 9.2% were Black or African American, 5.5% were Asian, and 1.7% were American Indian/Alaska Native.
Solicited Local and Systemic Adverse Reactions
Overall, among participants who received a booster dose in a subset from Study C4591001 (Study 2), the median age was 42 years (range 19 through 55 years of age), 45.8% were male and 54.2% were female, 81.4% were White, 27.8% were Hispanic/Latino, 9.2% were Black or African American, 5.2% were Asian, and 0.7% were American Indian/Alaska Native.
| Table 15: Study 2 – Frequency and Percentages of Participants With Solicited Local Reactions, By Maximum Severity, Within 7 Days After the Booster Dose of COMIRNATY – Booster Dose Safety Population* | |||||
|---|---|---|---|---|---|
| Local Reaction | COMIRNATY Booster Dose Na = 289 nb (%) | ||||
| Rednessc | |||||
| Any (>2 cm) | 17 (5.9) | ||||
| Severe | 0 | ||||
| Swellingc | |||||
| Any (>2 cm) | 23 (8.0) | ||||
| Severe | 1 (0.3) | ||||
| Pain at the injection sited | |||||
| Any | 240 (83.0) | ||||
| Severe | 1 (0.3) | ||||
Note: Reactions were collected in the electronic diary (e-diary) from Day 1 to Day 7 after the booster dose. Note: No Grade 4 solicited local reactions were reported. *Participants in the safety analysis population who received the booster dose of COMIRNATY. a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. n = Number of participants with the specified reaction. c. Mild: >2.0 to 5.0 cm; Moderate: >5.0 to 10.0 cm; Severe: >10.0 cm. d. Mild: does not interfere with activity; Moderate: interferes with activity; Severe: prevents daily activity. | |||||
In participants who received a booster dose the mean duration of pain at the injection site after the booster dose was 2.6 days (range 1 to 8 days), for redness 2.2 days (range 1 to 15 days), and for swelling 2.2 days (range 1 to 8 days).
| Table 16: Study 2 – Frequency and Percentages of Participants With Solicited Systemic Reactions, by Maximum Severity, Within 7 Days After the Booster Dose of COMIRNATY – Booster Dose Safety Population* | |||||
|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Booster Dose Na = 289 nb (%) | ||||
| Fever | |||||
| ≥38.0℃ | 25 (8.7) | ||||
| ≥38.0℃ to 38.4℃ | 12 (4.2) | ||||
| >38.4℃ to 38.9℃ | 12 (4.2) | ||||
| >38.9℃ to 40.0℃ | 1 (0.3) | ||||
| >40.0℃ | 0 | ||||
| Fatiguec | |||||
| Any | 184 (63.7) | ||||
| Severe | 13 (4.5) | ||||
| Headachec | |||||
| Any | 140 (48.4) | ||||
| Severe | 3 (1.0) | ||||
| Chillsc | |||||
| Any | 84 (29.1) | ||||
| Severe | 3 (1.0) | ||||
| Vomitingd | |||||
| Any | 5 (1.7) | ||||
| Severe | 0 | ||||
| Diarrheae | |||||
| Any | 25 (8.7) | ||||
| Severe | 0 | ||||
| New or worsened muscle painc | |||||
| Any | 113 (39.1) | ||||
| Severe | 4 (1.4) | ||||
| New or worsened joint painc | |||||
| Any | 73 (25.3) | ||||
| Severe | 1 (0.3) | ||||
| Use of antipyretic or pain medicationf | 135 (46.7) | ||||
Note: Events and use of antipyretic or pain medication were collected in the electronic diary (e-diary) from Day 1 to Day 7 after the booster dose. Note: No Grade 4 solicited systemic reactions were reported. *Randomized participants in the safety analysis population who received the booster dose of COMIRNATY. a. N = Number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. n = Number of participants with the specified reaction. c. Mild: does not interfere with activity; Moderate: some interference with activity; Severe: prevents daily activity. d. Mild: 1 to 2 times in 24 hours; Moderate: >2 times in 24 hours; Severe: requires intravenous hydration. e. Mild: 2 to 3 loose stools in 24 hours; Moderate: 4 to 5 loose stools in 24 hours; Severe: 6 or more loose stools in 24 hours. f. Severity was not collected for use of antipyretic or pain medication. | |||||
Unsolicited Adverse Events
Overall, participants who received a booster dose in Study C4591031 (Study 4), had a median follow-up time of 2.5 months after the booster dose to the cut-off date (October 5, 2021).
In an analysis of all unsolicited adverse events reported following the booster dose of COMIRNATY, through 1 month after the booster dose, in participants 16 through 87 years of age (N = 5,055), adverse reactions included headache (5%), fever (4.8%), lymphadenopathy (2.8%), decreased appetite (0.2%), malaise (0.7%), nausea (0.9%), and pain in extremity (1.1%).
Serious Adverse Events
Of the participants who received a booster dose of COMIRNATY or placebo (COMIRNATY = 5,055; placebo = 5,020) to the cut-off date (October 5, 2021), serious adverse events were reported by 0.3% of COMIRNATY recipients and 0.5% by placebo recipients. There were no notable patterns or numerical imbalances between treatment groups for specific categories of serious adverse events that would suggest a causal relationship to COMIRNATY. A 17-year-old male in Study 2 was diagnosed with myocarditis three days after receiving the booster dose (Dose 3). The participant was treated and recovered.
Adolescents 12 to 15 Years of Age – Primary Series (Two Doses)
Solicited Adverse Reactions
Table 17 and Table 18 present the frequency and severity of solicited local and systemic reactions, respectively, within 7 days following each dose of COMIRNATY and placebo in adolescents 12 to 15 years of age included in the safety population who were monitored for reactogenicity with an electronic diary.
| Table 17: Study 2 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose of COMIRNATY – Adolescents 12 to 15 Years of Age – Safety Population* | |||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | COMIRNATY Dose 1 Na=1,127 nb (%) | Placebo Dose 1 Na=1,127 nb (%) | COMIRNATY Dose 2 Na=1,097 nb (%) | Placebo Dose 2 Na=1,078 nb (%) | |||||||
| Redness | |||||||||||
| Any (>2 cm) | 65 (5.8) | 12 (1.1) | 55 (5.0) | 10 (0.9) | |||||||
| Severec | 1 (0.1) | 0 (0.0) | 0 (0.0) | 0 (0.0) | |||||||
| Swelling | |||||||||||
| Any (>2 cm) | 78 (6.9) | 11 (1.0) | 54 (4.9) | 6 (0.6) | |||||||
| Severec | 0 (0.0) | 0 (0.0) | 0 (0.0) | 0 (0.0) | |||||||
| Pain at the injection site | |||||||||||
| Any | 971 (86.2) | 263 (23.3) | 866 (78.9) | 193 (17.9) | |||||||
| Severed | 11 (1.0) | 0 (0.0) | 7 (0.6) | 0 (0.0) | |||||||
| Any local reactione | 976 (86.6) | 271 (24.0) | 872 (79.5) | 198 (18.4) | |||||||
Note: Reactions were collected in the electronic diary (e-diary) from Day 1 to Day 7 after vaccination. a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: >10.0 cm. d. Severe: prevents daily activity. e. Any local reaction: any redness >2.0 cm, any swelling >2.0 cm, or any pain at the injection site * Randomized participants in the safety analysis population who received at least 1 dose of the study intervention. | |||||||||||
| Table 18: Study 2 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose of COMIRNATY – Adolescents 12 to 15 Years of Age – Safety Population* | |||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Dose 1 Na=1,127 nb (%) | Placebo Dose 1 Na=1,127 nb (%) | COMIRNATY Dose 2 Na=1,097 nb (%) | Placebo Dose 2 Na=1,078 nb (%) | |||||||
| Fever | |||||||||||
| ≥38.0°C | 114 (10.1) | 12 (1.1) | 215 (19.6) | 7 (0.6) | |||||||
| >38.9°C | 11 (1.0) | 2 (0.2) | 25 (2.3) | 1 (0.1) | |||||||
| Fatigue | |||||||||||
| Any | 677 (60.1) | 457 (40.6) | 726 (66.2) | 264 (24.5) | |||||||
| Severec | 15 (1.3) | 8 (0.7) | 26 (2.4) | 4 (0.4) | |||||||
| Headache | |||||||||||
| Any | 623 (55.3) | 396 (35.1) | 708 (64.5) | 263 (24.4) | |||||||
| Severec | 11 (1.0) | 9 (0.8) | 22 (2.0) | 1 (0.1) | |||||||
| Chills | |||||||||||
| Any | 311 (27.6) | 109 (9.7) | 455 (41.5) | 73 (6.8) | |||||||
| Severec | 5 (0.4) | 2 (0.2) | 20 (1.8) | 0 (0.0) | |||||||
| Vomiting | |||||||||||
| Any | 31 (2.8) | 10 (0.9) | 29 (2.6) | 12 (1.1) | |||||||
| Severed | 1 (0.1) | 0 (0.0) | 0 (0.0) | 0 (0.0) | |||||||
| Diarrhea | |||||||||||
| Any | 90 (8.0) | 82 (7.3) | 65 (5.9) | 43 (4.0) | |||||||
| Severee | 0 (0.0) | 0 (0.0) | 0 (0.0) | 0 (0.0) | |||||||
| New or worsened muscle pain | |||||||||||
| Any | 272 (24.1) | 148 (13.1) | 355 (32.4) | 90 (8.3) | |||||||
| Severec | 2 (0.2) | 0 (0.0) | 6 (0.5) | 2 (0.2) | |||||||
| New or worsened joint pain | |||||||||||
| Any | 109 (9.7) | 77 (6.8) | 173 (15.8) | 51 (4.7) | |||||||
| Severec | 1 (0.1) | 0 (0.0) | 4 (0.4) | 0 (0.0) | |||||||
| Any systemic reactionsf | 877 (77.8) | 636 (56.4) | 904 (82.4) | 439 (40.7) | |||||||
| Use of antipyretic or pain medication | 413 (36.6) | 111 (9.8) | 557 (50.8) | 95 (8.8) | |||||||
Note: Events and use of antipyretic or pain medication were collected in the electronic diary (e-diary) from Day 1 to Day 7 after each dose. a. N = Number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: prevents daily activity. d. Severe: requires intravenous hydration. e. Severe: 6 or more loose stools in 24 hours. f. Any systemic reaction: any fever ≥38.0°C, any fatigue, any vomiting, any chills, any diarrhea, any headache, any new or worsened muscle pain, or any new or worsened joint pain * Randomized participants in the safety analysis population who received at least 1 dose of the study intervention. | |||||||||||
Unsolicited Adverse Events
In the analysis of Study 2 of all unsolicited adverse events reported following any dose, through 1 month after Dose 2, in adolescents 12 to 15 years of age (N=2260; 1,131 COMIRNATY group vs. 1,129 placebo group), those assessed as adverse reactions not already captured by solicited local and systemic reactions were lymphadenopathy (9 (0.8%) vs. 2 (0.2%)), and nausea (5 (0.4%) vs. 1 (0.1%)).
In analyses of all unsolicited adverse events in Study 2 from Dose 1 up to the participant unblinding date, 69.0% of study participants 12 through 15 years of age had at least 4 months of follow-up after Dose 2. Among participants 12 through 15 years of age who received at least one dose of study vaccine, 1,131 of whom received COMIRNATY and 1,129 of whom received placebo, unsolicited adverse events were reported by 95 (8.4%) participants in the COMIRNATY group and 113 (10.0%) participants in the placebo group.
Non-serious adverse events from Dose 1 through up to 30 days after Dose 2 in ongoing follow-up were reported by 5.8% of COMIRNATY recipients and by 5.8% of placebo recipients. From Dose 1 through 30 days after Dose 2, reports of lymphadenopathy plausibly related to the study intervention were imbalanced, with notably more cases in the COMIRNATY group (7) vs. the placebo group (1). In the analysis of blinded, placebo-controlled follow-up, there were no other notable patterns or numerical imbalances between treatment groups for specific categories of non-serious adverse events (including other neurologic or neuro-inflammatory, and thrombotic events) that would suggest a causal relationship to COMIRNATY. In the analysis of unblinded follow-up, there were no notable patterns of specific categories of non-serious adverse events that would suggest a causal relationship to COMIRNATY.
Serious Adverse Events
In Study 2, among participants 12 through 15 years of age who had received at least 1 dose of vaccine or placebo (COMIRNATY = 1,131; placebo = 1,129), serious adverse events from Dose 1 up to the participant unblinding date in ongoing follow-up were reported by 10 (0.9%) COMIRNATY recipients and 2 (0.2%) placebo recipients. In these analyses, 69.0% (786 COMIRNATY and 773 placebo) of study participants had at least 4 months of follow-up after Dose 2. In the analysis of blinded, placebo-controlled follow-up, there were no notable patterns between treatment groups for specific categories of serious adverse events (including neurologic, neuro-inflammatory, and thrombotic events) that would suggest a causal relationship to COMIRNATY. In the analysis of unblinded follow-up, there were no notable patterns of specific categories of serious adverse events that would suggest a causal relationship to COMIRNATY. In study 2, a 16-year-old male was diagnosed with myopericarditis 3 days after his 2nd dose. The participant was treated and recovered.
8.2.5 COMIRNATY (10 mcg)
Children 5 Years Through <12 Years of Age – Primary Series (Two Doses)
Solicited Adverse Reactions
Table 19 and Table 20 present the frequency and severity of solicited local and systemic reactions, respectively, within 7 days following each dose of COMIRNATY and placebo in children 5 years through <12 years of age included in the initial enrolment safety population who were monitored for reactogenicity with an electronic diary.
Table 19: Study 3 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose – Children 5 Years Through <12 Years of Age – Safety Population*
| Local Reaction | COMIRNATY Dose 1 Na=1,511 nc (%) | Placebo Dose 1 Na,b=748 nc (%) | COMIRNATY Dose 2 Na=1,501 nc (%) | Placebo Dose 2 Na,b=740 nc (%) |
|---|---|---|---|---|
| Rednessd | ||||
| Any (≥0.5 cm) | 222 (14.7) | 43 (5.7) | 278 (18.5) | 40 (5.4) |
| Severe | 0 (0.0) | 0 (0.0) | 3 (0.2) | 0 (0.0) |
| Swellingd | ||||
| Any (≥0.5 cm) | 158 (10.5) | 20 (2.7) | 229 (15.3) | 20 (2.7) |
| Severe | 1 (0.1) | 0 (0.0) | 0 (0.0) | 0 (0.0) |
| Pain at the injection sitee | ||||
| Any | 1,119 (74.1) | 234 (31.3) | 1,065 (71.0) | 218 (29.5) |
| Severe | 4 (0.3) | 0 (0.0) | 5 (0.3) | 0 (0.0) |
Note: Reactions were collected in the electronic diary (e-diary) from Day 1 to Day 7 after vaccination. a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. The denominators (N) used in the percentage calculations for redness and swelling were 749 after Dose 1 and 741 after Dose 2 in the placebo group, due to an e-diary error. c. n = Number of participants with the specified reaction. d. Severe: >7.0 cm. e. Severe: prevents daily activity. * Randomized participants who received at least 1 dose of the study intervention. | ||||
Table 20: Study 3 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose – Children 5 Years Through <12 Years of Age – Safety Population*
| Systemic Reaction | COMIRNATY Dose 1 Na=1,511 nc (%) | Placebo Dose 1 Na,b=748 nc (%) | COMIRNATY Dose 2 Na=1,501 nc (%) | Placebo Dose 2 Na,b=740 nc (%) |
|---|---|---|---|---|
| Fever | ||||
| ≥38.0℃ | 38 (2.5) | 10 (1.3) | 98 (6.5) | 9 (1.2) |
| >38.9℃ | 3 (0.2) | 1 (0.1) | 9 (0.6) | 1 (0.1) |
| Fatigued | ||||
| Any | 508 (33.6) | 234 (31.3) | 592 (39.4) | 180 (24.3) |
| Severe | 4 (0.3) | 1 (0.1) | 11 (0.7) | 1 (0.1) |
| Headached | ||||
| Any | 339 (22.4) | 180 (24.1) | 420 (28.0) | 138 (18.6) |
| Severe | 2 (0.1) | 4 (0.5) | 3 (0.2) | 0 (0.0) |
| Chillsd | ||||
| Any | 70 (4.6) | 35 (4.7) | 147 (9.8) | 32 (4.3) |
| Severe | 0 (0.0) | 0 (0.0) | 2 (0.1) | 1 (0.1) |
| Vomitinge | ||||
| Any | 33 (2.2) | 11 (1.5) | 28 (1.9) | 6 (0.8) |
| Severe | 0 (0.0) | 0 (0.0) | 0 (0.0) | 0 (0.0) |
| Systemic Reaction | COMIRNATY Dose 1 Na=1,511 nc (%) | Placebo Dose 1 Na,b=748 nc (%) | COMIRNATY Dose 2 Na=1,501 nc (%) | Placebo Dose 2 Na,b=740 nc (%) |
|---|---|---|---|---|
| Diarrheaf | ||||
| Any | 89 (5.9) | 31 (4.1) | 79 (5.3) | 35 (4.7) |
| Severe | 0 (0.0) | 0 (0.0) | 0 (0.0) | 0 (0.0) |
| New or worsened muscle paind | ||||
| Any | 137 (9.1) | 51 (6.8) | 175 (11.7) | 55 (7.4) |
| Severe | 1 (0.1) | 0 (0.0) | 1 (0.1) | 0 (0.0) |
| New or worsened joint paind | ||||
| Any | 50 (3.3) | 41 (5.5) | 78 (5.2) | 27 (3.6) |
| Severe | 0 (0.0) | 0 (0.0) | 0 (0.0) | 0 (0.0) |
| Use of antipyretic or pain medicationg | 217 (14.4) | 62 (8.3) | 296 (19.7) | 60 (8.1) |
Note: Events and use of antipyretic or pain medication were collected in the electronic diary (e-diary) from Day 1 to Day 7 after each dose. a. N = Number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. The denominators (N) used in the percentage calculations for fever and use of antipyretic or pain medication were 749 after Dose 1 and 741 after Dose 2 in the placebo group, due to an e-diary error. c. n = Number of participants with the specified reaction. d. Severe: prevents daily activity. e. Severe: requires intravenous hydration. f. Severe: 6 or more loose stools in 24 hours. g. Severity was not collected for use of antipyretic or pain medication. * Randomized participants who received at least 1 dose of the study intervention. | ||||
Unsolicited Adverse Events
In the analyses of Study 3 in children 5 years through <12 years of age (initial enrolment group: 1,518 COMIRNATY 10 mcg and 750 placebo), 99.5% of participants had at least 30 days and 95.7% of participants had at least 3 months follow-up after Dose 2.
Serious adverse events from Dose 1 through up to 30 days after Dose 2 in ongoing follow-up in the initial enrolment group were reported by 1 participant (0.1%) in each group after receiving the vaccine or placebo through the data cut-off date. No serious adverse events were reported that were considered related to vaccination.
Non-serious adverse events from Dose 1 through up to 30 days after Dose 2 in ongoing follow-up in the initial enrolment group were reported by 10.9% of COMIRNATY 10 mcg recipients and by 9.1% of placebo recipients. Lymphadenopathy was reported in 13 (0.9%) participants in the COMIRNATY 10 mcg group vs. 1 (0.1%) in the placebo group. All cases were considered to be mild, with a median onset of 3 days after Dose 1, and 2 days after Dose 2 in the vaccine group. The median duration was 3.5 days (ranged from 1 to 14 days) in the vaccine group. Skin and subcutaneous tissue disorders (including skin rash, dermatitis, eczema and urticaria) were reported in 17 (1.1%) participants in the vaccine group and 5 (0.7%) participants in the placebo group. Most of the events began from 3-11 days after the second dose and were characterized as mild and self-limited. There were no other notable patterns between treatment groups for specific categories of non-serious adverse events that would suggest a causal relationship to COMIRNATY. There were no reports of myocarditis/pericarditis or anaphylaxis by the study cut-off date.
Children 5 Years Through ˂12 Years of Age – After Booster Dose
A subset of Phase 2/3 participants 5 years through ˂12 years of age received a booster dose of COMIRNATY at least 5 months after completing the primary series (range 5 to 9 months, 86.8% of participants received the booster dose at least 8 months after Dose 2). Those participants vaccinated prior to February 22, 2022 provided the safety database (n=401), and had a median safety follow-up of 1.3 months from vaccination through the data cut-off date of March 22, 2022.
The median age of these 401 participants was 8.0 years (range 5 years through ˂12 years of age), 52.4% were male and 47.6% were female, 70.1% were White, 7.2% were Black or African American, 22.9% were Hispanic/Latino, 7.7% were Asian, and 2.0% were American Indian/Alaska Native.
Solicited Local and Systemic Adverse Reactions
Table 21 and Table 22 present the frequency and severity of reported solicited local and systemic reactions, respectively, within 7 days of a booster dose of COMIRNATY for Phase 2/3 participants 5 years through ˂12 years of age.
In participants who received a booster dose, the mean duration of pain at the injection site after the booster dose was 2.4 days (range 1 to 35 days), for redness 2.3 days (range 1 to 12 days), and for swelling 2.3 days (range 1 to 9 days).
Table 21: Study 3 – Frequency and Percentages of Participants With Solicited Local Reactions, By Maximum Severity, Within 7 Days After the Booster Dose of COMIRNATY– Children 5 Years through ˂12 Years of Age – Safety Population*
| Local Reaction | COMIRNATY Booster Na=371 nb (%) |
|---|---|
| Rednessc | |
| Any (≥0.5 cm) | 58 (15.6) |
| Severe | 1 (0.3) |
| Swellingc | |
| Any (≥0.5 cm) | 61 (16.4) |
| Severe | 0 |
| Pain at the injection sited | |
| Any | 274 (73.9) |
| Severe | 2 (0.5) |
* Randomized participants who received at least 1 dose of the study intervention. Note: Reactions were collected in the e-diary and unscheduled clinical assessments from Day 1 through Day 7 after vaccination. a. N = number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. n = Number of participants with the specified characteristic. c. Mild: ≥0.5 to 2.0 cm; moderate: >2.0 to 7.0 cm; severe: >7.0 cm. d. Mild: does not interfere with activity; moderate: interferes with activity; severe: prevents daily activity. | |
| Table 22: Study 3 – Frequency and Percentages of Participants With Solicited Systemic Reactions, by Maximum Severity, Within 7 Days After the Booster Dose of COMIRNATY– Children 5 Years through ˂12 Years of Age – Safety Population* | |||||
|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY Booster Na=371 nb (%) | ||||
| Fever | |||||
| ≥38.0℃ | 25 (6.7) | ||||
| >38.9℃ | 3 (0.8) | ||||
| Fatiguec | |||||
| Any | 169 (45.6) | ||||
| Severe | 7 (1.9) | ||||
| Headachec | |||||
| Any | 126 (34.0) | ||||
| Severe | 0 | ||||
| Chillsc | |||||
| Any | 39 (10.5) | ||||
| Severe | 1 (0.3) | ||||
| Vomitingd | |||||
| Any | 9 (2.4) | ||||
| Severe | 0 | ||||
| Diarrheae | |||||
| Any | 18 (4.9) | ||||
| Severe | 1 (0.3) | ||||
| New or worsened muscle painc | |||||
| Any | 68 (18.3) | ||||
| Severe | 0 | ||||
| New or worsened joint painc | |||||
| Any | 25 (6.7) | ||||
| Severe | 0 | ||||
| Use of antipyretic or pain medicationf | 114 (30.7) | ||||
* Randomized participants who received at least 1 dose of the study intervention. Note: Events and use of antipyretic or pain medication were collected in the e-diary and unscheduled clinical assessments from Day 1 through Day 7 after vaccination. a. N = number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. n = Number of participants with the specified characteristic. c. Mild: does not interfere with activity; moderate: some interference with activity; severe: prevents daily activity. d. Mild: 1 to 2 times in 24 hours; moderate: >2 times in 24 hours; severe: requires intravenous hydration. e. Mild: 2 to 3 loose stools in 24 hours; moderate: 4 to 5 loose stools in 24 hours; severe: 6 or more loose stools in 24 hours. f. Severity was not collected for use of antipyretic or pain medication. | |||||
Unsolicited Adverse Events
Overall, the 401 participants who received a booster dose of COMIRNATY had a median follow-up time of 1.3 months after the booster dose through the cut-off date.
In an analysis of all unsolicited adverse events reported in participants 5 years through ˂12 years of age (N = 401) through up to 1 month after the booster dose, lymphadenopathy (n = 10, 2.5%) was an adverse reaction not already captured by solicited local and systemic reactions.
Serious Adverse Events
No serious adverse events were reported after the booster dose of COMIRNATY through the cut-off date.
8.2.6 COMIRNATY (3 mcg)
Children 2 Through <5 Years of Age – Primary Series (Three Doses)
Solicited Adverse Reactions
Table 23 and Table 24 present the frequency of solicited local and systemic reactions, respectively, within 7 days following each dose of COMIRNATY and placebo in children 2 through <5 years of age who were monitored for reactogenicity with an electronic diary.
| Table 23: Study 3 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose – Children 2 Through <5 Years of Age – Safety Population* | ||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | COMIRNATY† Dose 1 Na=1,814 to 1,825 nb (%) | Placebo Dose 1 Na=905 to 909 nb (%) | COMIRNATY† Dose 2 Na=1,772 to 1,779 nb (%) | Placebo Dose 2 Na=877 to 878 nb (%) | COMIRNATY† Dose 3 Na=547 to 552 nb (%) | Placebo Dose 3 Na=262 nb (%) | ||||||||||||||||
| Redness | ||||||||||||||||||||||
| Any (≥0.5 cm) | 160 (8.8) | 77 (8.5) | 202 (11.4) | 50 (5.7) | 60 (10.9) | 9 (3.4) | ||||||||||||||||
| Severec | 1 (0.1) | 1 (0.1) | 1 (0.1) | 0 | 0 | 0 | ||||||||||||||||
| Swelling | ||||||||||||||||||||||
| Any (≥0.5 cm) | 67 (3.7) | 26 (2.9) | 102 (5.7) | 18 (2.1) | 17 (3.1) | 3 (1.1) | ||||||||||||||||
| Severec | 0 | 0 | 0 | 0 | 0 | 0 | ||||||||||||||||
| Pain at the injection site | ||||||||||||||||||||||
| Any | 559 (30.8) | 186 (20.6) | 550 (31.0) | 178 (20.3) | 146 (26.7) | 35 (13.4) | ||||||||||||||||
| Severed | 0 | 1 (0.1) | 0 | 1 (0.1) | 0 | 0 | ||||||||||||||||
* Randomized participants who received at least 1 dose of the study intervention. † COMIRNATY 3 mcg. Note: Reactions were collected in an electronic diary (e-diary) from Day 1 to Day 7 after vaccination. a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: >7.0 cm. d. Severe: prevents daily activity. | ||||||||||||||||||||||
| Table 24: Study 3 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose – Children 2 Through <5 Years of Age – Safety Population* | |||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY† Dose 1 Na=1,813 to 1,824 nb (%) | Placebo Dose 1 Na=905 to 909 nb (%) | COMIRNATY† Dose 2 Na=1,772 to 1,779 nb (%) | Placebo Dose 2 Na=877 to 878 nb (%) | COMIRNATY† Dose 3 Na=547 to 552 nb (%) | Placebo Dose 3 Na=262 nb (%) | |||||||||||||||||
| Fever | |||||||||||||||||||||||
| ≥38.0℃ | 95 (5.2) | 48 (5.3) | 88 (4.9) | 46 (5.2) | 28 (5.1) | 11 (4.2) | |||||||||||||||||
| >38.9℃ | 14 (0.8) | 8 (0.9) | 21 (1.2) | 8 (0.9) | 4 (0.7) | 3 (1.1) | |||||||||||||||||
| Fatigue | |||||||||||||||||||||||
| Any | 539 (29.7) | 277 (30.6) | 456 (25.7) | 201 (22.9) | 134 (24.5) | 57 (21.8) | |||||||||||||||||
| Severec | 6 (0.3) | 5 (0.6) | 8 (0.5) | 3 (0.3) | 2 (0.4) | 0 | |||||||||||||||||
| Headache | |||||||||||||||||||||||
| Any | 81 (4.5) | 44 (4.9) | 81 (4.6) | 36 (4.1) | 27 (4.9) | 11 (4.2) | |||||||||||||||||
| Severec | 0 | 1 (0.1) | 0 | 1 (0.1) | 0 | 0 | |||||||||||||||||
| Chills | |||||||||||||||||||||||
| Any | 41 (2.3) | 22 (2.4) | 53 (3.0) | 23 (2.6) | 18 (3.3) | 7 (2.7) | |||||||||||||||||
| Severec | 3 (0.2) | 0 | 0 | 0 | 1 (0.2) | 0 | |||||||||||||||||
| Vomiting | |||||||||||||||||||||||
| Any | 54 (3.0) | 24 (2.7) | 61 (3.4) | 29 (3.3) | 9 (1.6) | 10 (3.8) | |||||||||||||||||
| Severed | 0 | 0 | 0 | 0 | 0 | 0 | |||||||||||||||||
| Diarrhea | |||||||||||||||||||||||
| Any | 139 (7.7) | 72 (8.0) | 118 (6.7) | 64 (7.3) | 28 (5.1) | 13 (5.0) | |||||||||||||||||
| Severee | 0 | 0 | 1 (0.1) | 0 | 0 | 0 | |||||||||||||||||
| New or worsened muscle pain | |||||||||||||||||||||||
| Any | 43 (2.4) | 15 (1.7) | 46 (2.6) | 21 (2.4) | 11 (2.0) | 4 (1.5) | |||||||||||||||||
| Severec | 1 (0.1) | 0 | 0 | 0 | 0 | 0 | |||||||||||||||||
| New or worsened joint pain | |||||||||||||||||||||||
| Any | 14 (0.8) | 18 (2.0) | 24 (1.4) | 9 (1.0) | 7 (1.3) | 2 (0.8) | |||||||||||||||||
| Severec | 0 | 0 | 0 | 0 | 1 (0.2) | 0 | |||||||||||||||||
| Use of antipyretic or pain medicationf | 197 (10.8) | 83 (9.1) | 177 (9.9) | 74 (8.4) | 47 (8.5) | 18 (6.9) | |||||||||||||||||
* Randomized participants who received at least 1 dose of the study intervention. † COMIRNATY 3 mcg. Note: Events and use of antipyretic or pain medication were collected in an electronic diary (e-diary) from Day 1 to Day 7 after each dose. a. N = Number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: prevents daily activity. d. Severe: requires intravenous hydration. e. Severe: 6 or more loose stools in 24 hours. f. Severity was not collected for use of antipyretic or pain medication. | |||||||||||||||||||||||
Unsolicited Adverse Events
In the analyses of Study 3 in participants 2 through <5 years of age (606 COMIRNATY; 280 placebo), 76.6% of participants had at least 30 days of follow-up after Dose 3.
Serious adverse events from Dose 1 through 1 month after Dose 3, with an overall median of 1.4 months follow-up after Dose 3, were reported by 0.7% of COMIRNATY recipients and by 0.9% of placebo recipients. One serious adverse event of fever (maximum temperature 40.3°C) on Day 3 after Dose 2 in a 4-year-old was considered possibly related to vaccination.
Non-serious adverse events from Dose 1 through up to 30 days after Dose 3, in ongoing follow-up were reported by 18.5% of COMIRNATY recipients and by 18.5% of placebo recipients.
From Dose 1 through 30 days after Dose 3, lymphadenopathy was reported in 1 (0.1%) of COMIRNATY recipients vs. 0 (0.0%) of placebo recipients. There were no other notable patterns between treatment groups for specific categories of non-serious adverse events that would suggest a causal relationship to COMIRNATY.
Children 6 Months Through <2 Years of Age – Primary Series (Three Doses)
Solicited Adverse Reactions
Table 25 and Table 26 present the frequency of solicited local and systemic reactions, respectively, within 7 days following each dose of COMIRNATY and placebo in children 6 months through <2 years of age who were monitored for reactogenicity with an electronic diary.
| Table 25: Study 3 – Frequency of Solicited Local Reactions Within 7 Days After Each Dose – Children 6 Months Through <2 Years of Age – Safety Population* | |||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Local Reaction | COMIRNATY† Dose 1 Na=1,159 to 1,173 nb (%) | Placebo Dose 1 Na=591 to 595 nb (%) | COMIRNATY† Dose 2 Na=1,137 to 1,147 nb (%) | Placebo Dose 2 Na=590 to 591 nb (%) | COMIRNATY† Dose 3 Na=362 to 365 nb (%) | Placebo Dose 3 Na=170 nb (%) | |||||||||||||||||
| Redness | |||||||||||||||||||||||
| Any (≥0.5 cm) | 124 (10.6) | 44 (7.4) | 107 (9.3) | 39 (6.6) | 26 (7.1) | 9 (5.3) | |||||||||||||||||
| Severec | 0 | 0 | 0 | 0 | 1 (0.3) | 0 | |||||||||||||||||
| Swelling | |||||||||||||||||||||||
| Any (≥0.5 cm) | 46 (3.9) | 15 (2.5) | 45 (3.9) | 9 (1.5) | 10 (2.7) | 3 (1.8) | |||||||||||||||||
| Severec | 0 | 0 | 0 | 0 | 0 | 0 | |||||||||||||||||
| Tenderness at the injection site | |||||||||||||||||||||||
| Any | 192 (16.6) | 66 (11.2) | 171 (15.0) | 50 (8.5) | 58 (16.0) | 20 (11.8) | |||||||||||||||||
| Severed | 0 | 0 | 1 (0.1) | 0 | 0 | 0 | |||||||||||||||||
* Randomized participants who received at least 1 dose of the study intervention. † COMIRNATY 3 mcg. Note: Reactions were collected in an electronic diary (e-diary) from Day 1 to Day 7 after vaccination. a. N = Number of participants reporting at least 1 yes or no response for the specified reaction after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: >7.0 cm. d. Severe: causes limitation of limb movement. | |||||||||||||||||||||||
| Table 26: Study 3 – Frequency of Solicited Systemic Reactions Within 7 Days After Each Dose – Children 6 Months Through <2 Years of Age – Safety Population* | |||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Systemic Reaction | COMIRNATY† Dose 1 Na=1,159 to 1,173 nb (%) | Placebo Dose 1 Na=591 to 595 nb (%) | COMIRNATY† Dose 2 Na=1,137 to 1,147 nb (%) | Placebo Dose 2 Na=590 to 591 nb (%) | COMIRNATY† Dose 3 Na=362 to 365 nb (%) | Placebo Dose 3 Na=170 nb (%) | |||||||||||||
| Fever | |||||||||||||||||||
| ≥38.0℃ | 85 (7.2) | 43 (7.2) | 85 (7.4) | 36 (6.1) | 25 (6.8) | 10 (5.9) | |||||||||||||
| >38.9℃ to 40.0℃ | 20 (1.7) | 7 (1.2) | 24 (2.1) | 7 (1.2) | 6 (1.6) | 1 (0.6) | |||||||||||||
| Decreased appetite | |||||||||||||||||||
| Any | 257 (22.2) | 125 (21.2) | 252 (22.2) | 106 (18.0) | 73 (20.2) | 23 (13.5) | |||||||||||||
| Severec | 3 (0.3) | 1 (0.2) | 4 (0.4) | 1 (0.2) | 4 (1.1) | 0 | |||||||||||||
| Drowsiness | |||||||||||||||||||
| Any | 313 (27.0) | 173 (29.3) | 271 (23.8) | 125 (21.2) | 72 (19.9) | 22 (12.9) | |||||||||||||
| Severed | 2 (0.2) | 2 (0.3) | 4 (0.4) | 1 (0.2) | 1 (0.3) | 1 (0.6) | |||||||||||||
| Irritability | |||||||||||||||||||
| Any | 593 (51.2) | 279 (47.2) | 539 (47.4) | 240 (40.7) | 158 (43.6) | 64 (37.6) | |||||||||||||
| Severee | 7 (0.6) | 0 | 7 (0.6) | 5 (0.8) | 1 (0.3) | 0 | |||||||||||||
| Use of antipyretic or pain medicationf | 281 (24.0) | 117 (19.7) | 243 (21.2) | 111 (18.8) | 70 (19.2) | 28 (16.5) | |||||||||||||
* Randomized participants who received at least 1 dose of the study intervention. † COMIRNATY 3 mcg. Note: Events and use of antipyretic or pain medication were collected in an electronic diary (e-diary) from Day 1 to Day 7 after each dose. a. N = Number of participants reporting at least 1 yes or no response for the specified event after the specified dose. b. n = Number of participants with the specified reaction. c. Severe: refusal to feed. d. Severe: disabling; not interested in usual daily activity. e. Severe: inconsolable; crying cannot be comforted. f. Severity was not collected for use of antipyretic or pain medication. | |||||||||||||||||||
Unsolicited Adverse Events
In the analyses of Study 3 in participants 6 months through 2 years of age (386 COMIRNATY; 184 placebo), 83.7% of participants had at least 30 days of follow-up after Dose 3.
Serious adverse events from Dose 1 through 1 month after Dose 3, with an overall median of 1.3 months follow-up after Dose 3, were reported by 1.4% of COMIRNATY recipients and by 2.3% of placebo recipients. No serious adverse events were reported that were considered related to vaccination.
Non-serious adverse events from Dose 1 through up to 1 month after Dose 3, in ongoing follow-up were reported by 29.1% of COMIRNATY recipients and by 26.3% of placebo recipients.
From Dose 1 through 30 days after Dose 3, lymphadenopathy was reported in 2 (0.2%) of COMIRNATY recipients vs. 0 (0%) of placebo recipients. There were no other notable patterns between treatment groups for specific categories of non-serious adverse events that would suggest a causal relationship to COMIRNATY.
8.5 Post-Market Adverse Reactions
The following adverse reactions have been identified during post authorization use of COMIRNATY.
Cardiac disorders: myocarditis and/or pericarditis (see 7 WARNING AND PRECAUTIONS)
Immune system disorders: severe allergic reactions, including anaphylaxis
Musculoskeletal and connective tissue disorders: pain in extremity (arm)
Nervous system disorders: Facial paralysis / Bell’s Palsy, hypoesthesia, paresthesia, dizziness
Skin and subcutaneous tissue disorders and other hypersensitivity reactions: skin rash, pruritus, urticaria, angioedema, erythema multiforme
Because these reactions are reported voluntarily from a population of uncertain size, it is not always possible to reliably estimate their frequency or establish a causal relationship to product exposure. They are included because: a) they represent reactions that are known to occur following immunizations generally; b) they are potentially serious; or c) on the basis of their frequency of reporting.
9 DRUG INTERACTIONS
No interaction studies have been performed. There is no information on the co-administration of COMIRNATY Omicron XBB.1.5 with other vaccines.
Do not mix COMIRNATY Omicron XBB.1.5 with other vaccines/products in the same syringe.
10 CLINICAL PHARMACOLOGY
10.1 Mechanism of Action
The nucleoside-modified messenger RNA in raxtozinameran encodes for the viral spike (S) protein of SARS-CoV-2 Omicron variant lineage XBB.1.5. The mRNA is formulated in lipid nanoparticles, which enable delivery of the RNA into host cells to allow expression of the SARS-CoV-2 S antigen. The vaccine elicits both neutralizing antibody and cellular immune responses to the spike (S) antigen, which may contribute to protection against COVID-19 disease.
11 STORAGE, STABILITY AND DISPOSAL
During storage, minimize exposure to room light, and avoid exposure to direct sunlight and ultraviolet light.
Do not refreeze thawed vials.
For 12 Years and Older: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Gray Cap and Gray Label Border)
And
For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose or Multiple Dose Vials with Blue Cap and Blue Label Border)
Single Dose or Multiple Dose Vial Storage Prior to Use
Cartons of COMIRNATY Omicron XBB.1.5 single dose or multiple dose vials (for 12 years and older: DO NOT DILUTE) and single dose or multiple dose vials (for 5 years to <12 years): DO NOT DILUTE) may arrive frozen at ultra-cold conditions in thermal containers with dry ice.
Once received, frozen single dose or multiple dose vials may be immediately transferred to the refrigerator [2°C to 8°C (35°F to 46°F)], thawed and stored for a single period of up to 10 weeks within the 18-month shelf-life. The 10-week refrigerated expiry date should be recorded on the carton at the time of transfer.
A carton of 10 single dose vials may take up to 2 hours to thaw at this temperature.
A carton of 10 multiple dose vials may take up to 6 hours to thaw at this temperature.
Alternatively, frozen single dose or multiple dose vials may be stored in an ultra-low temperature freezer at -90°C to -60°C (-130°F to -76°F) within the expiry date printed on the single dose or multiple dose vials and cartons. Do not store single dose or multiple dose vials at -25°C to -15°C (-13°F to 5°F). Once single dose or multiple dose vials are thawed they should not be refrozen.
Cartons of COMIRNATY Omicron XBB.1.5 single dose or multiple dose vials (for 12 years and older: DO NOT DILUTE) and single dose or multiple dose vials (for 5 years to <12 years): DO NOT DILUTE) may also arrive at 2°C to 8°C (35°F to 46°F). If received at 2°C to 8°C, they should be stored at 2°C to 8°C. Check that the carton has been updated to reflect the 10-week refrigerated expiry date.
Regardless of storage condition, vaccine should be used within the expiry date printed on the single dose or multiple dose vials and cartons.
Single Dose or Multiple Dose Vial Storage During Use
If not previously thawed at 2°C to 8°C (35°F to 46°F), allow single dose or multiple dose vials to thaw at room temperature [up to 25°C (77°F)] for 30 minutes.
COMIRNATY Omicron XBB.1.5 single dose or multiple dose vials (for 12 years and older: DO NOT DILUTE) and single dose or multiple dose vials (for 5 years to <12 years): DO NOT DILUTE) may be stored at room temperature up to 25°C (77°F) for a total of 12 hours prior to the first puncture.
DO NOT DILUTE PRIOR TO USE.
After first puncture, the single dose or multiple dose vial should be stored at 2°C to 25°C (35°F to 77°F). Vials should be discarded 12 hours after first puncture.
Thawed single dose or multiple dose vials can be handled in room light conditions.
Transportation of Single Dose or Multiple Dose Vials
If local redistribution is needed, full cartons containing unpunctured single dose or multiple dose vials may be transported at -90°C to -60°C (-130°F to -76°F); full cartons or individual unpunctured single dose or multiple dose vials may also be transported at 2°C to 8°C (35°F to 46°F).
For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border)
And
For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border)
Multiple Dose Vial Storage Prior to Use
Cartons of COMIRNATY Omicron XBB.1.5 multiple dose vials (for age 5 years to <12 years) and COMIRNATY Omicron XBB.1.5 multiple dose vials (for age 6 months to <5 years) may arrive frozen at ultra-cold conditions in thermal containers with dry ice.
Once received, frozen multiple dose vials may be immediately transferred to the refrigerator [2°C to 8°C (35°F to 46°F)], thawed and stored for a single period of up to 10 weeks within the 18-month shelf-life. The 10-week refrigerated expiry date should be recorded on the carton at the time of transfer.
- COMIRNATY Omicron XBB.1.5 (for age 5 years to <12 years): A carton of 10 multiple dose vials may take up to 4 hours to thaw at this temperature.
- COMIRNATY Omicron XBB.1.5 (for age 6 months to <5 years): A carton of 10 multiple dose vials may take up to 2 hours to thaw at this temperature.
Alternatively, frozen multiple dose vials may be stored in an ultra-low temperature freezer at -90°C to -60°C (-130°F to -76°F) within the expiry date printed on the single dose or multiple dose vials and cartons. Do not store multiple dose vials at -25°C to -15°C (-13°F to 5°F). Once multiple dose vials are thawed they should not be refrozen.
Cartons of COMIRNATY Omicron XBB.1.5 (for age 5 years to <12 years) and COMIRNATY Omicron XBB.1.5 (for age 6 months to <5 years) may also arrive at 2°C to 8°C (35°F to 46°F). If multiple dose vials are received at 2°C to 8°C, they should be stored at 2°C to 8°C. Check that the carton has been updated to reflect the 10-week refrigerated expiry date.
Regardless of storage condition, vaccines should be used within the expiry date printed on the vial and cartons.
Multiple Dose Vial Storage During Use
If not previously thawed at 2°C to 8°C (35°F to 46°F), allow multiple dose vials to thaw at room temperature [up to 25°C (77°F)] for 30 minutes.
Multiple dose vials of COMIRNATY Omicron XBB.1.5 (for age 5 years to <12 years) and COMIRNATY Omicron XBB.1.5 (for age 6 months to <5 years) may be stored at temperatures up to 25°C (77°F) for a total of 12 hours prior to dilution.
After dilution the multiple dose vials should be stored at 2°C to 25°C (35°F to 77°F). Vials should be discarded 12 hours after dilution (i.e., the first puncture).
Thawed vials can be handled in room light conditions.
Transportation of Multiple Dose Vials
If local redistribution is needed, full cartons containing undiluted multiple dose vials may be transported at -90°C to -60°C (-130°F to -76°F); full cartons or individual undiluted multiple dose vials may also be transported at 2°C to 8°C (35°F to 46°F).
12 SPECIAL HANDLING INSTRUCTIONS
COMIRNATY Omicron XBB.1.5 single dose and multiple dose vials contain a frozen suspension that does not contain preservative and must be thawed and may require dilution prior to administration.
Careful attention should be paid to the vial cap colour and label border and information on the label, and the appropriate corresponding instructions must be followed. For important information on handling and preparation for administration, please refer to 11 STORAGE, STABILITY AND DISPOSAL and 4 DOSAGE AND ADMINISTRATION.
PART II: SCIENTIFIC INFORMATION
13 PHARMACEUTICAL INFORMATION
Drug Substance:
Proper name: COVID-19 Vaccine, mRNA
International nonproprietary name: Raxtozinameran (Omicron XBB.1.5 strain)
Product Characteristics:
COMIRNATY Omicron XBB.1.5 (COVID-19 mRNA Vaccine, Monovalent (Omicron XBB.1.5)) contains highly purified single-stranded, 5’-capped messenger RNA (mRNA) produced using a cell-free in vitro transcription from the corresponding DNA template encoding the viral spike (S) protein of the SARS-CoV-2 Omicron variant lineage XBB.1.5 (Omicron XBB.1.5).
This vaccine is a white to off-white frozen suspension.
For 12 Years and Older: DO NOT DILUTE (Single Dose Vials with Gray Cap and Gray Label Border)
One single dose vial contains 1 dose of 0.3 mL. Do not dilute prior to use. One dose (0.3 mL) contains 30 micrograms of COVID-19 mRNA (30 mcg Omicron XBB.1.5), embedded in lipid nanoparticles.
For 12 Years and Older: DO NOT DILUTE (Multiple Dose vials with Gray Cap and Gray Label Border)
One multiple dose vial (2.25 mL) contains 6† doses of 0.3 mL. Do not dilute prior to use. One dose (0.3 mL) contains 30 micrograms of COVID-19 mRNA (30 mcg Omicron XBB.1.5), embedded in lipid nanoparticles.
For Age 5 Years to <12 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Orange Cap and Orange Label Border)
One multiple dose vial (1.3 mL) contains 10* doses of 0.2 mL after dilution. One dose (0.2 mL) contains 10 micrograms of COVID-19 mRNA Vaccine (10 mcg Omicron XBB.1.5), embedded in lipid nanoparticles.
For Age 5 Years to <12 Years: DO NOT DILUTE (Single Dose Vials with Blue Cap and Blue Label Border)
One single dose vial contains 1 dose of 0.3 mL. Do not dilute prior to use. One dose (0.3 mL) contains 10 micrograms of COVID-19 mRNA (10 mcg Omicron XBB.1.5), embedded in lipid nanoparticles.
For Age 5 Years to <12 Years: DO NOT DILUTE (Multiple Dose Vials with Blue Cap and Blue Label Border)
One multiple dose vial (2.25 mL) contains 6† doses of 0.3 mL. Do not dilute prior to use. One dose (0.3 mL) contains 10 micrograms of COVID-19 mRNA (10 mcg Omicron XBB.1.5), embedded in lipid nanoparticles.
For Age 6 Months to <5 Years: DILUTE PRIOR TO USE (Multiple Dose Vials with Maroon Cap and Maroon Label Border)
One multiple dose vial (0.4 mL) contains 10* doses of 0.2 mL after dilution. One dose (0.2 mL) contains 3 micrograms of COVID-19 mRNA Vaccine (3 mcg Omicron XXB.1.5), embedded in lipid nanoparticles.
† Low dead volume syringes and/or needles can be used to extract 6 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract a 6th dose from a single vial.
* Low dead-volume syringes and/or needles can be used to extract 10 doses from a single vial. If standard syringes and needles are used, there may not be sufficient volume to extract 10 doses from a single vial.
14 CLINICAL TRIALS
14.1 Trial Design and Study Demographics
The safety and effectiveness of COMIRNATY Omicron XBB.1.5 for individuals 6 months of age and older are inferred from studies which evaluated the primary series and booster vaccination with COMIRNATY and supported by studies which evaluated a booster dose of COMIRNATY Original & Omicron BA.4/BA.5 in individuals 6 months of age and older.
14.1.1 COMIRNATY Original & Omicron BA.4/BA.5 (15/15mcg)
Relative vaccine immunogenicity in participants greater than 12 years of age – after a second booster dose of COMIRNATY bivalent vaccine
Study C4591044 (Study 5) is an ongoing Phase 2/3 study to evaluate the safety, tolerability, and immunogenicity of new bivalent vaccines including COMIRNATY Original & Omicron BA.4/BA.5. A subset of 107 Study 5 Phase 2/3 participants 12 through 17 years of age, 313 participants 18 through 55 years of age and 306 participants 56 years of age and older previously vaccinated with a 2-dose primary series and 1 booster dose of COMIRNATY (original vaccine), went on to receive a second booster dose with COMIRNATY Original & Omicron BA.4/BA.5 (15/15 mcg, bivalent vaccine). Participants received a second booster dose 11.1 months (median time; range 5.4 to 16.9 months) after receiving the first booster dose and had a median follow up time of 1.5 months up to a data cut-off date of 31 October 2022. The median age was 48.0 years, 42.7% were male, 57.3% were female, 80.6% were White, 11.4% were Hispanic/Latino, 5.9% were Asian, and 11.4% were Black or African American.
14.1.2 COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg)
Relative vaccine Immunogenicity in children 6 months through ˂5 years of age – after the booster (fourth dose)
Study C4591048 (Study 6) is a Phase 1/2/3 master study investigating the safety, tolerability, and immunogenicity of COMIRNATY Original & Omicron BA.4/BA.5 bivalent vaccine. In Study 6, a subset of 60 participants 6 months through <5 years of age received a booster dose (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 (1.5 mcg/1.5 mcg) after receiving 3 prior doses of COMIRNATY (3 mcg), with their last dose 60 to 240 days prior to enrollment. The evaluable immunogenicity population (with or without evidence of infection up to 1month post-Dose 4) included 58 participants ≥6 months through <5 years of age (23 were ≥6 months through <2 years of age and 35 were 2 through <5 years of age). A total of 50.0% of participants were male. Most participants were White (58.6%), with 5.2% Black or African American participants, 15.5% Asian participants, and 20.7% multiracial participants. There were 25.9% Hispanic/Latino participants. Median age at the fourth dose was 19.0 months for the ≥6 months through <2 years of age group and 2.0 years for the 2 through <5 years of age group. Overall, 8.6% of participants reported comorbidities. A total of 27.6% of participants had evidence of prior SARS-CoV-2 infection at the time of Dose 4 (“baseline positive”).
14.1.3 COMIRNATY (30 mcg)
The safety and efficacy of COMIRNATY were evaluated in Study 2, a multicenter, multinational, Phase 1/2/3, randomized, placebo-controlled, observer-blind, dose-finding, vaccine candidate-selection and efficacy study in participants 12 years of age and older. Randomization was stratified by age: 12 through 15 years of age, 16 through 55 years of age, or 56 years of age and older, with a minimum of 40% of participants in the ≥56 year stratum. The study excluded participants who were
immunocompromised and those who had previous clinical or microbiological diagnosis of COVID-19. Participants with pre-existing stable disease, defined as disease not requiring significant change in therapy or hospitalization for worsening disease during the 6 weeks before enrollment, were included as were participants with known stable infection with human immunodeficiency virus (HIV), hepatitis C virus (HCV), or hepatitis B virus (HBV).
In the Phase 2/3 portion of Study 2, based on data accrued through November 14, 2020, approximately 44,000 participants 12 years of age and older were randomized equally and received 2 doses of COMIRNATY or placebo. The efficacy analyses included participants that received their second vaccination within 19 to 42 days after their first vaccination. The majority (93.1%) of vaccine recipients received the second dose 19 days to 23 days after Dose 1. Participants are planned to be followed for up to 24 months, for assessments of safety and efficacy against COVID-19.
The population for the analysis of the primary efficacy endpoint included 36,621 participants 12 years of age and older (18,242 in the COMIRNATY group and 18,379 in the placebo group) who did not have evidence of prior infection with SARS-CoV-2 through 7 days after the second dose. Table 27 presents the specific demographic characteristics in the studied population.
| Table 27: Demographics (Population for the Primary Efficacy Endpoint)a (Data Accrued Through November 14, 2020) | |||||||||
|---|---|---|---|---|---|---|---|---|---|
| COMIRNATY (N=18,242) n (%) | Placebo (N=18,379) n (%) | ||||||||
| Sex | |||||||||
| Male | 9,318 (51.1) | 9,225 (50.2) | |||||||
| Female | 8,924 (48.9) | 9,154 (49.8) | |||||||
| Age (years) | |||||||||
| Mean (SD) | 50.6 (15.70) | 50.4 (15.81) | |||||||
| Median | 52.0 | 52.0 | |||||||
| Min, max | (12, 89) | (12, 91) | |||||||
| Age group | |||||||||
| 12 to 15 years | 46 (0.3) | 42 (0.2) | |||||||
| 16 to 64 years | 14,216 (77.9) | 14,299 (77.8) | |||||||
| 65 to 74 years | 3,176 (17.4) | 3,226 (17.6) | |||||||
| ≥75 years | 804 (4.4) | 812 (4.4) | |||||||
| Race | |||||||||
| White | 15,110 (82.8) | 15,301 (83.3) | |||||||
| Black or African American | 1,617 (8.9) | 1,617 (8.8) | |||||||
| American Indian or Alaska Native | 118 (0.6) | 106 (0.6) | |||||||
| Asian | 815 (4.5) | 810 (4.4) | |||||||
| Native Hawaiian or other Pacific Islander | 48 (0.3) | 29 (0.2) | |||||||
| Otherb | 534 (2.9) | 516 (2.8) | |||||||
| Ethnicity | |||||||||
| Hispanic or Latino | 4,886 (26.8) | 4,857 (26.4) | |||||||
| Not Hispanic or Latino | 13,253 (72.7) | 13,412 (73.0) | |||||||
| Not reported | 103 (0.6) | 110 (0.6) | |||||||
| Comorbiditiesc | |||||||||
| Table 27: Demographics (Population for the Primary Efficacy Endpoint)a (Data Accrued Through November 14, 2020) | |||
|---|---|---|---|
| COMIRNATY (N=18,242) n (%) | Placebo (N=18,379) n (%) | ||
| Yes | 8,432 (46.2) | 8,450 (46.0) | |
| No | 9,810 (53.8) | 9,929 (54.0) | |
a. All eligible randomized participants who receive all vaccination(s) as randomized within the predefined window, have no other important protocol deviations as determined by the clinician, and have no evidence of SARS-CoV-2 infection prior to 7 days after Dose 2. b. Includes multiracial and not reported. c. Number of participants who have 1 or more comorbidities that increase the risk of severe COVID-19 disease.
| |||
To assess boostability, a subset of Study 2 participants were enrolled in selected sites, and 306 participants aged 18 to 55 years were re-randomized to receive a booster dose approximately 6 months after completion of the two-dose regimen (median interval between dose 2 and booster dose – 6.8 months; range 4.8 to 8.0 months). The median age at the time of booster vaccination was 42.0 years, and 46.3% of participants were male.
In Study 4, a placebo-controlled booster study, 5,081 participants 16 years of age and older were recruited from Study 2 to receive a booster dose of COMIRNATY at least 6 months after the second dose. The median age at the time of booster vaccination was 53 years, and 49% of the participants were male.
14.1.4 COMIRNATY (10 mcg)
Participants 5 Through <12 Years of Age
Study 3 is a Phase 1/2/3 study comprised of an open-label vaccine dose finding portion (Phase 1) and a multicentre, multinational, randomized, saline placebo-controlled, observer-blind immunogenicity and efficacy portion (Phase 2/3) that has enrolled participants 6 months to <12 years of age.
Participants 5 Through <12 Years of Age: Demographic characteristics in Study 3 were generally similar with regard to age, gender, race, and ethnicity among participants 5 years through <12 years of age who received COMIRNATY 10 mcg and those who received placebo. Among the 1,518 participants (initial enrolment group) 5 years through <12 years of age who received at least 1 dose of COMIRNATY 10 mcg, 52.6% were male and 47.4% were female, 79.3% were White, 5.9% were Black or African American, 21.0% were Hispanic/Latino, 5.9% were Asian, and 0.8% were American Indian/Alaska Native.
A descriptive efficacy analysis of Study 3 has been performed in 1,968 children 5 years through ˂12 years of age without evidence of infection prior to 7 days after Dose 2. This analysis evaluated confirmed symptomatic COVID-19 cases accrued up to a data cut-off date of October 8, 2021.
Table 28 presents the specific demographic characteristics in participants who did not have evidence of prior infection with SARS-CoV-2 through 7 days after the second dose.
Table 28: Demographics Characteristics – Participants Without Evidence of Infection Prior to 7 Days After Dose 2 – Phase 2/3 – 5 Years Through ˂12 Years of Age – Evaluable Efficacy Population (Data Accrued Through October 8, 2021)
COMIRNATY 10 mcg/dose (Na=1,305) nb (%) | Placebo (Na=663) nb (%) | |||||||
|---|---|---|---|---|---|---|---|---|
| Sex | ||||||||
| Male | 679 (52.0) | 343 (51.7) | ||||||
| Female | 626 (48.0) | 320 (48.3) | ||||||
| Age at Vaccination | ||||||||
| Mean (SD) | 8.2 (1.93) | 8.1 (1.98) | ||||||
| Median | 8.0 | 8.0 | ||||||
| Min, max | (5, 11) | (5, 11) | ||||||
| Race | ||||||||
| White | 1,018 (78.0) | 514 (77.5) | ||||||
| Black or African American | 76 (5.8) | 48 (7.2) | ||||||
| American Indian or Alaska Native | <1.0% | <1.0% | ||||||
| Asian | 86 (6.6) | 46 (6.9) | ||||||
| Native Hawaiian or other Pacific Islander | <1.0% | <1.0% | ||||||
| Otherc | 110 (8.4) | 52 (7.8) | ||||||
| Ethnicity | ||||||||
| Hispanic or Latino | 243 (18.6) | 130 (19.6) | ||||||
| Not Hispanic or Latino | 1059 (81.1) | 533 (80.4) | ||||||
| Not reported | <1.0% | <1.0% | ||||||
| Comorbiditiesd | ||||||||
| Yes | 262 (20.1) | 133 (20.1) | ||||||
| No | 1043 (79.9) | 530 (79.9) | ||||||
a. N = number of participants in the specified group from the evaluable efficacy population with no evidence of SARS CoV-2 infection prior to 7 days after Dose 2. This value is the denominator for the percentage calculations. Evaluable efficacy population included all eligible randomized participants who received all vaccination(s) as randomized within the predefined window, had no other important protocol deviations as determined by the clinician. b. n = Number of participants with the specified characteristic. c. Includes multiracial and not reported. d. Number of participants who have 1 or more comorbidities that increase the risk of severe COVID-19 disease: defined as participants who had at least 1 of the prespecified comorbidities based on MMWR 69(32);1081-1088 and/or obesity (BMI ≥95th percentile). | ||||||||
14.2 Study Results
14.2.1 COMIRNATY Original & Omicron BA.4/BA.5 (15/15mcg)
14.2.1.1 Immunogenicity in Participants 12 Years of Age and Older – After Second Booster Dose (Fourth Dose)
In an analysis of a subset from Study 5, 105 participants 12 to 17 years of age, 297 participants 18 to 55 years of age, and 286 participants 56 years of age and older who had previously received a 2-dose primary series and 1 booster dose with COMIRNATY (original vaccine) received a second booster (fourth) dose of COMIRNATY Original & Omicron BA.4/BA.5 (15/15 mcg, bivalent vaccine). In participants 12 to 17 years of age, 18 to 55 years of age, and 56 years of age and older, 75.2%, 71.7% and 61.5% were positive for SARS-CoV-2 at baseline, respectively.
Analyses of 50% neutralizing antibody titers (NT50) against Omicron BA.4/BA.5 and against reference strain among participants 56 years of age and older who received a second booster dose of COMIRNATY Original & Omicron BA.4/5 in Study 5 compared to a subset of participants from Study 4 who received a second booster dose of COMIRNATY demonstrated superiority of COMIRNATY Original & Omicron BA.4/BA.5 to COMIRNATY based on geometric mean ratio (GMR) and noninferiority based on difference in seroresponse rates with respect to anti-Omicron BA.4/BA.5 response, and noninferiority of anti-reference strain immune response based on GMR (Table 29 and Table 30).
Analyses of NT50 against Omicron BA.4/BA.5 among participants 18 through 55 years of age compared to participants 56 years of age and older who received a second booster dose of COMIRNATY Original & Omicron BA.4/BA.5 in Study 5 demonstrated noninferiority of anti-Omicron BA.4/BA.5 response among participants 18 through 55 years of age compared to participants 56 years of age and older for both GMR and difference in seroresponse rates (Table 29 and Table 30).
The study also assessed the level of NT50 of the anti-Omicron BA.4/BA.5 SARS-COV-2 strains pre-vaccination and 1 month after vaccination in participants who received a second booster dose (Table 31).
| Table 29: Geometric Mean Ratios – Study 5 – Participants With or Without Evidence of Infection – Evaluable Immunogenicity Population | |||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| SARS-CoV-2 Neutralization Assay | Sampling Time Pointa | COMIRNATY Original & Omicron BA.4/BA.5 Study 5 | COMIRNATY Subset of Study 4 | Age Group Comparison | Vaccine Group Comparison | ||||||||||||||
| 18 Through 55 Years of Age | 56 Years of Age and Older | 56 Years of Age and Older | COMIRNATY Original & Omicron BA.4/BA.5 18 Through 55 Years of Age/≥ 56 Years of Age | ≥ 56 Years of age COMIRNATY Original & Omicron BA.4/BA.5 /COMIRNATY | |||||||||||||||
| nb | GMTc (95% CIc) | nb | GMTc (95% CIc) | nb | GMTc (95% CIc) | GMRd (95% CId) | GMRd (95% CId) | ||||||||||||
| Omicron BA.4/BA.5 – NT50 (titer)e | 1 month | 297 | 4455.9 (3851.7, 5154.8) | 284 | 4158.1 (3554.8, 4863.8) | 282 | 938.9 (802.3, 1098.8) | 0.98 (0.83, 1.16)f | 2.91 (2.45, 3.44)g | ||||||||||
Reference Strain – NT50 (titer)e | 1 month | - | - | 286 | 16250.1 (14499.2, 18212.4) | 289 | 10415.5 (9366.7, 11581.8) | - | 1.38 (1.22, 1.56)h | ||||||||||
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. a. Protocol-specified timing for blood sample collection. b. n = Number of participants with valid and determinate assay results for the specified assay at the given sampling time point. c. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. d. GMRs and 2-sided 95% Cis were calculated by exponentiating the difference of LS means and corresponding Cis based on analysis of logarithmically transformed neutralizing titers using a linear regression model with terms of baseline neutralizing titer (log scale) and vaccine group or age group. e. SARS-CoV-2 NT50 were determined using a validated 384-well assay platform (original strain [USA-WA1/2020, isolated in January 2020] and Omicron B.1.1.529 subvariant BA.4/BA.5). f. Noninferiority is declared if the lower bound of the 2-sided 95% CI for the GMR is greater than 0.67. g. Superiority is declared if the lower bound of the 2-sided 95% CI for the GMR is greater than 1. h. Noninferiority is declared if the lower bound of the 2-sided 95% CI for the GMR is greater than 0.67 and the point estimate of the GMR is ≥ 0.8. | |||||||||||||||||||
| Table 30: Difference in Percentages of Participants with Seroresponse – COMIRNATY Original & Omicron BA.4/BA.5 from Study 5 and COMIRNATY from Subset of Study 4 – Participants With or Without Evidence of Infection – Evaluable Immunogenicity Population | ||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| SARS-CoV-2 Neutralization Assay | Sampling Time Pointa | COMIRNATY Original & Omicron BA.4/BA.5 Study 5 | COMIRNATY Subset of Study 4 | Age Group Comparison | Vaccine Group Comparison | |||||||||||||
| 18 Through 55 Years of Age | 56 Years of Age and Older | 56 Years of Age and Older | COMIRNATY Original & Omicron BA.4/BA.5 18 Through 55 Years of Age/ ≥ 56 Years of Age | COMIRNATY Original & Omicron BA.4/BA.5 /COMIRNATY ≥ 56 Years of Age | ||||||||||||||
| nb | Nc (%) (95% CId) | nb | Nc (%) (95% CId) | nb | Nc (%) (95% CId) | Differencee (95% CIf) | Differencee (95% CIf) | |||||||||||
| Omicron BA.4/BA.5 – NT50 (titer)g | 1 month | 294 | 180 (61.2) (55.4, 66.8) | 282 | 188 (66.7) (60.8, 72.1) | 273 | 127 (46.5) (40.5, 52.6) | -3.03 (-9.68, 3.63)h | 26.77 (19.59, 33.95)i | |||||||||
Abbreviations: CI = confidence interval; LLOQ = lower limit of quantitation; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. Note: Seroresponse is defined as achieving a ≥ 4-fold rise from baseline. If the baseline measurement is below the LLOQ, a postvaccination assay result ≥ 4 × LLOQ is considered a seroresponse. a. Protocol-specified timing for blood sample collection. b. N = Number of participants with valid and determinate assay results for the specified assay at both the prevaccination time point and the given sampling time point. This value is the denominator for the percentage calculation. c. n = Number of participants with seroresponse for the given assay at the given sampling time point. d. Exact 2-sided CI, based on the Clopper and Pearson method. e. Difference in proportions, expressed as a percentage. f. 2-sided CI based on the Miettinen and Nurminen method stratified by baseline neutralizing titer category (< median, ≥ median) for the difference in proportions. The median of baseline neutralizing titers was calculated based on the pooled data in 2 comparator groups. g. SARS-CoV-2 NT50 were determined using a validated 384-well assay platform (Omicron B.1.1.529 subvariant BA.4/BA.5). h. Noninferiority is declared if the lower bound of the 2-sided 95% CI for the difference in percentages of participants with seroresponse is > -10%. i. Noninferiority is declared if the lower bound of the 2-sided 95% CI for the difference in percentages of participants with seroresponse is > -5%. | ||||||||||||||||||
| Table 31: Geometric Mean Titers by Baseline SARS-CoV-2 Status – COMIRNATY Original & Omicron BA.4/BA.5) Groups Subset of Study 5 – Prior to and 1 Month After Second Booster – Participants 12 Years of Age and Older – Evaluable Immunogenicity Population | ||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| SARS-CoV-2 Neutralization Assay | Baseline SARS-CoV-2 Status | Sampling Time Pointa | COMIRNATY Original & Omicron BA.4/BA.5 | |||||||||||||||||
| 12 Through 17 Years of Age | 18 Through 55 Years of Age | 56 Years of Age and Older | ||||||||||||||||||
| nb | GMTc (95% CIc) | nb | GMTc (95% CIc) | nb | GMTc (95% CIc) | |||||||||||||||
| Omicron BA.4/BA.5 – NT50 (titer)f | All | Pre- vaccination | 104 | 1105.8 (835.1, 1464.3) | 294 | 569.6 (471.4, 688.2) | 284 | 458.2 (365.2, 574.8) | ||||||||||||
| 1 Month | 105 | 8212.8 (6807.3, 9908.7) | 297 | 4455.9 (3851.7, 5154.8) | 284 | 4158.1 (3554.8, 4863.8) | ||||||||||||||
| Positived | Pre-vaccination | 78 | 1791.1 (1379.6, 2325.3) | 210 | 1181.4 (1005.3, 1388.3) | 174 | 1291.7 (1027.5, 1623.8) | |||||||||||||
| 1 Month | 79 | 9892.5 (8114.6, 12059.8) | 213 | 6031.6 (5203.9, 6991.0) | 176 | 6688.9 (5664.4, 7898.8) | ||||||||||||||
| Negativee | Pre-vaccination | 26 | 260.2 (157.1, 430.9) | 84 | 91.9 (71.5, 118.1) | 110 | 88.9 (69.8, 113.4) | |||||||||||||
| 1 Month | 26 | 4666.1 (3096.1, 7032.2) | 84 | 2067.7 (1530.2, 2793.9) | 108 | 1916.2 (1489.5, 2465.1) | ||||||||||||||
| Reference Strain – NT50 (titer)f | All | Pre-vaccination | 105 | 6863.3 (5587.8, 8430.1) | 296 | 4017.3 (3430.7, 4704.1) | 284 | 3690.6 (3082.2, 4419.0) | ||||||||||||
| 1 Month | 105 | 23641.3 (20473.1, 27299.8) | 296 | 16323.3 (14686.5, 18142.6) | 286 | 16250.1 (14499.2, 18212.4) | ||||||||||||||
| Positived | Pre-vaccination | 79 | 8685.4 (7062.7, 10680.9) | 213 | 7068.6 (6251.9, 7992.0) | 174 | 8082.1 (6843.6, 9544.8) | |||||||||||||
| 1 Month | 79 | 25991.8 (22377.5, 30189.8) | 212 | 19076.6 (17056.5, 21336.0) | 176 | 21273.3 (18604.2, 24325.3) | ||||||||||||||
| Negativee | Pre-vaccination | 26 | 3356.2 (2106.9, 5346.2) | 83 | 942.3 (705.6, 1258.3) | 110 | 1068.0 (835.9, 1364.6) | |||||||||||||
| 1 Month | 26 | 17725.2 (12376.4, 25385.7) | 84 | 11014.6 (8793.9, 13796.0) | 110 | 10560.6 (8827.1, 12634.5) | ||||||||||||||
Abbreviations: GMT = geometric mean titer; LLOQ = lower limit of quantitation; N-binding = SARS-CoV-2 nucleoprotein–binding; NAAT = nucleic acid amplification test; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. a. Protocol-specified timing for blood sample collection. b. n = Number of participants with valid and determinate assay results for the specified assay at the given sampling time point. c. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. d. Positive N-binding antibody result at baseline, positive NAAT result at baseline, or medical history of COVID-19. e. Negative N-binding antibody result at baseline, negative NAAT result at baseline, and no medical history of COVID-19. f. SARS-CoV-2 NT50 were determined using a validated 384-well assay platform (original strain [USA-WA1/2020, isolated in January 2020] and Omicron B.1.1.529 subvariant BA.4/BA.5). | ||||||||||||||||||||
14.2.2 COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg)
14.2.2.1 Immunogenicity in Participants 6 Months Through <5 Years of Age – After Booster (Fourth Dose)
In Study 6, a subset of 60 participants 6 months through <5 years of age received a booster (fourth dose) of COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) after receiving 3 prior doses of COMIRNATY 3 mcg. Neutralizing antibody levels following the fourth dose are presented in Table 32. Data from a subset of participants 6 months through <5 years of age in Study 3 who received 3 doses of COMIRNATY 3 mcg are included as a reference.
At 1 month after a booster dose (fourth dose), COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) elicited higher Omicron BA.4/BA.5 specific neutralizing titers (regardless of baseline SARS-CoV-2 status) compared with the titers in the comparator group who received 3 doses of COMIRNATY 3 mcg. COMIRNATY Original & Omicron BA.4/BA.5 (1.5/1.5 mcg) also elicited similar reference strain-specific titers compared with the titers in the comparator group.
| Table 32: Geometric Mean Titers – Study 6 Subset – Participants With or Without Evidence of Infection – 6 Months Through < 5 Years of Age – Evaluable Immunogenicity Population | ||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| SARS-CoV-2 Neutralization Assay | Age Group | Baseline (Dose 4 Study 6/ Dose 3 Study 3) SARS-CoV-2 Status | Sampling Time Pointa | Vaccine Group (as Assigned/Randomized) | ||||||||||||
Study 6 COMIRNATY Original & Omicron BA.4/BA.5 1.5/1.5 mcg Dose 4 and 1 Month After Dose 4 | Study 3 COMIRNATY 3 mcg Dose 3 and 1 Month After Dose 3 | |||||||||||||||
| nb | GMTc (95% CIc) | nb | GMTc (95% CIc) | |||||||||||||
| Omicron BA.4/BA.5 - NT50 (titer)d | 6 months through <5 years | Overall | Pre-vaccination | 54 | 192.5 (120.4, 307.8) | 54 | 70.5 (51.1, 97.2) | |||||||||
| 1 month | 58 | 1695.2 (1151.8, 2494.9) | 54 | 607.9 (431.1, 857.2) | ||||||||||||
| Positivee | Pre-vaccination | 16 | 1315.4 (789.1, 2192.8) | 15 | 351.7 (195.2, 633.8) | |||||||||||
| 1 month | 16 | 4897.7 (3085.5, 7774.1) | 15 | 1785.9 (1009.4, 3159.9) | ||||||||||||
| Negativef | Pre-vaccination | 38 | 85.7 (56.6, 129.8) | 36 | 38.2 (34.2, 42.8) | |||||||||||
| 1 month | 41 | 1116.0 (701.3, 1776.1) | 36 | 416.2 (287.8, 602.0) | ||||||||||||
Abbreviations: CI = confidence interval; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NAAT = nucleic acid amplification test; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. a. Protocol-specified timing for blood sample collection. b. n = Number of participants with valid and determinate assay results for the specified assay at the given sampling time point. c. GMTs and 2-sided 95% CIs were calculated by exponentiating the mean logarithm of the titers and the corresponding CIs (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. d. SARS-CoV-2 NT50 were determined using a validated 384-well assay platform (Omicron B.1.1.529 subvariant BA.4/BA.5). e. For Study 6: positive N-binding antibody result at Dose 4 visit, positive NAAT result at Dose 4 visit, or medical history of COVID-19. For Study 3: positive N-binding antibody result at Dose 1, 1-month post–Dose 2 (if available), or Dose 3 visits, positive NAAT result at Dose 1, Dose 2, Dose 3, or any unscheduled illness visit up to Dose 3 visit, or medical history of COVID-19. f. For Study 6: negative N-binding antibody result at Dose 4 visit, negative NAAT result at Dose 4 visit, and no medical history of COVID-19. For Study 3: negative N-binding antibody result at Dose 1, 1-month post–Dose 2 (if available), and Dose 3 visits, negative NAAT result at Dose 1, Dose 2, Dose 3, and any unscheduled illness visits up to Dose 3 visit, and no medical history of COVID-19. |
|---|
14.2.3 COMIRNATY (30 mcg)
14.2.3.1 Efficacy and Immunogenicity in Participants 16 Years of Age and Older
14.2.3.1.1 Efficacy in Participants 16 Years of Age and Older – After Two Doses
Primary Vaccine Efficacy Analysis (Based on Cut-off Date of November 14, 2020)
At the time of the primary efficacy analysis, participants had been followed for symptomatic COVID-19 for at least 2,214 person-years in the COMIRNATY group and at least 2,222 person-years in the placebo group.
There were no meaningful clinical differences in overall vaccine efficacy in participants who were at risk of severe COVID-19 including those with 1 or more comorbidities that increase the risk of severe COVID-19 [e.g., asthma, body mass index (BMI) ≥30 kg/m2, chronic pulmonary disease, diabetes mellitus, hypertension]. The primary endpoint was defined as any symptomatic COVID-19 case1 confirmed by Reverse Transcription-Polymerase Chain Reaction (RT-PCR). The population for the analysis of the primary efficacy endpoint included participants who did not have evidence of prior infection with SARS-CoV-2 through 7 days after the second dose (first primary efficacy endpoint), as well as participants with and without evidence of prior infections with SARS-CoV-2 through 7 days after the second dose (second primary efficacy endpoint). The pre-specified success criterion for vaccine efficacy was met. The vaccine efficacy information is presented in Table 33.
1 Case definition defined by Study 2 protocol: (at least 1 of) fever, new or increased cough, new or increased shortness of breath, chills, new or increased muscle pain, new loss of taste or smell, sore throat, diarrhea or vomiting.
| Table 33: Vaccine Efficacy – First COVID-19 Occurrence From 7 Days After Dose 2, by Age Subgroup – Participants Without Evidence of Infection and Participants With or Without Evidence of Infection Prior to 7 Days After Dose 2 – Evaluable Efficacy (7 Days) Population (Data Accrued Through November 14, 2020) | |||||||||
|---|---|---|---|---|---|---|---|---|---|
| First COVID-19 occurrence from 7 days after Dose 2 in participants without evidence of prior SARS-CoV-2 infection* | |||||||||
| Subgroup | COMIRNATY Na=18,198 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=18,325 Cases (n1b) Surveillance Timec (n2d) | Vaccine Efficacy % (95% CI) | ||||||
| All participantse | 8 2.214 (17,411) | 162 2.222 (17,511) | 95.0 (90.3, 97.6)f | ||||||
| 16 through 64 years | 7 1.706 (13,549) | 143 1.710 (13,618) | 95.1 (89.6, 98.1)g | ||||||
| 65 years and older | 1 0.508 (3,848) | 19 0.511 (3,880) | 94.7 (66.7, 99.9)g | ||||||
| First COVID-19 occurrence from 7 days after Dose 2 in participants with or without* evidence of prior SARS-CoV-2 infection | |||||||||
| Subgroup | COMIRNATY Na=19,965 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=20,172 Cases (n1b) Surveillance Timec (n2d) | Vaccine Efficacy % (95% CI) | ||||||
| All participantse | 9 2.332 (18,559) | 169 2.345 (18,708) | 94.6 (89.9, 97.3)f | ||||||
| 16 through 64 years | 8 1.802 (14,501) | 150 1.814 (14,627) | 94.6 (89.1, 97.7)g | ||||||
| 65 years and older | 1 0.530 (4,044) | 19 0.532 (4,067) | 94.7 (66.8, 99.9)g | ||||||
Note: Confirmed cases were determined by Reverse Transcription-Polymerase Chain Reaction (RT-PCR) and at least 1 symptom consistent with COVID-19 (symptoms included: fever; new or increased cough; new or increased shortness of breath; chills; new or increased muscle pain; new loss of taste or smell; sore throat; diarrhea; vomiting). Abbreviations: NAAT = nucleic acid amplification test; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2; VE = vaccine efficacy. * Participants who had no evidence of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative at Visit 1 and SARS-CoV-2 not detected by NAAT [nasal swab] at Visits 1 and 2), and had negative NAAT (nasal swab) at any unscheduled visit prior to 7 days after Dose 2 were included in the analysis. a. N = number of participants in the specified group. b. n1 = Number of participants meeting the endpoint definition. c. Total surveillance time in 1000 person-years for the given endpoint across all participants within each group at risk for the endpoint. Time period for COVID-19 case accrual is from 7 days after Dose 2 to the end of the surveillance period. d. n2 = Number of participants at risk for the endpoint. e. No confirmed cases were identified in adolescents 12 to 15 years of age. f. Two-sided credible interval for vaccine efficacy (VE) was calculated using a beta-binomial model with a beta (0.700102, 1) prior for θ=r(1-VE)/(1+r(1-VE)), where r is the ratio of surveillance time in the active vaccine group over that in the placebo group. g. Two-sided confidence interval (CI) for VE is derived based on the Clopper and Pearson method adjusted to the surveillance time. | |||||||||
Updated Vaccine Efficacy (Based on Cut-off Date of March 13, 2021)
Updated efficacy analyses were performed with additional confirmed COVID-19 cases accrued during blinded placebo-controlled follow-up through March 13, 2021, representing up to 6 months of follow-up after Dose 2 for participants in the efficacy population. There were 77 confirmed COVID-19 cases identified in the COMIRNATY and 850 in the placebo groups, respectively. In this analysis, compared to placebo, the vaccine efficacy of COMIRNATY in participants without evidence of SARS-CoV-2 infection prior to 7 days after Dose 2 was 91.3% (95% confidence interval of 89.0% to 93.2%); in participants 65 years of age and older without evidence of prior infection vaccine efficacy was 94.5% (two-sided 95% confidence interval 88.3% to 97.8%). The vaccine efficacy of COMIRNATY in participants with or without evidence of prior infection was 91.1% (95% confidence interval: 88.8% to 93.0%) with 81 COVID-19 cases in the COMIRNATY group compared to 873 cases in the placebo group.
Efficacy Against Severe COVID-19 (Based on Cut-off Date of March 13, 2021)
Secondary efficacy analyses in Study 2 supported benefit of COMIRNATY in preventing severe COVID-19. During blinded placebo-controlled follow-up through March 13, 2021, the vaccine efficacy against severe COVID 19 (as defined by the study protocol) in participants with or without evidence of SARS-CoV-2 infection prior to 7 days after Dose 2 was 95.3% (95% CI: 70.9%, 99.9%) with 1 and 21 cases in the vaccine and placebo groups, respectively. The COVID-19 case counts in participants without prior SARS-CoV-2 infection were the same as those in participants with or without prior SARS-CoV-2 infection in both the COMIRNATY and placebo groups.
14.2.3.1.2 Efficacy and Immunogenicity in Participants 16 Years of Age and Older – After Booster Dose
Immunogenicity in Participants 18 to 55 Years of Age – After Booster Dose
Noninferiority of immune responses 1 month after a COMIRNATY booster dose compared to 1 month after completion of the primary 2-dose series was assessed, in a subset of participants enrolled at selected sites in the US, by evaluating SARS-CoV-2 50% neutralizing titers (NT50) against the reference strain. Immunogenicity was evaluated in subjects who had no serological or virological evidence of past SARS-CoV-2 infection up to 1 month after the booster vaccination. The analysis demonstrated noninferior immune responses 1 month after a booster dose compared to 1 month after Dose 2 in individuals 18 through 55 years of age (Table 34).
| Table 34: SARS-CoV-2 neutralization assay – NT50 (titer)† – GMT and seroresponse rate comparison of 1 month after booster dose to 1 month after primary series – participants without evidence of infection up to 1 month after booster dose* – booster dose evaluable immunogenicity population | |||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|
| Assay | n | COMIRNATY Sampling Time Point | 1 month after booster dose/
(97.5% CI) | Met noninferiority objective (Y/N) | |||||||
1 month after booster dose (95% CI) | 1 month after Dose 2 (95% CI) | ||||||||||
| Geometric mean 50% neutralizing titer (GMTb) | 210a | 2,476.4b (2,210.1, 2774.9) | 753.7 b (658.2, 863.1) | 3.29c (2.76, 3.91) | Yd | ||||||
| Seroresponse rate (%) for 50% neutralizing titer | 198e | 197f 99.5% (97.2%, 100.0%) | 194f 98.0% (94.4%, 99.4%) | 1.5%g (-0.7%, 3.7%h) | Yi | ||||||
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; N-binding = SARS-CoV-2 nucleoprotein-binding; NAAT = nucleic acid amplification test; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2; Y/N = yes/no. † SARS-CoV-2 NT50 were determined using the SARS-CoV-2 mNeonGreen Virus Microneutralization Assay. The assay uses a fluorescent reporter virus derived from the USA_WA1/2020 strain and virus neutralization is read on Vero cell monolayers. The sample NT50 is defined as the reciprocal serum dilution at which 50% of the virus is neutralized. * Participants who had no serological or virological evidence (up to 1 month after receipt of a booster dose of COMIRNATY) of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative and SARS-CoV-2 not detected by NAAT [nasal swab]) and had a negative NAAT (nasal swab) at any unscheduled visit up to 1 month after the booster dose were included in the analysis. a. n = Number of participants with valid and determinate assay results at both sampling time points within specified window. b. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. c. GMRs and 2-sided 97.5% Cis were calculated by exponentiating the mean differences in the logarithms of the assay and the corresponding Cis (based on the Student t distribution). d. Noninferiority is declared if the lower bound of the 2-sided 97.5% CI for the GMR is >0.67 and the point estimate of the GMR is ≥0.80. e. n = Number of participants with valid and determinate assay results for the specified assay at baseline, 1 month after Dose 2 and 1 month after the booster dose within specified window. These values are the denominators for the percentage calculations. f. Number of participants with seroresponse for the given assay at the given dose/sampling time point. Exact 2-sided CI based on the Clopper and Pearson method. Note: Seroresponse is defined as achieving a ≥4-fold rise from baseline (before Dose 1). If the baseline measurement is below the LLOQ, a postvaccination assay result ≥4 × LLOQ is considered a seroresponse. g. Difference in proportions, expressed as a percentage (1 month after booster dose – 1 month after Dose 2). h. Adjusted Wald 2-sided CI for the difference in proportions, expressed as a percentage. i. Noninferiority is declared if the lower bound of the 2-sided 97.5% CI for the percentage difference is >-10%. | |||||||||||
Relative Vaccine Efficacy in Participants 16 Years of Age and Older – After Booster Dose
An interim efficacy analysis of Study 4, a placebo-controlled booster study was performed in approximately 10,000 participants 16 years of age and older who were recruited from Study 2 and evaluated confirmed COVID-19 cases accrued from at least 7 days after booster vaccination up to a data cut-off date of 5 October 2021, which represents a median of 2.5 months post-booster follow-up. Vaccine efficacy of the COMIRNATY booster dose after the primary series relative to the placebo booster group who only received the primary series dose was assessed. The relative vaccine efficacy information for participants 16 years of age and older is presented in Table 35.
| Table 35: Vaccine Efficacy – First COVID-19 Occurrence From 7 Days After Booster Vaccination – Participants 16 Years of Age and Older Without Evidence of Infection and Participants With or Without Evidence of Infection Prior to 7 Days After Booster Vaccination – Evaluable Efficacy Population | ||||||
|---|---|---|---|---|---|---|
| First COVID-19 occurrence from 7 days after booster dose in participants without evidence of prior SARS-CoV-2 infection* | ||||||
COMIRNATY Na=4,695 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=4,671 Cases (n1b) Surveillance Timec (n2d) | Relative Vaccine Efficacye % (95% CIf) | ||||
| First COVID-19 occurrence from 7 days after booster vaccination | 6 0.823 (4,659) | 123 0.792 (4,614) | 95.3 (89.5, 98.3) | |||
| First COVID-19 occurrence from 7 days after booster dose in participants with or without evidence of prior SARS-CoV-2 infection | ||||||
COMIRNATY Na=4,993 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=4,952 Cases (n1b) Surveillance Timec (n2d) | Relative Vaccine Efficacye % (95% CIf) | ||||
| First COVID-19 occurrence from 7 days after booster vaccination | 7 0.871 (4,934) | 124 0.835 (4,863) | 94.6 (88.5, 97.9) | |||
Note: Confirmed cases were determined by Reverse Transcription-Polymerase Chain Reaction (RT-PCR) and at least 1 symptom consistent with COVID-19 (symptoms included: fever; new or increased cough; new or increased shortness of breath; chills; new or increased muscle pain; new loss of taste or smell; sore throat; diarrhea; vomiting). * Participants who had no serological or virological evidence (prior to 7 days after receipt of the booster vaccination) of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative at Visit 1 and SARS-CoV-2 not detected by NAAT [nasal swab] at Visit 1, and had a negative NAAT [nasal swab] at any unscheduled visit prior to 7 days after booster vaccination) were included in the analysis. a. N = Number of participants in the specified group. b. n1 = Number of participants meeting the endpoint definition. c. Total surveillance time in 1000 person-years for the given endpoint across all participants within each group at risk for the endpoint. Time period for COVID-19 case accrual is from 7 days after the booster vaccination to the end of the surveillance period. d. n2 = Number of participants at risk for the endpoint. e. Relative vaccine efficacy of the COMIRNATY booster group relative to the placebo group (non-booster). f. Two-sided confidence interval (CI) for relative vaccine efficacy is derived based on the Clopper and Pearson method adjusted for surveillance time. | ||||||
14.2.3.2 Efficacy and Immunogenicity in Adolescents 12 to 15 Years of Age
14.2.3.2.1 Efficacy and Immunogenicity in Adolescents 12 to 15 Years of Age – After Two Doses
Efficacy
The vaccine efficacy in participants 12 to 15 years of age was evaluated on a subgroup analysis of Study 2 based on a cut-off date of March 13, 2021 (Table 36).
| Table 36: Vaccine Efficacy – First COVID-19 Occurrence From 7 Days After Dose 2, Without Evidence of Infection and With or Without Evidence of Infection Prior to 7 Days After Dose 2 – Blinded Placebo-Controlled Follow-up Period (Data Accrued Through March 13, 2021), Adolescents 12 to 15 Years of Age Evaluable Efficacy (7 Days) Population | |||||||
|---|---|---|---|---|---|---|---|
| First COVID-19 occurrence from 7 days after Dose 2 in adolescents 12 to 15 years of age without evidence of prior SARS-CoV-2 infection* | |||||||
COMIRNATY Na=1,005 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=978 Cases (n1b) Surveillance Timec (n2d) | Vaccine Efficacy % (95% CIe) | |||||
| Adolescents 12 to 15 Years of Age | 0 0.154 (1,001) | 16 0.147 (972) | 100.0 (75.3, 100.0) | ||||
| First COVID-19 occurrence from 7 days after Dose 2 in adolescents 12 to 15 years of age with or without* evidence of prior SARS-CoV-2 infection | |||||||
COMIRNATY Na=1,119 Cases (n1b) Surveillance Timec (n2d) | Placebo Na=1,110 Cases (n1b) Surveillance Timec (n2d) | Vaccine Efficacy % (95% CIe) | |||||
| Adolescents 12 to 15 Years of Age | 0 0.170 (1,109) | 18 0.163 (1094) | 100.0 (78.1, 100.0) | ||||
Note: Confirmed cases were determined by Reverse Transcription-Polymerase Chain Reaction (RT-PCR) and at least 1 symptom consistent with COVID-19 (symptoms included: fever; new or increased cough; new or increased shortness of breath; chills; new or increased muscle pain; new loss of taste or smell; sore throat; diarrhea; vomiting). * Participants who had no evidence of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative at Visit 1 and SARS-CoV-2 not detected by NAAT [nasal swab] at Visits 1 and 2), and had negative NAAT (nasal swab) at any unscheduled visit prior to 7 days after Dose 2 were included in the analysis. a. N = Number of participants in the specified group. b. n1 = Number of participants meeting the endpoint definition. c. Total surveillance time in 1000 person-years for the given endpoint across all participants within each group at risk for the endpoint. Time period for COVID-19 case accrual is from 7 days after Dose 2 to the end of the surveillance period. d. n2 = Number of participants at risk for the endpoint. e. Confidence interval (CI) for vaccine efficacy is derived based on the Clopper and Pearson method adjusted for surveillance time. | |||||||
An updated efficacy analysis of Study 2 has been performed in approximately 2,260 adolescents 12 through 15 years of age evaluating confirmed COVID-19 cases accrued up to a data cut-off date of September 2, 2021, representing up to 6 months of follow-up after Dose 2 for participants in the efficacy population.
In the adolescent group, in efficacy analyses in the evaluable efficacy population based on cases reported from at least 7 days after Dose 2 through the data cut-off date (02 September 2021), representing a median of 4.4 (range 0-10.8) months of follow-up after Dose 2, there were 0 confirmed COVID-19 cases identified in the COMIRNATY and 28 in the placebo groups, respectively. In this analysis, compared to placebo, the estimated VE against confirmed COVID-19 was 100% (95% CI: 86.8%, 100%) for individuals without evidence of prior SARS-CoV-2 infection before and during vaccination regimen. The estimated VE against confirmed COVID-19 was 100% (2-sided 95% CI: 87.5%, 100%) for those with or without evidence of prior SARS-CoV-2 infection before and during vaccination regimen, with 0 COVID-19 cases in the COMIRNATY group compared to 30 cases in the placebo group.
Among participants without and with or without evidence of SARS-CoV-2 infection before and during the vaccination regimen (evaluable efficacy population), VE against COVID-19 occurring at least 7 days after Dose 2 was evaluated for demographic and risk subgroups, and the estimated VE was 100.0% for all subgroups.
Immunogenicity – After Two Doses
In Study 2, an analysis of SARS-CoV-2 50% neutralizing titers (NT50) 1 month after Dose 2 in a randomly selected subset of participants demonstrated non-inferior immune responses (within 1.5-fold) comparing adolescents 12 through 15 years of age to participants 16 through 25 years of age who had no serological or virological evidence of past SARS-CoV-2 infection up to 1 month after Dose 2 (Table 37).
Table 37: Summary of Geometric Mean Ratio for 50% Neutralizing Titer – Comparison of Adolescents 12 Through 15 Years of Age to Participants 16 Through 25 Years of Age (Immunogenicity Subset) –Participants Without Evidence of Infection up to 1 Month After Dose 2 – Dose 2 Evaluable Immunogenicity Population
| COMIRNATY | 12 Through 15 Years/ 16 Through 25 Years | ||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|
12 Through 15 Years na=190 | 16 Through 25 Years na=170 | ||||||||||
| Assay | Time Pointb | GMTc (95% CIc) | GMTc (95% CIc) | GMRd (95% CId) | Met Noninferiority Objectivee (Y/N) | ||||||
| SARS-CoV-2 neutralization assay – NT50 (titer)f | 1 month after Dose 2 | 1,239.5 (1,095.5, 1,402.5) | 705.1 (621.4, 800.2) | 1.76 (1.47, 2.10) | Y | ||||||
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NAAT = nucleic-acid amplification test; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. Note: Participants who had no serological or virological evidence (up to 1 month after receipt of the last dose) of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative at Visit 1 and SARS-CoV-2 not detected by NAAT [nasal swab] at Visits 1 and 2), and had negative NAAT (nasal swab) at any unscheduled visit up to 1 month after Dose 2 were included in the analysis. a. n = Number of participants with valid and determinate assay results for the specified assay at the given dose/sampling time point. b. Protocol-specified timing for blood sample collection. c. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. | |||||||||||
d. GMRs and 2-sided 95% Cis were calculated by exponentiating the mean difference of the logarithms of the titers (Group 1 [12 through 15 years of age] – Group 2 [16 through 25 years of age]) and the corresponding CI (based on the Student t distribution). e. Noninferiority is declared if the lower bound of the 2-sided 95% CI for the GMR is greater than 0.67. f. SARS-CoV-2 NT50 were determined using the SARS-CoV-2 mNeonGreen Virus Microneutralization Assay. The assay uses a fluorescent reporter virus derived from the USA_WA1/2020 strain and virus neutralization is read on Vero cell monolayers. The sample NT50 is defined as the reciprocal serum dilution at which 50% of the virus is neutralized. |
|---|
14.2.4 COMIRNATY (10 mcg)
14.2.4.1 Efficacy and Immunogenicity in Children 5 Years Through <12 Years of Age
14.2.4.1.1 Efficacy and Immunogenicity in Children 5 Years Through <12 Years of Age – After Two Doses
Immunogenicity
In Study 3, an analysis of SARS-CoV-2 50% neutralizing titers (NT50) 1 month after Dose 2 in a randomly selected subset of participants demonstrated effectiveness by immunobridging of immune responses. Children 5 years through <12 years of age in the Phase 2/3 part of Study 3 were compared to participants 16 through 25 years of age in the Phase 2/3 part of Study 2 who had no serological or virological evidence of past SARS-CoV-2 infection up to 1 month after Dose 2. The study met the prespecified immunobridging criteria for both the geometric mean ratio (GMR) and the seroresponse difference with seroresponse defined as achieving at least 4-fold rise in SARS-CoV-2 NT50 from baseline (before Dose 1). The ratio of the SARS-CoV-2 NT50 in children 5 years through <12 years of age to that of young adults 16 to 25 years of age was 1.04 (2-sided 95% CI: 0.93, 1.18), meeting the 1.5-fold noninferiority criterion (the lower bound of the 2-sided 95% CI for the geometric mean ratio [GMR] >0.67). Results are presented in Table 38.
| Table 38: Summary of Geometric Mean Ratio for 50% Neutralizing Titer And Difference in Percentages of Participants with Seroresponse – Comparison of Children 5 Years Through < 12 Years of Age (Study 3) to Participants 16 Through 25 Years of Age (Study 2) – Participants Without Evidence of Infection up to 1 Month After Dose 2 – Immunobridging Subset – Dose 2 Evaluable Immunogenicity Population | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Geometric Mean Titers (NT50) | |||||||||||||
| COMIRNATY | 5 Years Through <12 Years/ 16 Through 25 Years | ||||||||||||
10 mcg/Dose 5 Years Through <12 Years Na=264 | 30 mcg/Dose 16 Through 25 Years Na=253 | ||||||||||||
| Assay | Time Pointb | GMTc (95% CIc) | GMTc (95% CIc) | GMRd (95% CId) | Met Immunobridging Objectivee (Y/N) | ||||||||
| SARS-CoV-2 neutralization assay – NT50 (titer)f | 1 month after Dose 2 | 1,197.6 (1,106.1, 1,296.6) | 1,146.5 (1,045.5, 1,257.2) | 1.04 (0.93, 1.18) | Y | ||||||||
| Seroresponse Rate | |||||
|---|---|---|---|---|---|
| COMIRNATY | 5 Years Through <12 Years/ 16 Through 25 Years | ||||
10 mcg/Dose 5 Years Through <12 Years Ng=264 | 30 mcg/Dose 16 Through 25 Years Ng=253 | ||||
| Assay | Time Pointb | nh (%) (95% CIi) | nh (%) (95% CIi) | Difference %j (95% CIk) | Met Immunobridging Objectivel (Y/N) |
| SARS-CoV-2 neutralization assay – NT50 (titer)f | 1 month after Dose 2 | 262 (99.2) (97.3, 99.9) | 251 (99.2) (97.2, 99.9) | 0.0 (-2.0, 2.2) | Y |
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NAAT = nucleic-acid amplification test; N-binding = SARS-CoV-2 nucleoprotein- binding; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. Note: Participants who had no serological or virological evidence (up to 1 month post-Dose 2 blood sample collection) of past SARS-CoV-2 infection (i.e., N-binding antibody [serum] negative at Visit 1 and 1 month after Dose 2, SARS-CoV-2 not detected by NAAT [nasal swab] at Visits 1 and 2, and negative NAAT (nasal swab) at any unscheduled visit up to 1 month after Dose 2 blood collection) and had no medical history of COVID-19 were included in the analysis. Note: Seroresponse is defined as achieving a ≥4-fold rise from baseline (before Dose 1). If the baseline measurement is below the LLOQ, a postvaccination assay result ≥4 × LLOQ is considered a seroresponse. a. n = Number of participants with valid and determinate assay results for the specified assay at the given dose/sampling time point. b. Protocol-specified timing for blood sample collection. c. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. d. GMRs and 2-sided 95% Cis were calculated by exponentiating the mean difference of the logarithms of the titers (Group 1[5 through <12 years of age] – Group 2 [16 through 25 years of age]) and the corresponding CI (based on the Student t distribution). e. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the GMR is greater than 0.67 and the point estimate of the GMR is ≥0.8. f. SARS-CoV-2 NT50 were determined using the SARS-CoV-2 mNeonGreen Virus Microneutralization Assay. The assay uses a fluorescent reporter virus derived from the USA_WA1/2020 strain and virus neutralization is read on Vero cell monolayers. The sample NT50 is defined as the reciprocal serum dilution at which 50% of the virus is neutralized. g. N = number of participants with valid and determinate assay results both before vaccination and at 1 month after Dose 2. These values are the denominators for the percentage calculations. h. n = Number of participants with seroresponse for the given assay at the given dose/sampling time point. i. Exact 2-sided CI based on the Clopper and Pearson method. j. Difference in proportions, expressed as a percentage (Group 1 [5 through < 12 years of age] – Group 2 [16 through 25 years of age]). k. 2-Sided CI, based on the Miettinen and Nurminen method for the difference in proportions, expressed as a percentage. l. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the difference in proportions is greater than -10.0%. | |||||
Among participants without prior evidence of SARS-CoV-2 infection up to 1 month after Dose 2, 99.2% of children 5 years through <12 years of age and 99.2% of participants 16 through 25 years of age had a seroresponse from before vaccination to 1 month after Dose 2. The difference in proportions of participants who had seroresponse between the 2 age groups (children – young adult) was 0.0% (2-sided 95% CI: -2.0%, 2.2%), meeting the -10% noninferiority criterion (the lower bound of the 2-sided 95% CI for the difference in seroresponse rate>-10%). Results are presented in Table 38.
Efficacy
An exploratory efficacy analysis (based on a cut-off date of October 8, 2021) in participants 5 to less than 12 years of age without evidence of SARS-CoV-2 infection prior to Dose 2 showed that the observed vaccine efficacy against confirmed COVID-19 occurring at least 7 days after Dose 2 was 90.7% (95% CI: 67.7%, 98.3%), with 3 COVID-19 cases in the vaccine group compared to 16 in the placebo group (2:1 randomization in vaccine group to placebo group).
No severe COVID-19 or multisystem inflammatory syndrome in children (MIS-C) were reported in children 5 to less than 12 years of age, as of the data cut-off date (October 8, 2021).
14.2.4.2 Immunogenicity in Children 5 Years Through <12 Years of Age – After Booster Dose
Immunogenicity of a booster dose administered 7 to 9 months after the second primary series dose was evaluated in a subset of 67 evaluable study participants with no evidence of prior SARS-CoV-2 infection up to 1 month after the booster dose, and descriptively compared to 96 subjects in the same age group (67 participants randomly selected from the 2-dose analysis set and 29 participants in the 3-dose analysis set) with evaluable immunogenicity data following 2 doses of 10 mcg BNT162b2.
Vaccine effectiveness of a booster dose of COMIRNATY was inferred based on a descriptive analysis of NT50 against the reference strain of SARS-CoV-2 (USA_WA1/2020). The NT50 GMT at 1 month after the booster dose was increased compared to before the booster dose and after dose 2. See Table 39.
| Table 39: Summary of Geometric Mean Ratios – NT50 – Comparison of 1-Month After Dose 3 With 1-Month After Dose 2 – Participants Without Evidence of Infection – Phase 2/3 – Immunogenicity Set – 5 to <12 Years of Age – Evaluable Immunogenicity Population |
|---|
Post-Dose 2 N = 96 GMT (95% CI) | Pre-Booster N = 67 GMT (95% CI) | Post-Booster N = 67 GMT (95% CI) | GMR* Post-Booster/Post- Dose 2 N = 96 GMT (95% CI) |
|---|---|---|---|
1,253.9 (1,116.0, 1,408.9) | 270.1 (229.1, 320.6) | 2,720.9 (2,280.1, 3,247.0) | 2.17 (1.76, 2.68) |
* GMR and confidence interval based on post-hoc descriptive analysis. Using a non-validated fluorescence focus reduction neutralization test assay against the Omicron variant of SARS-CoV-2 (B.1.1.529), the NT50 GMT at 1 month after the booster dose among a subset of 17 study participants (614.4 [95% CI: 410.7, 919.2]) was increased compared to the NT50 GMT at 1 month after dose 2 among a subset of 29 study participants (27.6 [95% CI: 22.1, 34.5]). |
|---|
14.2.5 COMIRNATY (3 mcg)
14.2.5.1 Immunogenicity in Children 6 Months Through <5 Years of Age
14.2.5.1.1 Immunogenicity in Children 6 Months Through <5 Years of Age – After a 3-Dose Primary Series
Effectiveness in individuals 6 months through <5 years of age is based on a comparison of immune responses in this age group to individuals 16 through 25 years of age.
Immunogenicity in Children 2 Through <5 Years of Age
Immunogenicity analyses have been performed in the immunobridging subset of 143 Study 3 participants 2 through <5 years of age without evidence of infection up to 1 month after Dose 3 based on a data cut-off date of April 29, 2022.
SARS-CoV-2 50% neutralizing antibody titers (NT50) were compared between an immunogenicity subset of Phase 2/3 participants 2 through <5 years of age from Study 3 at 1 month after the 3-dose primary course and a randomly selected subset from Study 2 Phase 2/3 participants 16 through 25 years of age at 1 month after the 2-dose primary series, using a microneutralization assay against the reference strain (USA_WA1/2020). The primary immunobridging analyses compared the geometric mean titers (using a geometric mean ratio [GMR]) and the seroresponse (defined as achieving at least 4-fold rise in SARS-CoV-2 NT50 from before Dose 1) rates in the evaluable immunogenicity population of participants without evidence of prior SARS-CoV-2 infection up to 1 month after Dose 3 in participants 2 through <5 years of age and up to 1 month after Dose 2 in participants 16 through 25 years of age. The prespecified immunobridging criteria were met for both the GMR and the seroresponse difference (Table 40).
| Table 40: SARS-CoV-2 GMTs (NT50) at 1 Month After Vaccination Series and Difference in Percentages of Participants with Seroresponse at 1 Month After Vaccination Series – Immunobridging Subset – Participants 2 Through <5 Years of Age (Study 3) 1 Month after Dose 3 and Participants 16 Through 25 Years of Age (Study 2) 1 Month After Dose 2 – Without Evidence of SARS-CoV-2 Infection – Evaluable Immunogenicity Population | ||||||||||
|---|---|---|---|---|---|---|---|---|---|---|
| Geometric Mean Titers (NT50) | ||||||||||
| COMIRNATY | GMR (95%CI) (2 Through <5 Years of Age/ 16 Through 25 Years of Age)c,d | |||||||||
3 mcg/Dose 2 Through <5 Years of Age (1 Month after Dose 3) na=143 | 30 mcg/Dose 16 Through 25 Years of Age (1 Month after Dose 2) na=170 | |||||||||
| Assay | GMTb (95% CIb) | GMTb (95% CIb) | ||||||||
| SARS-CoV-2 neutralization assay – NT50 (titer)e | 1,535.2 (1,388.2, 1,697.8) | 1,180.0 (1,066.6, 1305.4) | 1.30 (1.13, 1.50) | |||||||
| Seroresponse Rate | |||
|---|---|---|---|
| COMIRNATY | Difference in Seroresponse Rates %i (95% CIj) (2 Through <5 Years of Age minus 16 Through 25 years of age)k | ||
3 mcg/Dose 2 Through <5 Years of Age (1 Month After Dose 3) Nf=141 | 30 mcg/Dose 16 Through 25 Years of Age (1 Month after Dose 2) Nf=170 | ||
| Assay | ng (%) (95% CIh) | ng (%) (95% CIh) | |
| SARS-CoV-2 neutralization assay – NT50 (titer)e | 141 (100.0) (97.4, 100.0) | 168 (98.8) (95.8, 99.9) | 1.2 (-1.5, 4.2) |
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NAAT = nucleic acid amplification test; N-binding = SARS-CoV-2 nucleoprotein binding; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. Note: Participants who had no serological or virological evidence [(up to 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3) blood sample collection)] of past SARS-CoV-2 infection [(i.e. N-binding antibody [serum] negative at pre-Dose 1, pre-Dose 3 (Study 3) and 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3), SARS-CoV-2 not detected by NAAT [nasal swab] at pre-Dose 1, pre-Dose 2, and pre-Dose 3 (Study 3) study visits, and negative NAAT (nasal swab) at any unscheduled visit up to 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3) blood collection)] and had no medical history of COVID-19 were included in the analysis. Note: Seroresponse is defined as achieving a ≥ 4-fold rise from baseline (before Dose 1). If the baseline measurement is below the LLOQ, a postvaccination assay result ≥ 4 × LLOQ is considered a seroresponse. a. n = Number of participants with valid and determinate assay results for the specified assay at the given dose/sampling time point. b. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. c. GMRs and 2-sided 95% Cis were calculated by exponentiating the mean difference of the logarithms of the titers (2 through <5 years of age minus 16 through 25 years of age) and the corresponding CI (based on the Student t distribution). d. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the GMR ratio is greater than 0.67 and the point estimate of the GMR is ≥ 0.8. e. SARS-CoV-2 NT50 were determined using the SARS-CoV-2 mNeonGreen Virus Microneutralization Assay. The assay uses a fluorescent reporter virus derived from the USA_WA1/2020 strain and virus neutralization is read on Vero cell monolayers. The sample NT50 is defined as the reciprocal serum dilution at which 50% of the virus is neutralized. f. N = number of participants with valid and determinate assay results both before vaccination and at 1 month after Dose 2. These values are the denominators for the percentage calculations. g. n = Number of participants with seroresponse for the given assay at the given dose/sampling time point. h. Exact 2-sided CI based on the Clopper and Pearson method. i. Difference in proportions, expressed as a percentage (2 through <5 years of age minus 16 through 25 years of age). j. 2-sided CI, based on the Miettinen and Nurminen method for the difference in proportions, expressed as a percentage. k. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the difference in proportions is greater than -10.0% provided that the immunobridging criteria based on GMR were met. | |||
Using a non-validated fluorescence focus reduction neutralization test assay against the Omicron variant of SARS-CoV-2 (BA.1), the NT50 GMT at 1 month after Dose 3 among a subset of 34 study participants without evidence of prior SARS-CoV-2 infection (82.5 [2-sided 95% CI: 55.4, 122.9]) was increased compared to the NT50 GMT before Dose 3 (14.0 [2-sided 95% CI: 10.6, 18.5]).
Immunogenicity in Children 6 Months Through <2 Years of Age
Immunogenicity analyses have been performed in the immunobridging subset of 82 Study 3 participants 6 months through <2 years of age without evidence of infection up to 1 month after Dose 3 based on a data cut-off date of 29 April 2022.
SARS-CoV-2 50% neutralizing antibody titers (NT50) 1 month after the vaccination series were compared between an immunogenicity subset of Phase 2/3 participants 6 months through <2 years of age from Study 3 and a randomly selected subset from Study 2 Phase 2/3 participants 16 through 25 years of age, using a microneutralization assay against the reference strain (USA_WA1/2020). The primary immunobridging analyses compared the geometric mean titers (using a GMR) and the seroresponse (defined as achieving at least 4-fold rise in SARS-CoV-2 NT50 from before Dose 1) rates in the evaluable immunogenicity population of participants without evidence of prior SARS-CoV-2 infection up to 1 month after Dose 3 in participants 6 months through < 2 years of age and up to 1 month after Dose 2 in participants 16 through 25 years of age. The prespecified immunobridging criteria were met for both the GMR and the seroresponse difference (Table 41).
Table 41: SARS-CoV-2 GMTs (NT50) at 1 Month after Vaccination Series and Difference in Percentages of Participants With Seroresponse at 1 Month After Vaccination Series – Immunobridging Subset – Participants 6 Months Through <2 Years of Age (Study 3) 1 Month After Dose 3 and Participants 16 Through 25 Years of Age (Study 2) 1 Month After Dose 2 – Without Evidence of SARS-CoV-2 Infection– Evaluable Immunogenicity Population
| Geometric Mean Titers (NT50) | |||
|---|---|---|---|
| COMIRNATY | GMR (95%CI) (6 Months Through <2 Years of Age/ 16 Through 25 Years of Age)c,d | ||
3 mcg/Dose 6 Months Through <2 Years of Age (1 Month After Dose 3) na=82 | 30 mcg/Dose 16 Through 25 Years of Age (1 Month After Dose 2) na=170 | ||
| Assay | GMTb (95% CIb) | GMTb (95% CIb) | |
| SARS-CoV-2 neutralization assay – NT50 (titer)e | 1,406.5 (1,211.3, 1,633.1) | 1,180.0 (1,066.6, 1,305.4) | 1.19 (1.00, 1.42) |
| Seroresponse Rate | |||
|---|---|---|---|
| COMIRNATY | Difference in Seroresponse Rates %i (95% CIj) (6 Months Through <2 Years of Age minus 16 Through 25 Years of Age)k | ||
3 mcg/Dose 6 Months Through <2 Years of Age (1 Month After Dose 3) Nf=80 | 30 mcg/dose 16 Through 25 Years of Age (1 Month After Dose 2) Nf=170 | ||
| Assay | ng (%) (95% CIh) | ng (%) (95% CIh) | |
| SARS-CoV-2 neutralization assay – NT50 (titer)e | 80 (100.0) (95.5, 100.0) | 168 (98.8) (95.8, 99.9) | 1.2 (-3.4, 4.2) |
Abbreviations: CI = confidence interval; GMR = geometric mean ratio; GMT = geometric mean titer; LLOQ = lower limit of quantitation; NAAT = nucleic acid amplification test; N-binding = nucleoprotein binding; NT50 = 50% neutralizing titer; SARS-CoV-2 = severe acute respiratory syndrome coronavirus 2. Note: Participants who had no serological or virological evidence [(up to 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3) blood sample collection)] of past SARS-CoV-2 infection (i.e. N-binding antibody [serum] negative at pre-Dose 1, pre-Dose 3 (Study 3) and 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3), SARS-CoV-2 not detected by NAAT [nasal swab] at pre-Dose 1, pre-Dose 2, and pre-Dose 3 (Study 3) study visits, and negative NAAT (nasal swab) at any unscheduled visit up to 1 month after Dose 2 (Study 2) or 1 month after Dose 3 (Study 3) blood collection)] and had no medical history of COVID-19 were included in the analysis. Note: Seroresponse is defined as achieving a ≥ 4-fold rise from baseline (before Dose 1). If the baseline measurement is below the LLOQ, a postvaccination assay result ≥ 4 × LLOQ is considered a seroresponse. a. n = Number of participants with valid and determinate assay results for the specified assay at the given dose/sampling time point. b. GMTs and 2-sided 95% Cis were calculated by exponentiating the mean logarithm of the titers and the corresponding Cis (based on the Student t distribution). Assay results below the LLOQ were set to 0.5 × LLOQ. c. GMRs and 2-sided 95% Cis were calculated by exponentiating the mean difference of the logarithms of the titers (6 months through < 2 years of age minus 16 through 25 years of age) and the corresponding CI (based on the Student t distribution). d. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the GMR ratio is greater than 0.67 and the point estimate of the GMR is ≥0.8. e. SARS-CoV-2 NT50 were determined using the SARS-CoV-2 mNeonGreen Virus Microneutralization Assay. The assay uses a fluorescent reporter virus derived from the USA_WA1/2020 strain and virus neutralization is read on Vero cell monolayers. The sample NT50 is defined as the reciprocal serum dilution at which 50% of the virus is neutralized. f. N = number of participants with valid and determinate assay results both before vaccination and at 1 month after Dose 2. These values are the denominators for the percentage calculations. g. n = Number of participants with seroresponse for the given assay at the given dose/sampling time point. h. Exact 2-sided CI based on the Clopper and Pearson method. i. Difference in proportions, expressed as a percentage (6 months through <2 years of age minus 16 through 25 years of age). j. 2-sided CI, based on the Miettinen and Nurminen method for the difference in proportions, expressed as a percentage. k. Immunobridging is declared if the lower bound of the 2-sided 95% CI for the difference in proportions is greater than -10.0% provided that the immunobridging criteria based on GMR were met. | |||
Using a non-validated fluorescence focus reduction neutralization test assay against the Omicron variant of SARS-CoV-2 (BA.1), the NT50 GMT at 1 month after Dose 3 among a subset of 32 study participants without evidence of prior SARS-CoV-2 infection (127.5 [2-sided 95% CI: 90.2, 180.1]) was increased compared to the NT50 GMT before Dose 3 (16.3 [2-sided 95% CI: 12.8, 20.8]).
15 MICROBIOLOGY
No microbiological information is required for this product.
16 NON-CLINICAL TOXICOLOGY
Non-clinical data reveal no special hazard for humans based on conventional studies of repeat dose toxicity.
General Toxicology:
In a repeat-dose toxicity study, rats were administered three once weekly doses of 30 mcg/animal (0.06 mL of a vaccine formulation containing the same quantity of nucleoside-modified messenger ribonucleic acid (mRNA) and other ingredients included in a single human dose) of COMIRNATY by intramuscular injection. Vaccine administration resulted in transient erythema and edema at the site of injection, as well as increased cellularity in draining and inguinal lymph nodes, spleen, and bone marrow, along with transiently increased body temperature, increased white blood counts, and decreased reticulocyte counts coupled with decreased red blood cell mass. Clinical chemistry changes (e.g., increased acute phase protein levels) indicated an acute phase response. These changes are consistent with an expected immunostimulatory response following intramuscular administration of a vaccine. Transient periportal hepatocyte vacuolation was also observed without evidence of liver injury. Full or partial recovery from all findings was observed following a 3-week recovery period.
Carcinogenicity:
Carcinogenic potential was not assessed, as carcinogenicity studies were not considered relevant to this vaccine.
Genotoxicity:
Genotoxic potential was not assessed, as genotoxicity studies were not considered relevant to this vaccine.
Reproductive and Developmental Toxicology:
In a reproductive and developmental toxicity study, 30 mcg/animal (0.06 mL of a vaccine formulation containing the same quantity of nucleoside-modified messenger ribonucleic acid (mRNA) and other ingredients included in a single human dose) of COMIRNATY was administered to female rats by the intramuscular route on four occasions: 21 and 14 days prior to mating, and on gestation days 9 and 20. No vaccine-related adverse effects on female fertility, fetal development, or postnatal development were reported in the study.