


Product Monograph - Euflex
ACTION and CLINICAL PHARMACOLOGY
Flutamide demonstrates potent antiandrogenic effects by inhibiting androgen uptake and/or inhibiting nuclear binding of androgen in target tissues. In adult male rats, ventral prostate weights and seminal vesicle weights were markedly reduced by daily administration of flutamide.
Pharmacokinetics
Analysis of plasma, urine, and feces following a single oral 200 mg dose of tritium-labelled flutamide to human volunteers showed that the drug is rapidly and completely absorbed. It is excreted mainly in the urine with 4.2% of the dose excreted in the faeces over 72 hours. The composition of plasma radioactivity showed that flutamide is rapidly and extensively metabolized, with flutamide comprising 2.5% of plasma radioactivity one hour after administration. At least six metabolites have been identified in plasma. The major plasma metabolite is a biologically active alpha-hydroxylated derivative, which accounts for 23% of the plasma tritium one hour after drug administration. The major urinary metabolite is 2-amino-5-nitro-4-(trifluoromethyl)phenol.
Following a single 250 mg oral dose to normal adult volunteers, low plasma levels of varying amounts of flutamide were detected. The biologically active alpha-hydroxylated metabolite reaches maximum plasma levels in about two hours, indicating that it is rapidly formed from flutamide. The plasma half-life for this metabolite is about 6 hours.
Following multiple oral dosing of 250 mg three times a day in normal geriatric volunteers, flutamide and its active metabolite approached steady-state plasma levels (based on pharmacokinetic simulations) after the fourth flutamide dose. The half-life of the active metabolite in geriatric volunteers after a single flutamide dose is about 8 hours and at steady-state is 9.6 hours.
Flutamide, in vivo, at steady-state plasma concentrations of 24 to 78 ng/mL is 94% to 96% bound to plasma proteins. The active metabolite of flutamide, in vivo, at steady-state plasma concentrations of 1556 to 2284 ng/mL, is 92% to 94% bound to plasma proteins.
In male rats neither flutamide nor any of its metabolites are preferentially accumulated in any tissue except the prostate after an oral 5 mg/kg dose of 14C-flutamide. Total drug levels were
highest 6 hours after drug administration in all tissues. Levels declined at roughly similar rates to low levels at 18 hours. The major metabolite was present at higher concentrations than flutamide in all tissues studied.
Elevations of plasma testosterone and estradiol levels have been noted following flutamide administration.
INDICATIONS AND CLINICAL USE
EUFLEX® (flutamide) Tablets are indicated for use in combination with LHRH agonistic analogues (such as leuprolide acetate) for the treatment of metastatic prostatic carcinoma (stage D2). To achieve the benefit of the adjunctive therapy with EUFLEX®, treatment must be started simultaneously using both drugs. EUFLEX® Tablets are also indicated as an adjunctive therapy to orchiectomy, in order to achieve complete androgen blockade.
EUFLEX® Tablets in combination with LHRH agonists are also indicated prior to and during definitive external beam radiotherapy for patients with bulky locally advanced Stage B2 and Stage C prostatic carcinoma. (See Dosage and Administration).
CONTRAINDICATIONS
EUFLEX® (flutamide) Tablets are contraindicated in patients who have shown hypersensitivity to flutamide or any component of this preparation.
EUFLEX® is contraindicated in patients with severe hepatic impairment.
EUFLEX® has not been studied in women and is not indicated for this population, particularly for nonserious or nonthreatening conditions.
WARNINGS
General
- Gynecomastia occurred in 9% of patients receiving flutamide together with medical castration.
- Physicians must familiarize themselves with the proper use of LHRH before combination medication is contemplated.
Antiandrogen Withdrawal Syndrome
In some patients with metastatic prostate cancer, antiandrogens (steroidal or non-steroidal), may promote, rather than inhibit, the growth of prostate cancer. A decrease in PSA and/or clinical improvement following the discontinuation of antiandrogens have been reported. It is recommended that patients prescribed an antiandrogen, who have PSA progression, should
have the antiandrogen discontinued immediately and be monitored for 6-8 weeks for a withdrawal response prior to any decision to proceed with other prostate cancer therapy.
Cardiovascular
EUFLEX® is indicated for use in combination with an LHRH analogue or orchiectomy. Based on evidence from the published literature, combined androgen blockade with an anti-androgen plus LHRH analogue increases risk of cardiovascular disease (heart attack, cardiac failure, sudden cardiac death) and adversely affects independent cardiovascular risk factors (serum lipoproteins, insulin sensitivity and obesity). Physicians should carefully consider whether the benefits of combined androgen blockade outweigh the potential cardiovascular risk. Assessment of cardiovascular risk factors, monitoring for signs and symptoms suggestive of development of cardiovascular disease, and management according to local clinical practice and guidelines should be considered.
Effect on QT/QTc interval
EUFLEX® is indicated for use in combination with an LHRH analogue or orchiectomy. The potential for QT/QTc prolongation has not been studied with EUFLEX®. Combined androgen blockade studies with other anti-androgen plus LHRH analogue or surgical castration have been associated with the potential to prolong QT/QTc interval on ECG. Physicians should consider whether the benefits of androgen deprivation therapy outweigh the potential risk in patients with congenital long QT syndrome, electrolyte abnormalities, or congestive heart failure and in patients taking Class IA (e.g. quinidine, procainamide), Class III (e.g. amiodarone, sotalol, dofetilide, ibutilide)), or Class IC (e.g. flecainide, propafenone) antiarrhythmic medications.
Endocrine and Metabolism
A reduction in glucose tolerance and/or glycated hemoglobin (HbAlc) has been observed in males receiving combined androgen blockade. This may manifest as diabetes or loss of glycemic control in those with pre-existing diabetes. Consideration should therefore be given to monitoring blood glucose and/or glycated hemoglobin (HbAlc) in patients receiving EUFLEX® in combination with LHRH analogues.
Hepatic Injury
There have been postmarketing reports of hospitalization and rarely death due to liver failure in patients taking flutamide. Evidence of hepatic injury included elevated serum transaminase levels, jaundice, hepatic encephalopathy, and death related to acute hepatic failure. The hepatic injury was reversible after prompt discontinuation of therapy in some patients. Approximately half of the reported cases occurred within the initial 3 months of treatment with flutamide.
Serum transaminase levels should be measured prior to starting treatment with flutamide. Flutamide is not recommended in patients whose ALT values exceed twice the upper limit of normal. Serum transaminase levels should then be measured monthly for the first 4 months of therapy, and periodically thereafter. Liver function tests also should be obtained at the first signs and symptoms suggestive of liver dysfunction, e.g. nausea, vomiting, abdominal pain,
fatigue, anorexia, “flu-like” symptoms, hyperbilirubinuria, jaundice, or right upper quadrant tenderness. If at any time a patient has jaundice, or their ALT rises above 2 times the upper limit of normal, flutamide should be immediately discontinued with close follow-up of liver function tests until resolution.
Hematologic
Anemia is a known physiologic consequence of testosterone suppression. Assessment of anemia risk and management according to local clinical practice and guidelines should be considered.
Monitoring and Laboratory Tests
Regular assessments of serum Prostate Specific Antigen (PSA) may be helpful in monitoring patients' response.
Anemia has been observed in patients treated with EUFLEX®. Hemoglobin levels should be monitored.
Assessment of cardiovascular risk factors, monitoring for signs and symptoms suggestive of development of cardiovascular disease, and management according to local clinical practice and guidelines should be considered. Monitoring of ECG and serum electrolyte levels during treatment should also be considered for those at risk for electrolyte abnormality and QTc prolongation.
Serum transaminase levels should be measured prior to starting treatment with flutamide, then monthly for the first 4 months of therapy, and periodically thereafter. Liver function tests also should be obtained at the first signs and symptoms suggestive of liver dysfunction.
Consideration should be given to monitoring blood glucose and/or glycated hemoglobin (HbAlc) in patients receiving EUFLEX® in combination with LHRH analogues.
Assessment of osteoporosis risk and management according to clinical practice and guidelines should be considered.
Musculoskeletal
Changes in Bone Density
EUFLEX® is indicated for use in combination with an LHRH analogue or orchiectomy. Based on studies conducted in the literature, decreased bone mineral density can be anticipated with long term combined androgen blockade with an anti-androgen plus LHRH analogue. Combined androgen blockade is associated with increased risks of osteoporosis and skeletal bone fractures. The risk of skeletal fracture increases with the duration of combined androgen blockade. Assessment of osteoporosis risk and management according to clinical practice and guidelines should be considered.
In patients with significant risk factors for decreased bone mineral content and/or bone mass such as chronic alcohol and/or tobacco use, presumed or strong family history of
osteoporosis or chronic use of drugs that can reduce bone mass such as anticonvulsants or corticosteroids, combined androgen blockade may pose an additional risk. In these patients, risk versus benefit must be weighed carefully before therapy is instituted.
Use in Pregnancy and Lactation
No studies have been conducted in pregnant or lactating women. Therefore, the possibility that EUFLEX® may cause fetal harm if administered to a pregnant woman, or may be present in the breast milk of lactating women must be considered.
There was decreased 24-hour survival in the offspring of rats treated with flutamide at doses of 30, 100, or 200 mg/kg/day (approximately 3, 9, and 19 times the human dose) during pregnancy. A slight increase in minor variations in the development of the sternebra and vertebra was seen in fetuses of rats at the two higher doses. Feminization of the males also occurred at the two higher dose levels. There was a decreased survival rate in the offspring of rabbits receiving the highest dose (15 mg/kg/day; equal to 1.4 times the human dose).
PRECAUTIONS
Periodic liver function tests and sperm count determinations must be performed in patients on long-term treatment with flutamide.
After long-term administration in rats, flutamide produced testicular interstitial cell adenomas and dose-related increases in mammary gland adenomas or carcinomas. The relevance of these findings to humans is unknown. It should be noted that few cases of malignant breast neoplasms have been reported in male patients receiving EUFLEX® (flutamide); causality has not been established.
Since flutamide tends to elevate plasma testosterone and estradiol levels, fluid retention may occur. Accordingly, flutamide should be used with caution in those patients with cardiac disease.
Hepatic Injury: Treatment with EUFLEX® should not be initiated in patients with serum transaminase levels exceeding 2 to 3 times the upper limit of normal.
Since transaminase abnormalities, cholestatic jaundice, hepatic necrosis and hepatic encephalopathy have been reported with the use of flutamide, periodic liver function tests must be performed in all patients.
Appropriate laboratory testing should be done monthly for the first 4 months, and periodically thereafter and at the first symptom/sign of liver dysfunction (e.g., pruritus, dark urine, persistent anorexia, jaundice, right upper quadrant tenderness or unexplained "flu-like" symptoms).
If the patient has laboratory evidence of liver injury or jaundice, in the absence of biopsy-confirmed liver metastases, flutamide therapy should be discontinued if the patient develops
jaundice or if serum transaminase levels rise to 2 to 3 times the upper limit of normal, even in clinically asymptomatic patients.
The hepatic injury is usually reversible after discontinuation of therapy and in some patients, after dosage reduction. However, there have been reports of death following severe hepatic injury associated with the use of flutamide.
Drug interactions: Interactions between EUFLEX® Tablets and leuprolide have not occurred. In patients receiving long-term oral-anticoagulant therapy, increases in prothrombin time have been reported after flutamide monotherapy was initiated. Therefore close monitoring of prothrombin time is recommended and adjustment of the anticoagulant dose may be necessary when flutamide tablets are administered concomitantly.
Cases of increased theophylline plasma concentrations have been reported in patients receiving concomitant theophylline and flutamide tablets. Theophylline is primarily metabolized by CYP 1A2, which is the primary enzyme responsible for the conversion of flutamide to its active agent 2-hydroxyflutamide.
Information for Patients: Patients should be informed prior to initiating this medication, of the possibility of its causing hepatic dysfunction. Instruct the patient to consult the doctor immediately if symptoms of hepatic dysfunction appear. These include itching of the skin, dark urine (amber or yellow-green urine is not a cause of concern), nausea, vomiting, persistent lack of appetite, yellow eyes or skin, tenderness in the right upper abdomen, or “flu-like” symptoms.
EUFLEX® (flutamide) is indicated only for use in male patients.
Patients should be informed that EUFLEX® and the drug used for medical castration should be administered concomitantly, and that they should not interrupt their dosing or stop taking these medications without consulting their physician.
ADVERSE REACTIONS
The most frequently reported adverse reactions to EUFLEX® (flutamide) Tablets monotherapy are gynecomastia and/or breast tenderness, sometimes accompanied by galactorrhoea. These reactions disappear upon discontinuation of treatment or reduction in dosage. The incidence of gynecomastia is reduced greatly when EUFLEX® Tablets are administered concomitantly with an LHRH agonist.
The most frequently reported (greater than 5%) adverse experiences during treatment with EUFLEX® Tablets in combination with a LHRH agonist are listed in the table below. For comparison, adverse experiences seen with a LHRH agonist and placebo are also listed in the following table.
(n=294) (n=285)
Flutamide + Placebo +
LHRH-agonist LHRH-agonist
% All % All
Hot Flashes 61 57
Loss of Libido 36 31
Impotence 33 29
Diarrhea 12 4
Nausea/Vomiting 11 10
Gynecomastia 9 11
Other 7 9
Other Gastro-Intestinal 6 4
As shown in the table, for both treatment groups, the most frequently occurring adverse experiences (hot flashes, loss of libido, impotence) were those known to be associated with low serum androgen levels and known to occur with LHRH-agonists alone.
The only notable difference between these treatment groups was the higher incidence of diarrhea in the flutamide + LHRH-agonist group (12%; severe in 5%) as compared to the placebo + LHRH-agonist group (4%; were severe in less than 1%).
In addition, the following adverse reactions were reported during treatment with flutamide +LHRH-agonist. No causal relatedness of these reactions to drug treatment has been made, and some of the adverse experiences reported are those that commonly occur in elderly patients.
Cardiovascular System: hypertension in 1% of patients. Rarely thrombophlebitis, pulmonary embolism, myocardial infarction.
Central Nervous System: CNS (drowsiness/confusion/depression/anxiety/nervousness) reactions occurred in 1% of patients. Rarely insomnia, tiredness, headache, dizziness, weakness, malaise, blurred vision and decreased libido have been reported.
Endocrine System: gynecomastia in 9% of patients. Rarely breast tenderness sometimes accompanied by galactorrhoea.
Gastrointestinal System: nausea/vomiting occurred in 11%; diarrhea 12%, anorexia 4%, and other gastro-intestinal disorders occurred in 6% of patients. Increased appetite, indigestion and constipation have also been reported.
Hematopoietic System: anaemia occurred in 6% of patients, leukopenia 3%, thrombocytopenia 1%.
Liver and Biliary System: clinically evident hepatitis and jaundice occurred in <1% of patients.
Skin: irritation at the injection site and rash occurred in 3% of patients. Photosensitivity reactions have been reported in five patients.
Other: Pruritus, ecchymosis, herpes zoster, thirst, lymphedema, lupus-like syndrome, hematuria, reduced sperm counts have been reported rarely in long-term treatment. Edema occurred in 4% of patients; neuromuscular, genitourinary symptoms occurred in 2% of patients. Interstitial lung disease occurred in <1% of patients.
Additional Adverse Experiences: In addition, the following adverse experiences have been reported during world-wide marketing of EUFLEX® Tablets: hemolytic anemia, macrocytic anemia, methemoglobinemia, sulfhemoglobinemia, photosensitivity reactions -- including erythema, ulcerations, bullous eruptions, and epidermal necrolysis -- and change in urine color to an amber or yellow-green appearance, which can be attributed to flutamide and/or its metabolites. Also observed were cholestatic jaundice, hepatic encephalopathy and hepatic necrosis. The hepatic conditions were usually reversible after discontinuing therapy; however, there have been reports of death following severe hepatic injury associated with use of flutamide. Cardiac failure, sudden cardiac death have been reported. Hyperglycemia and aggravated diabetes mellitus have been reported very rarely.
Two reports of malignant male breast neoplasms in patients being dosed with EUFLEX® have been reported. One involved aggravation of a pre-existing nodule which was first detected three to four months before initiation of EUFLEX® monotherapy in a patient with benign prostatic hypertrophy. After excision, this was diagnosed as a poorly differentiated ductal carcinoma. The other report involved gynecomastia and a nodule noted two and six months respectively, after initiation of EUFLEX® monotherapy for treatment of advanced prostatic carcinoma. Nine months after the initiation of therapy, the nodule was excised and diagnosed as a moderately differentiated invasive ductal tumor staged T4N0M0, G3, no metastases had advanced.
Laboratory Values: Reported abnormal laboratory test results include elevated SGOT (AST), SGPT (ALT); elevated blood urea nitrogen (BUN) and bilirubin levels; less frequently, elevated serum creatinine levels and elevated gamma-glutamyl transferase levels have been reported.
DRUG INTERACTIONS
Drug-Drug Interactions
EUFLEX® is indicated for use in combination with an LHRH analogue or orchiectomy. The
potential for QT/QTc prolongation has not been studied with EUFLEX®. Since combined androgen blockade prolongs the QTc interval, the concomitant use of EUFLEX® with medicinal products known to prolong the QTc interval or medicinal products able to induce torsades de pointes should be carefully evaluated. Such medicinal products include but are not limited to the examples that follow: Class IA (e.g. quinidine, disopyramide), Class III (e.g. amiodarone, sotalol, dofetilide, ibutilide, dronedarone), or Class IC (e.g. flecainide, propafenone) antiarrhythmic medicinal products, antipsychotics (e.g. chlorpromazine), antidepressants (e.g. amitriptyline, nortriptyline), opioids (e.g. methadone), macrolide antibiotics and analogues (e.g. erythromycin, clarithromycin, azithromycin), quinolone antibiotics (e.g. moxifloxacin), antimalarials (e.g. quinine), azole antifungals, 5-hydroxytryptamine (5-HT3) receptor antagonists (e.g. ondansetron), and beta-2 adrenoceptor agonists (e.g. salbutamol).
SYMPTOMS AND TREATMENT OF OVERDOSE
In animal studies with flutamide alone, signs of overdose included hypoactivity, piloerection, slow respiration, ataxia, and/or lacrimation, anorexia, tranquillization, emesis and methemoglobinemia.
Clinical trials have been conducted with flutamide in doses up to 1500 mg per day for periods up to 36 weeks with no serious adverse effects reported. Those adverse reactions reported included gynecomastia, breast tenderness and some increases in SGOT. The single dose of flutamide ordinarily associated with symptoms of overdose or considered to be life-threatening has not been established.
Since flutamide is highly protein bound, dialysis may not be of any use as treatment for overdose. As in the management of overdosage with any drug, it should be borne in mind that multiple agents may have been taken. Gastric lavage may be considered. General supportive care, including frequent monitoring of the vital signs and close observation of the patient, is indicated.
DOSAGE AND ADMINISTRATION
The recommended dosage of EUFLEX® (flutamide) in combination with orchiectomy or in combination with an LHRH agonist is one 250 mg tablet three times a day at eight-hour intervals. In combination with an LHRH agonist, either the two agents may be initiated simultaneously, or EUFLEX® tablet therapy may be started 24 hours prior to initiation of the LHRH agonist.
In the management of bulky locally advanced Stage B2 and Stage C prostatic carcinoma, the recommended dosage is one 250 mg tablet, three times a day at eight-hour intervals. EUFLEX® should be started simultaneously or 24 hours prior to initiation of the LHRH
agonist. Administration of EUFLEX® should begin eight weeks prior to external beam radiation therapy and continue through the course of radiation therapy. 
PHARMACEUTICAL INFORMATION
Flutamide:
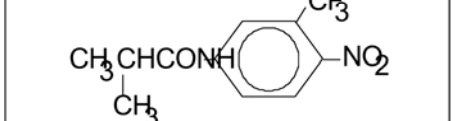
Molecular Formula: C11H11N2O3F3
Molecular Weight: 276.21
Chemical Name: 2-methyl-N-(4-nitro-3-(trifluoromethyl) phenyl) propanamide.
Description: Buff to yellow powder.
AVAILABILITY OF DOSAGE FORMS
Each round, biconvex, pale yellow, compressed tablet, engraved with "EUFLEX" on one face, and a single score on the other with the "SP" logo engraved on each side of the score line, contains: flutamide 250 mg. Nonmedicinal ingredients: cellulose, lactose, magnesium stearate, pregelatinized starch, silicon dioxide and sodium lauryl sulfate. Tartrazine-free. Bottles of 100. Store between 15° and 30°C. Protect from light and excessive moisture.