


Product Monograph - Bryhali
1 INDICATIONS
BRYHALI (halobetasol propionate) is a high to super-high potency topical corticosteroid indicated for corticosteroid-responsive dermatoses and the topical treatment of plaque psoriasis.
1.1 Pediatrics
Pediatrics (< 18 years of age): No data are available to Health Canada; therefore, Health Canada has not authorized an indication for pediatric use.
1.2 Geriatrics
Geriatrics (≥ 65 years of age): A limited number of subjects aged ≥ 65 years have been treated with BRYHALI in clinical trials, therefore the safety and efficacy have not been established in this patient population (see 7.1.4 Geriatrics).
2 CONTRAINDICATIONS
BRYHALI is contraindicated in patients who are hypersensitive to this drug or to any ingredient in the formulation, including any non-medicinal ingredient. For a complete listing, see 6 DOSAGE FORMS, STRENGTHS, COMPOSITION AND PACKAGING.
BRYHALI is also contraindicated:
- in patients who are hypersensitive to other corticosteroids.
- in viral diseases of the skin including herpes simplex, vaccinia and varicella.
- in untreated bacterial, tubercular and fungal infections involving the skin.
4 DOSAGE AND ADMINISTRATION
4.1 Dosing Considerations
BRYHALI is for dermatological use only.
4.2 Recommended Dose and Dosage Adjustment
Health Canada has not authorized an indication for pediatric use (see 1.1 Pediatrics and 7.1.3 Pediatrics).
BRYHALI should not be used in children. Because of the higher ratio of skin surface area to body mass, children are at greater risk for hypothalamic pituitary adrenal (HPA) axis suppression, glucocorticoid insufficiency after withdrawal of treatment and Cushing's syndrome while on treatment.
The total dosage of BRYHALI should not exceed approximately 50 g per week because of the potential for the drug to suppress the hypothalamic-pituitary-adrenal (HPA) axis.
Apply a thin layer of BRYHALI once daily to cover only affected areas and rub in gently. If a bath or shower is taken prior to application, the skin should be dry before applying the lotion. Do not use occlusive dressing, unless recommended by a physician.
Periodic evaluation for evidence of HPA axis suppression is recommended, particularly in patients who are using BRYHALI uninterrupted for up to 8 weeks (see 7 WARNINGS
AND PRECAUTIONS).
Treatment should be discontinued when control has been achieved. BRYHALI uninterrupted treatment beyond 8 weeks is not recommended. If no improvement is seen within 8 weeks of treatment, reassessment of the diagnosis may be necessary.
The efficacy and safety of BRYHALI in patients with more than 12% of body surface area affected by plaque psoriasis has not been established (see 14 CLINICAL TRIALS).
4.3 Reconstitution
This information is not available for this drug product.
4.4 Administration
BRYHALI should not be used with occlusive dressings. BRYHALI is for external use only.
Keep away from the eyes, nose, mouth, and other mucous membranes. In the event of contact with the eye, flush with cold water.
BRYHALI should not be used on the face, scalp, groin, or in the axillae.
5 OVERDOSAGE
Topically applied BRYHALI can be absorbed in sufficient amounts to produce systemic effects including reversible HPA axis suppression with the potential for glucocorticosteroid insufficiency after withdrawal of treatment. If HPA axis suppression is noted, withdraw the drug gradually by reducing the amount and frequency of application. Recovery of HPA axis function is generally prompt and complete upon discontinuation of topical corticosteroids. Infrequently, signs and symptoms of glucocorticosteroid insufficiency may occur requiring supplemental systemic corticosteroids.
| For management of a suspected drug overdose, contact your regional poison control centre. |
|---|
6 DOSAGE FORMS, STRENGTHS, COMPOSITION AND PACKAGING
Table – Dosage Forms, Strengths, Composition and Packaging.
*Does not include any fragrance, colorant or alcohol
BRYHALI is a white to off-white, lightweight, non-greasy lotion supplied 45 g, 60 g and 100 g aluminium tubes providing 0.1 mg of halobetasol per gram. Physicians’ samples are supplied in 3 g white aluminum tubes. It is provided in a formulation with known hydrating and moisturizers such as diethyl sebacate, light mineral oil and sorbitol solution which may alleviate dryness of skin.
7 WARNINGS AND PRECAUTIONS
General
Significant systemic absorption may occur when steroids are applied over large areas of the body. To minimize this possibility, when long term therapy is anticipated, interrupt treatment periodically or treat one area of the body at a time.
Laboratory Tests: Patients receiving a large dose of a high potency topical steroid applied to a large surface area should be evaluated periodically for evidence of HPA axis suppression. This may be done by using the ACTH stimulation, A.M. plasma cortisol and urinary free-cortisol tests. Patients receiving super-potent corticosteroids should not be treated for more than 8 weeks at a time and it is recommended that only small areas be treated at any one time due to the increased risk of HPA suppression.
Carcinogenesis and Mutagenesis
Long-term animal studies have not been performed to evaluate the carcinogenic potential of halobetasol propionate. Positive mutagenicity studies were observed in two genotoxicity assays. Halobetasol was positive in a Chinese hamster micronucleus test in vivo and in a mouse lymphoma gene mutation assay in vitro. In other genotoxicity tests including Ames/Salmonella assay, sister chromatid exchange test, chromosome aberration studies of germinal and somatic cells of rodents and in a mammalian spot test for point mutations, halobetasol propionate was not found to be genotoxic.
Endocrine and Metabolism
Halobetasol has been shown to suppress the HPA axis. Systemic effects of topical corticosteroids may include reversible HPA axis suppression with the potential for glucocorticosteroid insufficiency. This may occur during treatment or upon withdrawal of treatment with the topical corticosteroid.
| Route of Administration | Dosage Form / Strength/Composition | Non-medicinal Ingredients* |
| Topical | Lotion 0.01 % w/w | Carbomer Copolymer Type B, Carbomer Homopolymer Type A, Diethyl Sebacate, Edetate Disodium Dihydrate, Light Mineral Oil, Methylparaben, Propylparaben, Purified Water, Sodium Hydroxide, Sorbitan Monooleate and Sorbitol Solution, 70%. |
The potential for HPA axis suppression with BRYHALI was evaluated in a study of 19 adult subjects with moderate to severe plaque psoriasis involving ≥20% of their body surface area (BSA). HPA axis suppression was reported for 1 (5.6%) subject at Week 4 and for 3 (15.8%) subjects at Week 8. All 3 subjects had normal HPA axis suppression test with discontinuation of treatment (see 14 CLINICAL TRIALS).
Because of the potential for systemic absorption, use of topical corticosteroids, including BRYHALI, may require that patients be evaluated periodically for evidence of HPA axis suppression. Factors that predispose a patient using a topical corticosteroid to HPA axis suppression include the use of more potent corticosteroids, use over large surface areas, occlusive use, use on an altered skin barrier, concomitant use of multiple corticosteroid-containing products, liver failure, and young age. An adrenocorticotropic hormone (ACTH) stimulation test may be helpful in evaluating patients for HPA axis suppression.
If HPA axis suppression is documented, attempt to gradually withdraw the drug, reduce the frequency of application, or substitute a less potent steroid. Manifestations of adrenal insufficiency may require supplemental systemic corticosteroids. Recovery of HPA axis function is generally prompt and complete upon discontinuation of topical corticosteroids.
Systemic effects of topical corticosteroids may also include Cushing’s syndrome, hyperglycemia, and glucosuria. Use of more than one corticosteroid-containing product at the same time may increase the total systemic exposure to corticosteroids. Pediatric patients may be more susceptible than adults to systemic toxicity from the use of topical corticosteroids due to their larger surface-to-body mass ratios (see 7.1.3 Pediatrics).
Hepatic
There are no adequate and well controlled studies of BRYHALI use in patients with hepatic impairment. As corticosteroids undergo hepatic metabolism, BRYHALI should be used with caution in patients with hepatic impairment.
Immune
Allergic contact dermatitis with corticosteroids is usually diagnosed by observing failure to heal rather than noting a clinical exacerbation. Consider confirmation of a clinical diagnosis of allergic contact dermatitis by appropriate patch testing. Discontinue BRYHALI if allergic contact dermatitis occurs.
Ophthalmic
The product is not formulated for ophthalmic use and should not be used in or near the eyes.
Use of topical corticosteroids may increase the risk of posterior subcapsular cataracts and glaucoma. Cataracts and glaucoma have been reported in post-marketing experience with the use of topical corticosteroid products. Advise patients to report any visual symptoms and consider referral to an ophthalmologist for evaluation.
Sexual Health
Reproduction/Fertility
There are no data on the effects of halobetasol propionate on human fertility (see section 16 NON-CLINICAL TOXICOLOGY for fertility studies in animals).
Skin
Prolonged use of topical corticosteroid products may produce atrophy of the skin and subcutaneous tissues. If this occurs, treatment should be discontinued.
Topical corticosteroids should be used with caution in patients with stasis dermatitis and other skin diseases associated with impaired circulation, hypersensitive patients and patients with glaucoma.
Use an appropriate antimicrobial agent if a skin infection is present or develops. If a favorable response does not occur promptly, discontinue use of BRYHALI until the infection has been adequately treated.
7.1 Special Populations
7.1.1 Pregnant Women
There are no available data on BRYHALI use in pregnant women to inform a drug-associated risk of major birth defects, miscarriage, or adverse maternal or fetal outcomes. Therefore, BRYHALI use in pregnant women is not recommended. If the patient becomes pregnant while using this drug, treatment should be discontinued.
Animal Data
Corticosteroids have been shown to be teratogenic and embryotoxic in laboratory animals at low doses when administered systemically. Some corticosteroids have been shown to be teratogenic after topical application. Halobetasol propionate has been shown to cause malformations in rats and rabbits when given orally during organogenesis at doses of 0.04 to 0.1 mg/kg/day in rats and 0.01 mg/kg/day in rabbits. Halobetasol propionate was embryotoxic in rabbits but not in rats. Cleft palate was observed in both rats and rabbits. Omphalocele was seen in rats but not in rabbits. The human topical dose of BRYHALI was embryotoxic in rabbits.
7.1.2 Breast-feeding
There are no data on the presence of halobetasol propionate or its metabolites in human milk, the effects on the breastfed infant, or the effects on milk production after treatment with BRYHALI.
Systemically administered corticosteroids appear in human milk and could suppress growth, interfere with endogenous corticosteroid production, or cause other untoward effects. It is not known whether topical administration of corticosteroids could result in sufficient systemic absorption to produce detectable quantities in human milk.
The developmental and health benefits of breastfeeding should be considered along with the mother’s clinical need for BRYHALI and any potential adverse effects on the breastfed child from BRYHALI.
Advise breastfeeding women not to apply BRYHALI directly to the nipple and areola to avoid direct infant exposure.
7.1.3 Pediatrics
Safety and effectiveness of BRYHALI in pediatric patients under the age of 18 years have not been evaluated.
Because of higher skin surface area to body mass ratios, pediatric patients are at a greater risk than adults of HPA axis suppression and Cushing’s syndrome when they are treated with topical corticosteroids. They are therefore also at greater risk of adrenal insufficiency during or after withdrawal of treatment. Adverse reactions including striae have been reported with use of topical corticosteroids in infants and children (see 7 WARNINGS AND PRECAUTIONS, Endocrine and Metabolism).
HPA axis suppression, Cushing’s syndrome, linear growth retardation, delayed weight gain, and intracranial hypertension have been reported in children receiving topical corticosteroids. Manifestations of adrenal suppression in children include low plasma cortisol levels and an absence of response to ACTH stimulation. Manifestations of intracranial hypertension include bulging fontanelles, headaches, and bilateral papilledema (see 7 WARNINGS AND PRECAUTIONS, Endocrine and Metabolism).
7.1.4 Geriatrics
A limited number of subjects aged ≥ 65 years have been treated with BRYHALI in clinical trials, therefore the safety and efficacy have not been established in this patient population.
8 ADVERSE EVENTS
8.1 Adverse Reaction Overview
Local adverse reactions from topical corticosteroids may include atrophy, striae, telangiectasias, burning, itching, irritation, dryness, folliculitis, acneiform eruptions, hypopigmentation, perioral dermatitis, allergic contact dermatitis, secondary infection, and miliaria. These may be more likely with occlusive use, prolonged use, or use of higher potency corticosteroids, including BRYHALI. Some local adverse reactions may be irreversible.
8.2 Clinical Trial Adverse Reactions
Because clinical trials are conducted under very specific conditions, the adverse reaction rates observed in the clinical trials may not reflect the rates observed in practice and should not be compared to the rates in the clinical trials of another drug. Adverse reaction information from clinical trials is useful for identifying drug-related adverse events and for approximating rates.
In randomized, double-blind, multicenter, vehicle-controlled clinical trials, 426 adults with plaque psoriasis were treated with BRYHALI and had post-baseline safety data. Subjects applied BRYHALI once daily for up to eight weeks. Table 1 presents adverse events that occurred in at least 1% of subjects treated with BRYHALI and more frequently than in vehicle-treated subjects.
Table 1: Adverse Events Occurring in ≥1% of the Subjects Treated with BRYHALI through Week 8
BRYHALI (N=284) | Vehicle (N=142) | |
Infections and infestations Upper Respiratory Tract Infection Gastroenteritis | 2.5% 1.1% | 1.4%
|
Respiratory, thoracic and mediastinal disorders Cough | 1.1% | 0.7% |
Vascular disorders Hypertension | 1.4% | 0.7% |
8.2.1 Clinical Trial Adverse Reactions (Pediatrics)
No clinical trials were carried out in pediatric population.
8.3 Less Common Clinical Trial Adverse Reactions (<1%)
General disorders and administration site conditions: Application site dermatitis, application site discolouration, application site pruritus.
Infections and infestations: Application site infection.
8.4 Abnormal Laboratory Findings: Hematologic, Clinical Chemistry and Other Quantitative Data
There were no findings related to hematology or chemistry parameters that appeared to be treatment-related in any of the studies that included subjects with plaque psoriasis.
9 DRUG INTERACTIONS
9.2 Drug Interactions Overview
No formal drug-drug interaction studies were conducted with BRYHALI.
10 ACTION AND CLINICAL PHARMACOLOGY
10.1 Mechanism of Action
Halobetasol propionate belongs to the superpotent class of topical corticosteroids. Like other topical corticosteroids, halobetasol propionate has anti-inflammatory, antipruritic and vasoconstrictive actions. The mechanism of the anti-inflammatory activity of the topical corticosteroids, in general, is unclear. However, corticosteroids are thought to act by the induction of phospholipase A2 inhibitory proteins, collectively called lipocortins. It is postulated that these proteins control the biosynthesis of potent mediators of inflammation such as prostaglandins and leukotrienes by inhibiting the release of their common precursor arachidonic acid. Arachidonic acid is released from membrane phospholipids by phospholipase A2.
10.2 Pharmacodynamics
A vasoconstrictor assay in healthy subjects with BRYHALI indicated that the formulation is in the potent to superpotent range of potency as compared to other topical corticosteroids.
BRYHALI has been shown to suppress the HPA axis.
HPA axis suppression was evaluated in Study 501 after repeated once daily treatment of the final to be marketed formulation of BRYHALI for 57 days. The incidence of HPA axis suppression was low, which included 1 (5.6%) subject in on Day 29 and 3 (15.8%) subjects on Day 57 following BRYHALI treatment. All subjects had normal HPA axis suppression tests at the unscheduled follow up visit.
10.3 Pharmacokinetics
The maximum concentrations of and exposure to halobetasol propionate after administrations of halobetasol propionate lotion were similar between Days 14-15 and 28-29. Mean Cmax values for halobetasol propionate were 8.60, 31.2, and 28.7 pg/mL on Days 1-2, Days 14-15, and Days 28-29, respectively. Mean AUC (0 t) values for halobetasol propionate were 678, 2160, and 2060 pg*hr/mL on Days 1-2, Days 14-15, and Days 28-29, respectively. Median Tmax values were 4.95 hours, 4.83 hours, and 5.55 hours for Days 1-2, Days 14-15, and Days 28-29, respectively. Mean (SD) Cmin values were 0 pg/mL, 6.55 (29.3) pg/mL, and 5.37 (23.4) pg/mL (29.3) pg/mL for Days 1--2, Days 14--15, and Days 28--29, 29, respectively.
Mean (SD) AUC (0 t) values were 678 pg*hr/mL, 2160 (2440) pg*hr/mL, and 2060 (2760) pg*hr/mL for Days 1-2, Days 14-15, and Days 28-29, respectively. All of these values should be interpreted with caution, however, since these values were calculated from only 1 subject on Days 1-2, and only 2 subjects on each of Days 14-15 and Days 28-29. No value for
AUC (0-24h) could be calculated on either of Days 1-2 or Days 28-29 due to the low exposure and high number of values below the limit of quantification (BLQ). An AUC (0 24h) value of 3890 pg*hr/mL was calculated for Days 14-15; however, this value was based on 1 subject. The calculated mean AUC values do not reflect the mean concentration time curve profiles since these plots included zero values in the mean and standard errors and since AUCs were determined only for subjects with three consecutives measurable (not BLQ) HP concentrations.
Pharmacokinetics parameters for halobetasol propionate after administration the lotion are summarized in Table 2.
Table 2: Summary of HP Pharmacokinetic Parameters for BRYHALI
| Parameter | N | Days 1-2 Mean (SD) | N | Days 14-5 Mean (SD) | N | Days 28-29 Mean (SD) |
| Cmax (pg/mL) | 21 | 8.60 (28.5) | 20 | 31.2 (62.2) | 19 | 28.7 (71.2) |
| Cmin (pg/mL) | 21 | 0 (0) | 20 | 6.55 (29.3) | 19 | 5.37 (23.4) |
| Tmax (hr) | 2 | 4.95 (4.17) | 5 | 4.83 (5.01) | 5 | 5.55 (4.36) |
| AUC(0-t) (pg*hr/mL) | 1 | 678 (NA) | 2 | 2160 (2440) | 2 | 2060 (2760) |
| AUC(0-24h) (pg*hr/mL) | 0 | - | 1 | 3890 (NA) | 0 | 0 |
SD = Standard deviation
11 STORAGE, STABILITY AND DISPOSAL
Store at controlled room temperature between 15°C and 30°C. Protect from freezing. The in-use period is 8 weeks. Stability data beyond 8 weeks after opening is not available.
12 SPECIAL HANDLING INSTRUCTIONS
This information is not available for this drug product.
PART II: SCIENTIFIC INFORMATION
13 PHARMACEUTICAL INFORMATION
BRYHALI contains halobetasol propionate as the active ingredient in a white to off-white, lightweight, non-greasy lotion formulation intended for topical use. The BRYHALI formulation uses a polymeric emulsification system (PRISMATREXTM) which maintains the emulsion droplets size distribution stable across time and temperature. This system provides stable emulsions by anchoring its hydrophobic portions and forming an adsorbed gel layer around each oil droplet. The product’s target pH (5.0 – 6.0) is controlled by the amount of polymer and base (sodium hydroxide) present in the formulation. This pH range is consistent with the pH tolerated by the skin without inducing irritation.
Drug Substance
Proper name: Halobetasol Propionate
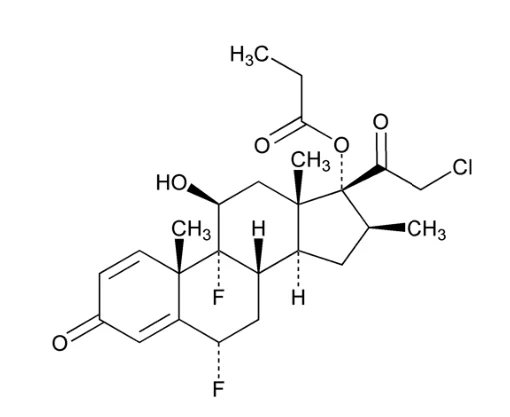
Chemical name: 21-Chloro-6α,9-difluoro-11β-hydroxy-16β-methyl-pregna-1,4-diene-3-20-dione,17-propionate
Molecular formula: C25H31ClF2O5
Molecular mass: 484.96 g/mol
Structural formula:
Physicochemical properties
Description: Halobetasol propionate is a white crystalline powder or solid
Solubility: Insoluble in water, soluble in ethanol (37%), dimethyl sulfoxide (>50%), soluble in diesters (e.g., dibutyl adipate), slightly soluble in long-chain monoesters (isopropyl myristate).
Melting Point: The melting range is between 200°C and 216°C with decomposition.
14 CLINICAL TRIALS
14.1 Trial Design and Study Demographics
Table 3: Summary of patient demographics for clinical trials for BRYHALI
| Study # | Trial design | Dosage, route of administration and duration | Study subjects (n) | Mean age (Range) | Sex |
| Trial 1 | A Phase 3, Multicenter, Double-Blind, Randomized, Vehicle Controlled Clinical Study to Assess the Safety and Efficacy of BRYHALI in the Treatment of Plaque Psoriasis | BRYHALI (HP 0.01%) Reference Therapy: Vehicle Lotion Test product and reference therapy were applied topically, once daily for 8 weeks. | BRYHALI: 143 subjects Vehicle: 74 subjects | BRYHALI: (20 - 88) Vehicle: (20 – 85) | BRYHALI: 85 M/58 F Vehicle: 39 M/35 F |
| Trial 2 | A Phase 3, Multicenter, Double-Blind, Randomized, Vehicle Controlled Clinical Study to Assess the Safety and Efficacy of BRYHALI in the Treatment of Plaque Psoriasis | BRYHALI (HP 0.01%) Reference Therapy: Vehicle Lotion Test product and reference therapy were applied topically, once daily for 8 weeks. | BRYHALI: 142 subjects Vehicle: 71 subjects | BRYHALI: (19 – 83) Vehicle: (22 – 80) | BRYHALI: 87 M/55 F Vehicle: 42 M/29 F |
| Study # | Trial design | Dosage, route of administration and duration | Study subjects (n) | Mean age (Range) | Sex |
| Trial 3 | A Phase 2, Multi-Center, Double-Blind, Randomized, Vehicle-Controlled Study to Compare the Safety and Efficacy of BRYHALI to HP cream, 0.05% in the Treatment of Plaque Psoriasis | Test Product: BRYHALI (HP 0.01%) Reference Therapy: (HP) Cream, 0.05% Vehicle Lotion Vehicle Cream The test product and reference therapy were applied topically, once daily for 2 weeks. | BRYHALI: 60 subjects HP Cream: 57 subjects Vehicle Lotion: 17 subjects Vehicle Cream: 16 subjects | BRYHALI: (18 – 76) HP Cream: (26 – 72) Vehicle Lotion: (30 – 60) Vehicle Cream: (23 – 72) | BRYHALI: 35 M/25 F HP Cream: 30 M/27 F Vehicle Lotion: 11 M/6 F Vehicle Cream: 4 M/12 F |
(Trial 1 = V01-122A-301; Trial 2 = V01-122A-302; Trial 3 = V01-122A-203)
Halobetasol was evaluated for the treatment of moderate to severe plaque psoriasis in two pivotal prospective, multicenter, randomized, double-blind clinical trials (Trial 1 and Trial 2). These trials were conducted in 430 subjects 18 years of age and older with moderate to severe plaque psoriasis that covered a body surface area (BSA) between 3% and 12% excluding the face, scalp, palms, soles, axillae, and intertriginous areas. Subjects applied the lotion or vehicle to all affected areas once daily for up to 8 weeks.
Efficacy was based primarily on results of the Investigator’s Global Assessment (IGA). The IGA is a scale that scores the overall areas affected with plaque psoriasis ranging from 0 to 4 defined as 0 (clear), 1 (almost clear), 2 (mild), 3 (moderate), and 4 (severe). In addition, evaluations included improvement in the signs of psoriasis (erythema, plaque elevation, and scaling), using scales that ranged from 0 (none) to 4 (severe). Assessments of improvements in the percent body surface area (BSA) affected by psoriasis also were conducted, and subjects completed the Dermatology Life Quality Index (DLQI) questionnaire, a skin disease-specific instrument validated for use in patients with psoriasis.
The Phase 2 study (Trial 3) included the same efficacy assessments as in the pivotal Phase 3 studies and used the same definition of treatment success (IGA). Two-grade improvements in each of the signs of psoriasis and changes to the percent BSA affected by psoriasis were evaluated. The study had a 2-week treatment duration, consistent with the recommended duration of use for the lotion.
14.2 1Study Results
The primary efficacy endpoint was the percent of subjects with treatment success at Week 8, where treatment success was defined as at least a 2-grade improvement from Baseline in IGA score and an IGA score equating to “Clear” or “Almost Clear.” Table 3 lists the primary efficacy results for Trials 1 and 2. For both trials treatment success in the BRYHALI treatment arm was statistically significant to vehicle (p<0.001). The secondary efficacy endpoints similarly evaluated the percent of subjects with treatment success but used a gated sequential analysis procedure to analyze the results at Weeks 12 (4-week follow-up), 6, 4, and 2. Figure 1 shows the secondary analysis of the primary efficacy endpoints.
Table 4: Primary Efficacy Results at Week 8 in Subjects with Moderate to Severe Plaque Psoriasis
| Trial 1 | Trial 2 | |||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| BRYHALI | Vehicle | BRYHALI | Vehicle | |||||||||||
IGA Treatment Success at Week 8* | N = 143 | N = 74 | N = 142 | N = 71 | ||||||||||
| 37 % | 8 % | 38 % | 12 % | |||||||||||
| P-Value | <0.001% | <0.001% | ||||||||||||
* Intent-to-Treat population. Treatment success was defined as at least a 2-grade improvement from baseline in IGA score and an IGA score equating to “clear” or “almost clear”. Clear = no evidence of scaling, no evidence of erythema, no evidence of plaque elevation above normal skin level. Almost clear = some plaques with fine scales, faint pink/light red erythema on most plaques, slight or barely perceptible elevation of plaques above normal skin level.
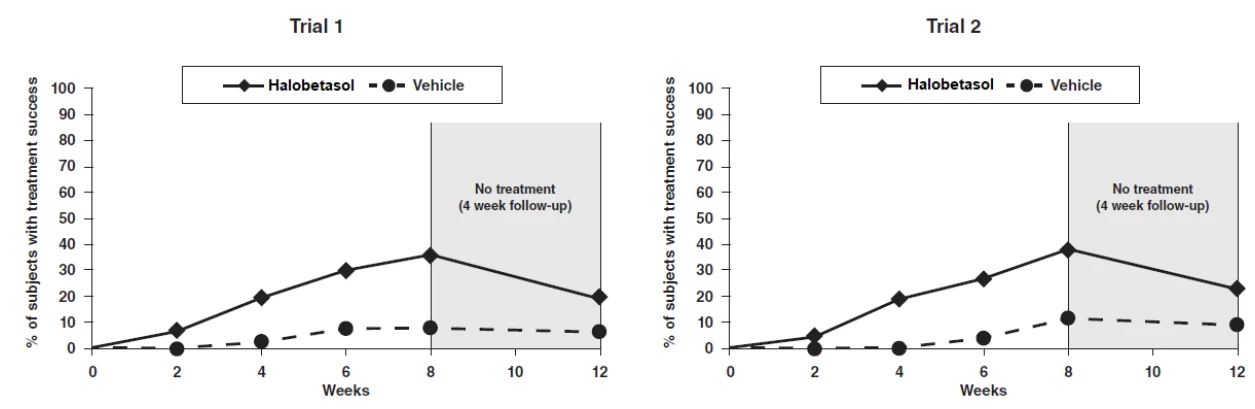 *The treatment difference at Week 2 in Trial 2 was not statistically significant.
Halobetasol = BRYHALI
Figure 1: Efficacy Results* over Time: IGA Treatment Success
15 MICROBIOLOGY
This information is not available for this drug product.
16 NON-CLINICAL TOXICOLOGY
General Toxicology
Carcinogenicity
Long-term animal studies have not been performed to evaluate the carcinogenic potential of halobetasol propionate.
Genotoxicity
Ames/Salmonella
Halobetasol propionate was tested for mutagenic potential in a screening Ames/Salmonella microsome plate assay. An epoxide hydratase inhibitor and glutathione depleter, 1,1,1-trichloropropene 2,3-oxide was also included in the assay in order to increase the sensitivity of the assay for mutagenic epoxide potential. The results of this study showed halobetasol propionate to be nonmutagenic to bacterial cells with or without metabolic activation under the conditions of this assay.
Nuclear Anomaly Test in Somatic Interphase Nuclei of Chinese Hamster
Halobetasol propionate was tested to evaluate the potential mutagenic effects on somatic interphase cells in the bone marrow of Chinese hamsters. Halobetasol propionate was administered at single oral dose levels of 750, 1500, 3000, and 6000 mg/kg daily for 2 consecutive days to study groups of 6 or 8/sex/group. The hamsters were sacrificed 24 hours after the second dose, and bone marrow smears were prepared. Control groups received the vehicle (0.5% CMC), and a positive control group was dosed with cyclophosphamide at 128 mg/kg. Nuclear anomalies were increased significantly in the bone marrow cells from animals dosed with the three lower doses of halobetasol propionate as compared to the number of nuclear anomalies in the controls. Under the experimental conditions, halobetasol propionate exerted a mutagenic action on hamster bone marrow somatic cells.
Another potent steroid, clobetasol propionate, in a similar study, was also found to produce nuclear anomalies in the bone marrow cells of Chinese hamsters at dose levels of 1250, 2500, and 5000 mg/kg.
Sister Chromatid Exchange
Halobetasol propionate was tested to evaluate the potential mutagenic effect on somatic cells (bone marrow) by the induction of sister chromatid exchange (SCE). Halobetasol propionate was administered as a single oral dose at levels of 1500, 3000, and 6000 mg/kg at a dose volume of 20 ml/kg in 0.5% CMC vehicle to Chinese hamster study groups of 4/sex/group. Two hours prior to dosing, the animals received a subcutaneous implant of a 45 mg tablet of 5-bromodeoxyuridine. After 24 hours, the hamsters were dosed i.p. with 10 mg/kg of colcemid and sacrificed, with bone marrow preparations made and stained for SCE evaluation. Control groups consisting of the vehicle and a positive control (100 mg/kg DMBA) were similarly evaluated. The results of this study showed no significant increase in the number of SCE found in comparison to the negative (vehicle) control.
Mouse Lymphoma
Halobetasol propionate was tested for mutagenic effects on L5178Y/TK +/- mouse lymphoma cells in vitro with and without microsomal activation. Results were expressed by the number of induced TK-/- mutants/106 surviving cells. Initially in the assay tested with microsomal activation, the two low dose levels of 8.125 and 16.25 mg/ml did not produce increased mutant frequencies. In the 3 upper dose levels tested, 32.5-130 mg/ml, increased mutant frequencies were observed. Similar effects occurred when the dose levels were reassayed in the cells. Mutation frequency values were elevated at all 5 dose levels tested in the presence of metabolic activation. Again, precipitation of the test article was observed after a 4-hour treatment period. Under the experimental conditions, halobetasol propionate exerted mutagenic activity in the mouse lymphoma forward mutation system with and without metabolic activation. Clobetasol propionate was subsequently evaluated in a similar mouse lymphoma test and did not show evidence of mutagenic effects. When tested soluble concentrations did not produce a marked increase in the mutant frequency compared with the control. Concentrations in excess of 56 mg/ml without microsomal activation proved to be cytotoxic to the mouse lymphoma cells. Mutation frequencies were, therefore, not determined at this concentration.
Chromosome Studies on the Male Germinal Epithelium/ Spermatocytes – Mouse
Halobetasol propionate was tested for mutagenic effect on the germinal epithelium, particularly on the potential formation of chromosomal aberrations in spermatocytes of mice. Halobetasol propionate was administered orally at daily dose levels of 333 and 1,000 mg/kg at a dose volume of 20 ml/kg in 0.5% CMC. Dosing was conducted intermittently for 5 days (days 0, 2, 3, 5, and 9) to study groups of 15 male mice/group including vehicle control. Three days later, the groups were dosed with 10 mg/kg of colcemid and were killed. Drop preparations were made of the testicular parenchyma with 100 metaphases scored per animal. Results from this study indicated no evidence of mutagenic activity of halobetasol propionate in mouse spermatocytes. There were no dose-related increases in the frequency of chromosomal aberrations, however, there was an occurrence of a quadrivalent exchange figure in the low dose. Mortalities occurred at both the low and high-dose groups.
Clobetasol propionate was subsequently evaluated in a comparative chromosome study in mouse spermatocytes and did not produce any chromosomal aberrations.
Chromosome Studies on Male Germinal Epithelium/ Spermatogonia – Mouse
Halobetasol propionate was tested for mutagenic effect on the germinal epithelium particularly on the potential formation of chromosomal aberrations in spermatogonia of mice. Halobetasol propionate was administered orally for 5 consecutive days at daily dose levels of 1,667 and 5,000 mg/kg in a dose volume of 20 ml/kg in 0.5% CMC to study groups of 12 males/group including a vehicle control group. The mice were sacrificed one day after the last dose after receiving 10 mg/kg of colcemid. Drop-preparations were made of the testicular parenchyma, with 100 metaphases scored per animal. The results from this assay revealed no mutagenic activity of halobetasol propionate in mouse spermatogonia.
In a similar study clobetasol propionate was subsequently evaluated for chromosomal aberrations in the spermatogonia of mice and did not produce any chromosomal aberrations.
Chromosome Studies on Somatic Cells - Chinese Hamster
Halobetasol propionate was tested to evaluate the potential mutagenic effects on somatic cells (bone marrow) in Chinese hamsters. Halobetasol propionate was administered orally as a single dose for 2 consecutive days at dose levels of 1250, 2500, and 5000 mg/kg in 20 ml/kg of 0.5% CMC to study groups of 4/sex/group. Control groups of the vehicle and positive control (cyclophosphamide) were similarly tested. Colcemid (10 mg/kg) was dosed 2 hours after the second dose to the study groups, and all animals were killed 4 hours later. Chromosomal preparations were made from the bone marrow, with 100 metaphase plates examined from 2/sex/group. Frequencies of chromosomal aberrations and aberrant metaphases were similar in the treated and negative control groups. Therefore, the results of this study indicated no evidence of mutagenic effects of halobetasol propionate in the somatic cells of Chinese hamsters.
In a parallel study clobetasol propionate was similarly evaluated in the Chinese hamster and did not show evidence of mutagenic effects.
Mammalian Spot Test – Mouse
Halobetasol propionate was tested to evaluate the potential mutagenic effects on somatic cells in vivo. The test permits the detection of induced point mutations and other genetic events in the melanoblasts of embryos exposed in utero to the test material. Mutation induction is monitored postnatally for the presence on the fur of young mice for recessive spots (RS). Halobetasol propionate was administered as a single i.p. dose at 6 levels ranging from 18.75 to 600 mg/kg in 10 ml/kg of sesame oil on the 10th day of pregnancy to study groups of 71-73 pregnant female C57B1/6J mice each. A vehicle control and positive control (50 mg/kg - N-nitroso-n-ethyl urea (EMA)) were similarly evaluated.
Post-natal examinations of the fur were recorded at the age of 12-14 days and twice weekly for 3 weeks for the presence of RS as well as for cytotoxic effects on melanocytes by recording of white mid-ventral spots (WMVS). In the 3 highest dose levels, a high percentage of mortality and embryotoxicity was observed. The results of this study revealed no evidence of mutagenic effects observed in the surviving offspring. Dose-related cytotoxic effects on melanocytes, as well as embryotoxic effects were observed.
Similar results were observed with clobetasol propionate in a subsequent mammalian spot test study in mice.
Micronucleus Test in Mice
Halobetasol propionate was evaluated in this in vivo micronucleus test to determine its potential to damage chromosomes of bone marrow cells or damage the mitotic spindle apparatus in these cells. Halobetasol propionate was administered to groups of mice (5 males and 5 females per group) by intraperitoneal injection for two consecutive days. Nominal halobetasol propionate doses were 7.5, 40 and 75 mg/kg per day. The animals were sacrificed at 24 and 48 hours after the second injection. Animals administered the negative (vehicle) control, i.e., DMSO and corn oil, were also sacrificed at 24 and 48 hours after the second injection. Positive control mice (triethylenemelamine at 0.5 mg/kg) were sacrificed only at 24 hours after the second dose administration.
Slides were prepared from the bone marrow of the femurs of each animal. The slides were subsequently stained, blind coded, and microscopically evaluated for the incidence of micronucleated polychromatic erythrocytes (PCE). Also scored was the incidence of normochromatic erythrocytes in order to estimate PCE/NCE ratios, a measure of toxicity to the hemopoietic system. Halobetasol propionate did not induce any statistically significant increases in the number of micronucleated PCEs. The mean PCE/NCE ratio for all halobetasol propionate treated groups was significantly reduced when compared to vehicle controls at both the 24- and 48-hours time points. The lower PCE/NCE ratios are indicative of toxicity to the bone marrow hemopoietic cells.
In conclusion, halobetasol propionate is negative in this in vivo mouse bone marrow micronucleus assay, when tested up to toxic dose levels.
CHO/HGPRT Mammalian Cell Forward Gene Mutation Assay
Halobetasol propionate was evaluated for mutagenic potential in this forward gene mutation assay. This assay, conducted in vitro in Chinese hamster ovary fibroblasts, measures the ability of a test article to induce a deletion, frame shift, or base pair substitution.
Halobetasol propionate was evaluated for a five-hour dosing period at nominal concentrations of 25, 50, 75, and 125 mcg/ml of treatment medium. All dose levels were evaluated with and without Aroclor 1254 induced rat liver S-9 activation. Mutant frequencies among the halobetasol propionate treated cultures were all at or below negative (vehicle, DMSO) control values for the three lower doses. The mutation frequency at 125 mcg /ml dose level was increased above vehicle and negative control values. However, this dose exceeded the limit of solubility in the treatment medium.
Clobetasol propionate was included in the study as a reference agent at nominal concentrations of 25, 50, 75, and 125 mcg/ml of treatment medium. Clobetasol propionate was evaluated only in the absence of S-9 metabolic activation. Mutant frequencies observed in the clobetasol propionate treated culture while slightly elevated over negative controls did not exhibit dose dependency and thus this compound was considered negative in this assay.
Reproductive Studies and Developmental Toxicology
Fertility and Reproduction
Rat
Halobetasol propionate was studied in rats for potential effects on fertility, general reproductive pregnancy and prenatal development. Halobetasol propionate was orally administered at daily dose levels of 0.008, 0.020, and 0.050 mg/kg/day (as a lactose premix suspended in CMC) to study groups of 20 male and 20 female rats. Similarly, groups were treated with the vehicle, and clobetasol propionate was dosed at 0.05 mg/kg/day and served as a reference control. The males were treated for 60 days and the females for 14 days prior to mating until termination of a 12-day mating period. Treatment was continued until Day 15 of pregnancy.
The results from the reproductive study on halobetasol propionate included males and females of the dose groups reacting to the treatment by a reduction in body weight gain and, to a lesser extent, food consumption in a dose-related fashion. Fertility and reproductive performance remained unchanged. At 0.050 mg/kg, some tendency to an increased pre-implantation and, in particular, post-implantation rate of embryonic death was noted. In relation to maternal toxicity, the average weight of the fetuses examined near term was significantly diminished in the three experimental groups, in a dose-related fashion. Parallel to the reduction in fetal body weight, there were indications of delay of skeletal maturation at 0.020 mg/kg and, in particular, 0.050 mg/kg. Occasional anomalies and/or malformations were recorded for all groups. These included two fetuses from one litter of the 0.008 mg/kg dose group and one fetus of the 0.050 mg/kg dose group showing an omphalocele being consistent with a disturbance of ventral closure of the embryo. A "visceral anomaly" dilatation of the pelvic cavity of kidneys was also found in two fetuses from the litter the omphalocele was recorded for. Abnormal ossification of sternebrae was observed in all groups, including the vehicle control.
Based on the results of this study on halobetasol propionate, neither fertility nor general reproductive performance were impaired in the rat under the experimental conditions employed.
The treatment of rats with 0.050 mg/kg of the reference compound, clobetasol propionate, produced results similar to those recorded for halobetasol propionate at doses of 0.020 mg/kg
(body weight of adults) or 0.050 mg/kg (fetal body weight). The live fetuses examined near term exhibited one instance each of cleft palate and dilatation of renal pelvic cavity as well as three instances of abnormal ossification of sternebrae.
Teratology
Oral – Rat
Halobetasol propionate was administered orally at dose levels of 0.008, 0.040, and 0.100 mg/kg/day from days 6 to 15 of pregnancy to study groups of 24 pregnant female rats each. Similar study groups were dosed with the vehicle (0.5% CMC), and a reference control group received clobetasol propionate at 0.100 mg/kg.
The results of this teratology study indicated a high degree of embryotoxicity associated with clobetasol propionate. Halobetasol propionate was, in contrast to the reference compound, devoid of an embryotoxic activity in the rat under the experimental conditions. At 0.040 and 0.100 mg/kg of halobetasol propionate, omphalocele and cleft palate were observed.
Cleft palate was observed to be slightly higher (2.1%) in the high dose group compared to the clobetasol propionate treated group. The omphalocele malformation was at a comparable rate in all drug treated groups. The marginal teratogenic action of halobetasol propionate as well as of clobetasol propionate was associated with maternal toxicity and fetotoxicity.
Oral – Rabbit
In a preliminary teratology study, halobetasol propionate was orally administered at dose levels of 0.01 and 0.04 mg/kg to study groups of 6 pregnant rabbits each from days 6 to 18 of pregnancy. In a similar study group of pregnant rabbits, clobetasol propionate was dosed at 0.02 mg/kg and served as the reference control.
The results from the screening teratology study in rabbits included dose related maternal and embryotoxicity. Halobetasol propionate was teratogenic in the rabbit at a dose producing maternal toxicity (0.01 mg/kg). The type of malformation observed was predominantly cleft palate. Omphalocele and malrotation of the fore limbs were also observed. Clobetasol propionate, which was used as a reference compound, also produced cleft palate and malrotation of fore limbs at a similar incidence. The dose administered (0.02 mg/kg) also induced maternal toxicity.
Failure of palatal closure is well known to occur in the rabbit fetus after treatment of the dams with a variety of corticosteroids. The mechanism of teratogenic action is considered to involve a disturbance of collagen synthesis in the embryonic tissues at a critical phase of development.
No instance of cleft palate was recorded to occur in a cumulative control population of the breed of rabbits used for this experiment.
The teratogenic action of halobetasol propionate as well as of the reference compound clobetasol propionate is attributed to the specific pharmacodynamic properties of these products. In the view of the clear-cut teratogenic effect that could be established in this preliminary study which allows also a qualitative and quantitative comparative assessment of teratogenic potency of the test substance and the reference compound, it was decided by the sponsor upon recommendation by the management of the testing facility not to conduct the planned main study.
Special Toxicology Studies
Special Toxicology studies were performed with the 0.05% Halobetasol Propionate cream and 0.05% Halobetasol Propionate ointment products and not with the lotion 0.01% Halobetasol Propionate
Acute Toxicity Studies on 0.05% Halobetasol Propionate Cream
Rabbit
The ocular irritation potential of 0.05% halobetasol propionate cream or cream vehicle was determined in rabbits. 0.1 g of the test material or vehicle was instilled into the conjunctival eye sac in the eyes of 6 rabbits. After 1 minute, the eyes of 3 rabbits were washed out and all animals graded for irritation at 1, 24, 48 and 72 hours, and at 6 and 7 days. No irritation was observed. 0.05% halobetasol propionate cream was not classified as an eye irritant.
Rabbit
The primary skin irritation potential of 0.05% halobetasol propionate cream was determined in rabbits. Topical applications of 0.5 ml of the test material was applied to the intact or abraded skin sites of rabbits under 24-hour patch occlusion. A primary skin irritation index of 1.21 (mild irritation) was obtained and 0.05% halobetasol propionate cream was not classified as a primary skin irritant.
Rabbit
The dermal irritation potential of 0.05% halobetasol propionate cream or cream vehicle after repeated application was determined in rabbits. The test materials were topically applied at a dose level of 3.0 g to the depilated backs of rabbits (5/sex/group) under 24-hour occlusion daily for 5 days. The fifth dressing was removed after 8 hours. Minimal irritation was observed during the first 2 days and none thereafter. There was a slight body weight increase. There were no mortalities observed during the study.
Rabbit
The dermal irritation potential of 0.05% halobetasol propionate cream, after repeated application for 5 days and a follow-up period of 3 days, was determined in rabbits. The test material was topically applied at 0.5 g under 24-hour patch occlusion to the backs of rabbits (3/sex) for days 1-4, with the fifth dressing removed after 8 hours. Moderate irritation was observed during the study which appeared to clear after 3 days of non-treatment. There was a slight body weight reduction. No mortalities were observed during the study.
Acute Toxicity Studies on 0.05% Halobetasol Propionate Ointment
Rat
The acute oral (LD50) toxicity of 0.05% halobetasol propionate ointment was determined in rats. 0.05%, halobetasol propionate ointment was orally administered undiluted at a dose level of 7.5 mg/kg to a study group of 5/sex. No mortalities or other signs of toxicity were observed. The acute oral (LD50) of 0.05% halobetasol propionate ointment was greater than 7.5 mg/kg.
Rabbit
The ocular irritation potential of 0.05% halobetasol propionate ointment or vehicle was determined in rabbits. 0.1 g of the test material or vehicle was instilled into the conjunctival eye sac in the eyes of 6 rabbits. After 1 minute, the eyes of 3 rabbits were washed out and all animals graded for irritation at 1, 6, 24, 48, and 72 hours, and at 6 and 8 days. No irritation was observed. 0.05% halobetasol propionate ointment was not classified as an eye irritant.
Rabbit
The ocular irritation potential of 0.05% halobetasol propionate ointment was determined in rabbits. 0.1 ml of the test material or vehicle was instilled into the conjunctival eye sac in the eyes of 8 rabbits. After 2 seconds, the eyes of 2 rabbits were washed out, and all animals were graded for irritation at 24, 48, and 72 hours and at 7 days. Slight irritation was observed. 0.05% halobetasol propionate ointment was not classified as an eye irritant.
Rabbit
The primary skin irritation potential of 0.05% halobetasol propionate ointment was determined in rabbits. Topical applications of 0.5 ml of the test material was applied to the intact or abraded skin sites of rabbits under 24-hour patch occlusion. A primary skin irritation index of 0.42 (minimal irritation) was obtained and 0.05% halobetasol propionate ointment was not classified as a primary skin irritant.
Rabbit
The dermal irritation potential of 0.05% halobetasol propionate ointment or ointment vehicle after repeated application was determined in rabbits. The test materials were topically applied at a dose level of 5.0 g to the depilated backs of rabbits (3/sex/group) under 24-hour occlusion daily for 5 days. The fifth dressing was removed after 8 hours. There was a slight body weight decrease. There were no mortalities or irritation observed during the study.
Sensitization Studies on 0.05% Halobetasol Propionate Cream and Ointment
Guinea Pig
The sensitization potential of 0.05% halobetasol propionate cream or vehicle was determined in guinea pigs. To groups of 10/sex/group, topical applications of the test materials were made under occlusion on sites injected with adjuvant. At the second induction during week 2, the test materials were reapplied under occlusion on the sites pretreated with a 10% sodium lauryl sulfate. Challenge applications to test and control study groups were made 14 days later. Under the conditions of the modified maximization test in guinea pigs, 0.05% halobetasol propionate cream was non-sensitizing.
Guinea Pig
The sensitization potential of 0.05% halobetasol propionate ointment or vehicle was determined in guinea pigs. To groups of 10/sex/group, topical applications of the test materials were made under occlusion on sites injected with adjuvant. At the second induction during week 2, the test materials were reapplied under occlusion on the sites pretreated with 10% sodium lauryl sulfate. Challenge applications to test and control study groups were made 14 days later. Under the conditions of the modified maximization test in guinea pigs, 0.05% halobetasol propionate ointment was non-sensitizing.
Subacute Toxicity Studies on Halobetasol Propionate (as pure compound and ointment)
Oral – Rat
The subacute oral toxicity potential of halobetasol propionate was determined in rats. Halobetasol propionate was administered orally by gavage at dose levels of 0.01, 0.1 and 1.0 mg/kg/day (in 0.5% CMC) at a dose volume of 10 ml/kg to study groups of 10 or 15/sex/group for 3 months. A similar test group of animals received the vehicle, 0.5% CMC, and served as a control. At termination of treatment, a one-month recovery period was conducted involving 5/sex/group from the 0.1 and 1.0 mg/kg dose groups and of the vehicle control group.
During the study, 1 male rat of the high dose group was found dead (Day 40). Test material related changes included piloerection, reduction of spontaneous activity, muscle hypotonia, and loss of hair in all dose groups. These symptoms except for hair loss were, in general, reversible. Body weights were markedly reduced in all dose levels, with some improvement in weight gain during recovery. Food consumption values were similarly reduced during the study. Clinical chemistry effects included increased but reversible ALAT (SGPT) values in the mid and high dose groups. Hematological changes were reduced packed cell volume and dose related lymphopenia and neutrophilia which were reversible during recovery. Urinalysis revealed blood and protein in the urine which were present after the recovery. Organ weights of the liver, heart and kidney were increased, with decreased adrenal, spleen and thymus weights. The organ weight changes in the mid and high dose groups were generally correlated with microscopic evaluations.
Histopathological examination revealed expected dose related corticosteroid related changes including ballooned hepatocytes, with marked adrenal, thymic and splenic atrophy, and respiratory tract infections. Most changes were reduced or reversible after recovery. On the basis of numerous observed effects including body weight reduction, minimal liver effects, reduced lymphatic elements in axillary lymph nodes in the 0.01 mg/kg dose group, the no effect level is less than 0.01 mg/kg/day.
In a study performed in parallel to the above, the reference control steroid, clobetasol propionate, was similarly evaluated for subacute oral toxicity potential at a dose level of 0.1 mg/kg/day to a study group of 15/sex for 3 months, followed by a one-month recovery period. Test material effects were similar in type and comparable to the steroid related effects reported on halobetasol propionate (at 0.1 mg/kg). The effects produced were reversible.
Oral – Dog
The subacute oral toxicity potential of halobetasol propionate was determined in beagle dogs. Halobetasol propionate was administered orally via gelatin capsules to dogs at dose levels of 0.01, 0.03, and 0.1 mg/kg/day to study groups of 3/sex/group (low and mid dose) and 6/sex/group (high dose) for 3 months. The test material was administered as a premix with lactose. A control group of 3/sex was administered 10 mg/kg/day of lactose in gelatin capsules.
At termination of treatment, a 1-month recovery period was conducted involving 3/sex/group of the high dose group dogs.
No mortalities related to test material administration were observed during the study. Changes associated with administration of halobetasol propionate included general signs of soft, bloody feces in the high dose group, diarrhea, decreased body weight in the high dose, no changes in food consumption, increased but reversible ALAT levels in mid and high dose levels, increased alkaline phosphatase values at high dose, marked with reversible dose dependent increase of an alpha 3 globulins with a slight decrease in other globulins, decreased but slightly reversible dose related cortisol levels, increased but reversible triglycerides in the high dose group, hematological changes included decreased hemoglobin, rbc, and hematocrit in the high dose group, dose related lymphopenia and neutrophilia, with eosinopenia, and bacteria in urine (high dose). Organ weight changes included reduced adrenals, non or minimally reversible, increased liver, increased spleen and kidneys (high dose). Histopathological examination revealed marked atrophy of the adrenal and lymphatic tissues, ballooning and/or vacuolization of the hepatocytes, and inflammation of the urogenital tract. Most changes were dose dependent or occurring at the high dose and were reduced or reversible following recovery other than the adrenals. There was no ocular, auditory, or neurological changes observed during the study.
In the high dose group (0.1 mg/kg), an increased incidence of slight deviations in repolarization (diphasic or notched T-wave, reversion of polarity) was detected in 11 of 12 animals. Alternations continued through the one-month recovery period in 3 of 6 animals.
Clobetasol propionate, the reference steroid, was similarly evaluated in a 3-month subacute oral toxicity study at a dose level of 0.03 mg/kg/day (as a lactose premix) in gelatin capsules to a study group of 3 males and female dogs. 2/3 male dogs showed variations of the T-wave polarity.
The dogs receiving oral doses of both steroids reacted to the treatment by developing stress along with a suppression of the lymphatic system and apparently increased susceptibility to infectious diseases in a dose related manner. The deviation in the form of the T-Waves was therefore not considered to indicate drug related cardiomyopathy.
In the absence of cardiac lesions found at necropsy, the absence of electrocardiographic changes other than non-specific T-wave changes, presence of toxic effects that can produce non-specific T-wave effects in dogs, the electrocardiographic findings are not considered indicative of cardiotoxicity.
Other results of this study revealed comparable steroid related effects of clobetasol propionate to that of halobetasol propionate, other than a decrease in body weight present in the clobetasol propionate group and in liver changes which were more pronounced in the dogs treated with halobetasol propionate.
Dermal – Rat
The subacute dermal toxicity potential of halobetasol propionate ointment was determined in rats for 3 months. The test material was topically administered to study groups of 6/sex/group for 3 months at daily dose levels of 0.05%, 0.1% and 0.2% halobetasol propionate ointment at a dose volume of 400 mg/kg of ointment (equivalent to 0.2, 0.4, and 0.8 mg/kg/day of halobetasol propionate). Control groups were similarly treated with the vehicle ointment and 0.05% clobetasol propionate ointment served as a reference control. Separate study groups of 4/sex/group of rats were also treated with test or control ointments for 3 months and were maintained on study for 1 month without treatment for recovery.
During the study there were no treatment related mortalities observed in the rats. Test material related changes observed during the study included skin changes (red/blue colouration), decreased body weights in the males, decreased food consumption, increased ALAT (SGPT) levels, hematological changes consisting of lymphopenia and neutrophilia, slight anemia, hematuria, and hypo gamma globulinemia, the organ weight of the adrenals and thymus were reduced. At necropsy, the condition of the animals was considered cachectic. Histopathological examination revealed expected steroid related changes including epidermal thinning, distended macrophages in the lungs, vacuolated and distended hepatocytes of the liver, moderate adrenocortical atrophy and atrophy of the lymphoid organs, and hyperplasia of the islets of Langerhans in the pancreas.
The changes observed were, in general, dose related and reversible or reduced by the end of the recovery study. Similar changes, which were less severe, were observed in the 0.05% clobetasol propionate ointment test groups.
Dermal – Dog
The subacute dermal toxicity potential of halobetasol propionate ointment was determined in dogs following topical administrations for 3 months. The test material was topically administered to study groups of 6/sex/group for 3 months at daily dose levels of 0.05%, 0.1%, and 0.2% halobetasol propionate ointment at a dose volume of 400 mg/kg/day of ointment (equivalent to 0.2, 0.4, and 0.8 mg/kg/day of halobetasol propionate). All application sites were occluded daily for 6 hours. Control groups were similarly treated with the vehicle ointment, and 0.05% clobetasol propionate ointment served as a reference control. After 3 months, study groups of 2/sex/group of dogs were maintained on study for 1 month without treatment for recovery.
During the study, 1 dog in the mid and high dose was killed due to severely infected skin wounds. Another dog was killed in the low dose due to self-inflicted injury. Test material related changes observed during the study included skin lesions, erythema, hair loss, papules/scabs and abscesses, no remarkable body weight or food consumption changes, increased ALAT, alkaline phosphatase, and alpha 2 globulins, decreased cortisol, with hypogammaglobulinemia, dose related anemia in the halobetasol propionate treated animals, hematological changes included increased ESR, lymphopenia and neutrophilia, eosinopenia, hematuria and hemoglobinuria, with no other changes observed during ophthalmic, auditory or electrocardiac examinations. Organ weight changes revealed decreased adrenals and increased liver weights. Histopathological examination revealed expected steroid related changes consisting of adrenocortical and marked thymic atrophy, cytoplasmic vacuolation of the hepatocytes, dermal changes were thinning of the stratum corneum and focal panniculus myopathy of the skin, and an increased incidence of granulomatous reactions associated with helminth larvae in the lungs, liver and lymph nodes.
Most changes observed were dose related and reversible or reduced following a 1-month recovery period. Comparable changes were observed in the 0.05% clobetasol propionate ointment dose group.
Study evaluating the dermal and systemic toxicity of an enhanced strength of halobetasol propionate lotion (0.02%)
Göttingen minipigs were dosed dermally with halobetasol propionate lotion (enhanced) once daily for 90/91 days to 10% total BSA at 0.01 mL/cm2 (n = 3/sex). The selected application rate covered the entire skin dose site without lotion run off. Control groups included sham (n = 2/sex) and vehicle (n = 4/sex); statistical comparisons were made against the vehicle group. The endpoints evaluated included clinical signs, dermal scores, body weights, food consumption, clinical pathology, toxicokinetics, ophthalmology, electrocardiography, gross necropsy findings, organ weights and histopathology.
All animals survived to scheduled termination. In the sham, vehicle, and halobetasol propionate lotion (enhanced) groups, there were only some instances of transient, very slight skin irritation. A moderate decrease in mean body weights (17% to 19% on Day 89) was observed in the halobetasol propionate lotion (enhanced) group when compared with the vehicle group, with no gender differences. There were no table food consumption differences between groups. Likewise, there were no toxicologically meaningful differences in hematology, coagulation, clinical chemistry, electrocardiography, or ophthalmology data.
Blue discoloration of the nose/snout was noted in the halobetasol propionate lotion (enhanced) group, which is attributed to halobetasol propionate as other test article groups containing HP evaluated in this study presented the same observations.
Adrenal absolute weights were decreased in males and females treated with halobetasol propionate lotion (enhanced), compared with vehicle. Thymus absolute weights were also decreased, respectively. In most cases, both adrenal and thymus relative organ weights were lower when compared with vehicle.
Microscopically, halobetasol propionate lotion (enhanced) administration was associated with minimal hyperkeratosis in the skin and minimal to moderate cortical atrophy in the adrenal glands of all animals. Cortical adrenal atrophy correlated with decreased adrenal weights and was characterized microscopically by decreased size of zona fasciculata cells, resulting in an overall decreased thickness of the adrenal cortex. There were no other microscopic test article-related findings.